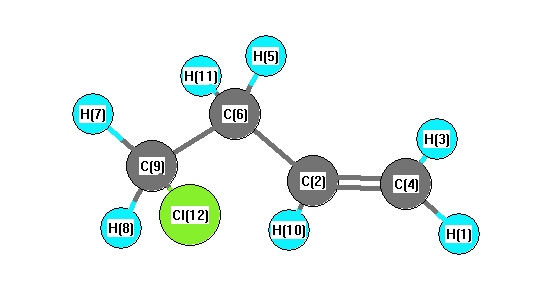
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3166 |
3147 |
12.73 |
|
|
|
| 2 |
A |
3089 |
3071 |
3.58 |
|
|
|
| 3 |
A |
3078 |
3060 |
8.57 |
|
|
|
| 4 |
A |
3066 |
3048 |
8.94 |
|
|
|
| 5 |
A |
3014 |
2997 |
26.42 |
|
|
|
| 6 |
A |
2999 |
2982 |
3.80 |
|
|
|
| 7 |
A |
2954 |
2937 |
15.45 |
|
|
|
| 8 |
A |
1672 |
1662 |
6.69 |
|
|
|
| 9 |
A |
1405 |
1397 |
2.16 |
|
|
|
| 10 |
A |
1393 |
1385 |
10.82 |
|
|
|
| 11 |
A |
1375 |
1367 |
3.21 |
|
|
|
| 12 |
A |
1299 |
1291 |
1.49 |
|
|
|
| 13 |
A |
1267 |
1260 |
10.50 |
|
|
|
| 14 |
A |
1250 |
1243 |
27.02 |
|
|
|
| 15 |
A |
1208 |
1201 |
0.58 |
|
|
|
| 16 |
A |
1132 |
1126 |
0.37 |
|
|
|
| 17 |
A |
1096 |
1090 |
6.84 |
|
|
|
| 18 |
A |
1002 |
997 |
5.25 |
|
|
|
| 19 |
A |
994 |
988 |
13.77 |
|
|
|
| 20 |
A |
964 |
959 |
3.74 |
|
|
|
| 21 |
A |
898 |
893 |
29.92 |
|
|
|
| 22 |
A |
871 |
866 |
15.11 |
|
|
|
| 23 |
A |
794 |
790 |
6.63 |
|
|
|
| 24 |
A |
651 |
647 |
22.49 |
|
|
|
| 25 |
A |
644 |
641 |
14.06 |
|
|
|
| 26 |
A |
443 |
441 |
2.34 |
|
|
|
| 27 |
A |
342 |
340 |
0.42 |
|
|
|
| 28 |
A |
240 |
239 |
1.83 |
|
|
|
| 29 |
A |
119 |
118 |
2.22 |
|
|
|
| 30 |
A |
76 |
75 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21251.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 21127.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.030 |
|
|
|
| 2 |
C |
-0.082 |
|
|
|
| 3 |
H |
0.018 |
|
|
|
| 4 |
C |
-0.005 |
|
|
|
| 5 |
H |
0.053 |
|
|
|
| 6 |
C |
0.009 |
|
|
|
| 7 |
H |
0.076 |
|
|
|
| 8 |
H |
0.083 |
|
|
|
| 9 |
C |
-0.082 |
|
|
|
| 10 |
H |
0.008 |
|
|
|
| 11 |
H |
0.026 |
|
|
|
| 12 |
Cl |
-0.135 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.140 |
1.414 |
-0.228 |
1.830 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.533 |
-1.053 |
-0.530 |
| y |
-1.053 |
-35.806 |
-0.445 |
| z |
-0.530 |
-0.445 |
-38.869 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.196 |
-1.053 |
-0.530 |
| y |
-1.053 |
3.895 |
-0.445 |
| z |
-0.530 |
-0.445 |
-0.699 |
|
| Polar |
|---|
| 3z2-r2 | -1.398 |
|---|
| x2-y2 | -4.727 |
|---|
| xy | -1.053 |
|---|
| xz | -0.530 |
|---|
| yz | -0.445 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.970 |
0.307 |
-0.976 |
| y |
0.307 |
7.964 |
-0.152 |
| z |
-0.976 |
-0.152 |
5.712 |
<r2> (average value of r
2) Å
2
| <r2> |
202.277 |
| (<r2>)1/2 |
14.222 |