return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (3-Hexyne)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-233.889366
Energy at 298.15K-233.898451
HF Energy-232.992601
Nuclear repulsion energy205.248317
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3154 3026 0.00      
2 A 3147 3020 9.34      
3 A 3103 2977 0.00      
4 A 3065 2940 0.15      
5 A 3060 2936 59.66      
6 A 2340 2245 0.03      
7 A 1503 1442 1.35      
8 A 1493 1432 0.00      
9 A 1483 1423 2.38      
10 A 1418 1360 1.11      
11 A 1365 1309 0.07      
12 A 1293 1240 0.00      
13 A 1104 1059 1.55      
14 A 1103 1058 0.00      
15 A 1016 975 0.00      
16 A 786 754 0.00      
17 A 696 668 0.10      
18 A 423 405 6.03      
19 A 373 358 0.00      
20 A 234 224 0.00      
21 A 89 85 1.44      
22 A 7 7 0.00      
23 B 3155 3027 58.43      
24 B 3147 3019 60.20      
25 B 3103 2977 19.48      
26 B 3064 2940 55.66      
27 B 3061 2936 0.82      
28 B 1503 1442 0.85      
29 B 1493 1433 9.77      
30 B 1481 1421 2.17      
31 B 1417 1360 0.96      
32 B 1371 1315 45.40      
33 B 1296 1244 0.00      
34 B 1173 1125 0.01      
35 B 1120 1075 0.98      
36 B 1094 1050 2.43      
37 B 944 906 0.03      
38 B 789 757 2.03      
39 B 536 515 0.41      
40 B 294 282 2.99      
41 B 208 200 0.09      
42 B 156 149 4.17      

Unscaled Zero Point Vibrational Energy (zpe) 31329.5 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 30057.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.30891 0.04604 0.04132

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.000 0.613 -0.573
C2 0.000 -0.613 -0.573
C3 0.000 2.094 -0.553
C4 -0.000 -2.094 -0.553
C5 -0.001 2.666 0.876
C6 0.001 -2.666 0.876
H7 0.886 2.462 -1.104
H8 -0.886 2.462 -1.105
H9 -0.886 -2.462 -1.104
H10 0.886 -2.462 -1.105
H11 -0.001 3.770 0.852
H12 -0.895 2.329 1.430
H13 0.893 2.328 1.431
H14 0.001 -3.770 0.852
H15 0.895 -2.329 1.430
H16 -0.893 -2.328 1.431

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.22561.48152.70702.51313.58462.11822.11823.24373.24373.46422.78552.78554.60893.66963.6696
C21.22562.70701.48153.58462.51313.24373.24372.11822.11824.60893.66963.66963.46422.78552.7855
C31.48152.70704.18831.53964.96991.10631.10634.67424.67422.18712.18862.18866.03044.92904.9290
C42.70701.48154.18834.96991.53964.67424.67421.10631.10636.03044.92904.92902.18712.18862.1886
C52.51313.58461.53964.96995.33132.17912.17915.56755.56811.10491.10451.10456.43605.10405.1034
C63.58462.51314.96991.53965.33135.56755.56812.17912.17916.43605.10405.10341.10491.10451.1045
H72.11823.24371.10634.67422.17915.56751.77225.23364.92422.51433.10012.53836.59165.41945.7043
H82.11823.24371.10634.67422.17915.56811.77224.92425.23322.51432.53833.10016.59215.70495.4204
H93.24372.11824.67421.10635.56752.17915.23364.92421.77226.59165.41945.70432.51433.10012.5383
H103.24372.11824.67421.10635.56812.17914.92425.23321.77226.59215.70495.42042.51432.53833.1001
H113.46424.60892.18716.03041.10496.43602.51432.51436.59166.59211.79241.79247.54056.19136.1908
H122.78553.66962.18864.92901.10455.10403.10012.53835.41945.70491.79241.78746.19134.98914.6570
H132.78553.66962.18864.92901.10455.10342.53833.10015.70435.42041.79241.78746.19084.65704.9874
H144.60893.46426.03042.18716.43601.10496.59166.59212.51432.51437.54056.19136.19081.79241.7924
H153.66962.78554.92902.18865.10401.10455.41945.70493.10012.53836.19134.98914.65701.79241.7874
H163.66962.78554.92902.18865.10341.10455.70435.42042.53833.10016.19084.65704.98741.79241.7874

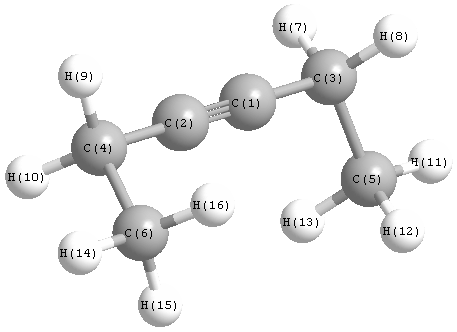
picture of 3-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 179.224 C1 C3 C5 112.566
C1 C3 H7 109.014 C1 C3 H8 109.014
C2 C1 C3 179.224 C2 C4 C6 112.566
C2 C4 H9 109.014 C2 C4 H10 109.014
C3 C5 H11 110.517 C3 C5 H12 110.660
C3 C5 H13 110.661 C4 C6 H14 110.517
C4 C6 H15 110.660 C4 C6 H16 110.661
C5 C3 H7 109.804 C5 C3 H8 109.804
C6 C4 H9 109.804 C6 C4 H10 109.804
H7 C3 H8 106.447 H9 C4 H10 106.447
H11 C5 H12 108.442 H11 C5 H13 108.442
H12 C5 H13 108.035 H14 C6 H15 108.442
H14 C6 H16 108.442 H15 C6 H15 0.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability