return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6O (3-Methylfuran)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-268.612408
Energy at 298.15K-268.618948
HF Energy-267.680912
Nuclear repulsion energy219.000681
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3290 3136        
2 A' 3279 3125        
3 A' 3253 3101        
4 A' 3133 2986        
5 A' 3041 2899        
6 A' 1651 1573        
7 A' 1547 1475        
8 A' 1488 1419        
9 A' 1419 1353        
10 A' 1403 1337        
11 A' 1281 1221        
12 A' 1252 1193        
13 A' 1204 1147        
14 A' 1088 1037        
15 A' 1044 995        
16 A' 1006 959        
17 A' 967 921        
18 A' 879 838        
19 A' 637 607        
20 A' 309 294        
21 A" 3115 2969        
22 A" 1473 1404        
23 A" 1052 1003        
24 A" 818 779        
25 A" 767 731        
26 A" 717 684        
27 A" 604 576        
28 A" 595 568        
29 A" 258 246        
30 A" 120 115        

Unscaled Zero Point Vibrational Energy (zpe) 21345.7 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 20344.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.29213 0.11023 0.08127

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.698 -1.478 0.000
C2 -1.085 -0.160 0.000
C3 0.672 -1.473 0.000
C4 0.000 0.688 0.000
C5 1.161 -0.188 0.000
C6 -0.011 2.196 0.000
H7 -2.161 0.019 0.000
H8 1.150 -2.454 0.000
H9 2.214 0.105 0.000
H10 -1.047 2.582 0.000
H11 0.503 2.599 0.893
H12 0.503 2.599 -0.893

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
O11.37401.37032.27542.26293.73742.09322.08983.31464.07504.34304.3430
C21.37402.19401.37652.24652.58841.09113.20293.30942.74193.30583.3058
C31.37032.19402.26311.37493.73233.20241.09072.20664.40484.17254.1725
C42.27541.37652.26311.45481.50812.26243.34542.28922.16452.16882.1688
C52.26292.24651.37491.45482.65713.32912.26551.09273.54313.00013.0001
C63.73742.58843.73231.50812.65713.05954.79253.05321.10531.10671.1067
H72.09321.09113.20242.26243.32913.05954.13264.37612.79493.81483.8148
H82.08983.20291.09073.34542.26554.79254.13262.77175.49445.17185.1718
H93.31463.30942.20662.28921.09273.05324.37612.77174.09493.15323.1532
H104.07502.74194.40482.16453.54311.10532.79495.49444.09491.78911.7891
H114.34303.30584.17252.16883.00011.10673.81485.17183.15321.78911.7861
H124.34303.30584.17252.16883.00011.10673.81485.17183.15321.78911.7861

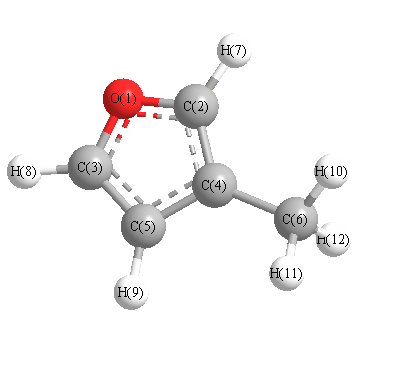
picture of 3-Methylfuran state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C4 111.635 O1 C2 H7 115.761
O1 C3 C5 111.038 O1 C3 H8 115.777
C2 O1 C3 106.157 C2 C4 C5 104.984
C2 C4 C6 127.550 C3 C5 C4 106.186
C3 C5 H9 126.441 C4 C2 H7 132.604
C4 C5 H9 127.373 C4 C6 H10 110.893
C4 C6 H11 111.144 C4 C6 H12 111.144
C5 C3 H8 133.185 C5 C4 C6 127.466
H10 C6 H11 107.955 H10 C6 H12 107.955
H11 C6 H12 107.598
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability