return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6O (3-Methylfuran)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-268.578640
Energy at 298.15K-268.585283
HF Energy-267.682397
Nuclear repulsion energy219.580562
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3310 3176 1.00      
2 A' 3299 3165 1.25      
3 A' 3273 3140 1.39      
4 A' 3148 3020 18.88      
5 A' 3058 2934 33.38      
6 A' 1685 1616 0.51      
7 A' 1578 1514 17.96      
8 A' 1500 1439 5.25      
9 A' 1436 1378 3.15      
10 A' 1420 1362 1.45      
11 A' 1295 1243 5.19      
12 A' 1268 1216 1.39      
13 A' 1223 1173 23.05      
14 A' 1101 1056 14.25      
15 A' 1059 1016 23.59      
16 A' 1019 977 2.57      
17 A' 976 937 2.04      
18 A' 891 854 21.90      
19 A' 644 618 2.49      
20 A' 314 301 0.81      
21 A" 3130 3003 20.07      
22 A" 1483 1422 5.03      
23 A" 1065 1021 0.83      
24 A" 853 818 0.39      
25 A" 794 762 50.18      
26 A" 748 717 5.69      
27 A" 614 589 5.85      
28 A" 610 585 13.82      
29 A" 264 253 2.93      
30 A" 132 127 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 21592.9 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 20716.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.29370 0.11078 0.08169

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -0.700 -1.471 0.000
C2 -1.082 -0.155 0.000
C3 0.666 -1.471 0.000
C4 0.000 0.685 0.000
C5 1.158 -0.195 0.000
C6 -0.004 2.191 0.000
H7 -2.156 0.028 0.000
H8 1.140 -2.452 0.000
H9 2.210 0.095 0.000
H10 -1.036 2.583 0.000
H11 0.512 2.591 0.892
H12 0.512 2.591 -0.892

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
O11.37041.36602.26702.25443.72782.08902.08553.30474.06784.33224.3322
C21.37042.18791.36972.24022.58181.08933.19583.30122.73813.29813.2981
C31.36602.18792.25671.36823.72333.19491.08912.19954.39694.16244.1624
C42.26701.36972.25671.45421.50622.25403.33742.28712.16242.16602.1660
C52.25442.24021.36821.45422.65393.32132.25691.09103.53972.99602.9960
C63.72782.58183.72331.50622.65393.05154.78183.04891.10391.10531.1053
H72.08901.08933.19492.25403.32133.05154.12444.36622.79003.80603.8060
H82.08553.19581.08913.33742.25694.78184.12442.76205.48485.15975.1597
H93.30473.30122.19952.28711.09103.04894.36622.76204.08973.14823.1482
H104.06782.73814.39692.16243.53971.10392.79005.48484.08971.78651.7865
H114.33223.29814.16242.16602.99601.10533.80605.15973.14821.78651.7836
H124.33223.29814.16242.16602.99601.10533.80605.15973.14821.78651.7836

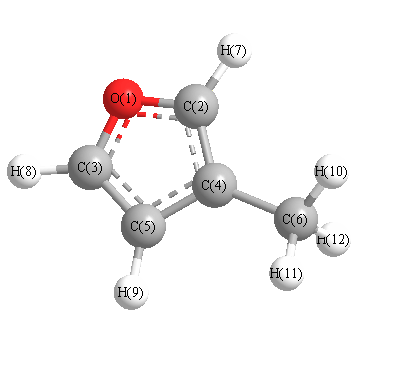
picture of 3-Methylfuran state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C4 111.653 O1 C2 H7 115.801
O1 C3 C5 111.076 O1 C3 H8 115.849
C2 O1 C3 106.179 C2 C4 C5 104.952
C2 C4 C6 127.658 C3 C5 C4 106.139
C3 C5 H9 126.490 C4 C2 H7 132.545
C4 C5 H9 127.371 C4 C6 H10 110.944
C4 C6 H11 111.149 C4 C6 H12 111.149
C5 C3 H8 133.074 C5 C4 C6 127.390
H10 C6 H11 107.934 H10 C6 H12 107.934
H11 C6 H12 107.577
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability