return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H4O2 (4-Cyclopentene-1,3-dione)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-342.443017
Energy at 298.15K-342.447676
Nuclear repulsion energy265.338471
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3260 3128 1.18      
2 A1 3104 2978 0.08      
3 A1 1855 1780 24.20      
4 A1 1652 1584 2.27      
5 A1 1421 1363 18.45      
6 A1 1276 1224 121.02      
7 A1 1068 1025 9.35      
8 A1 857 823 1.82      
9 A1 572 549 3.43      
10 A1 388 372 15.54      
11 A2 1145 1099 0.00      
12 A2 997 957 0.00      
13 A2 646 620 0.00      
14 A2 242 232 0.00      
15 B1 3160 3032 0.43      
16 B1 967 927 23.09      
17 B1 803 770 23.41      
18 B1 441 423 0.13      
19 B1 84 80 13.68      
20 B2 3237 3106 0.52      
21 B2 1821 1747 401.75      
22 B2 1354 1299 11.97      
23 B2 1252 1201 0.22      
24 B2 1150 1103 1.30      
25 B2 832 798 3.05      
26 B2 691 663 0.83      
27 B2 521 500 0.88      

Unscaled Zero Point Vibrational Energy (zpe) 17398.0 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 16691.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.21962 0.07235 0.05499

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.095
C2 0.000 1.210 0.152
C3 0.000 -1.210 0.152
C4 0.000 0.677 -1.258
C5 0.000 -0.677 -1.258
O6 0.000 2.383 0.467
O7 0.000 -2.383 0.467
H8 0.892 0.000 1.744
H9 -0.892 0.000 1.744
H10 0.000 1.341 -2.128
H11 0.000 -1.341 -2.128

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 O7 H8 H9 H10 H11
C11.53351.53352.44822.44822.46402.46401.10361.10363.49053.4905
C21.53352.41941.50732.35551.21473.60632.18972.18972.28373.4214
C31.53352.41942.35551.50733.60631.21472.18972.18973.42142.2837
C42.44821.50732.35551.35432.42613.51283.20463.20461.09442.1979
C52.44822.35551.50731.35433.51282.42613.20463.20462.19791.0944
O62.46401.21473.60632.42613.51284.76572.84692.84692.79624.5391
O72.46403.60631.21473.51282.42614.76572.84692.84694.53912.7962
H81.10362.18972.18973.20463.20462.84692.84691.78384.19404.1940
H91.10362.18972.18973.20463.20462.84692.84691.78384.19404.1940
H103.49052.28373.42141.09442.19792.79624.53914.19404.19402.6827
H113.49053.42142.28372.19791.09444.53912.79624.19404.19402.6827

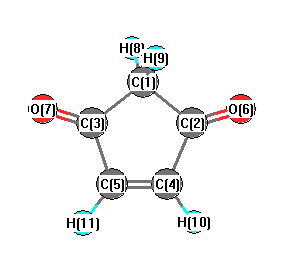
picture of 4-Cyclopentene-1,3-dione state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 107.233 C1 C2 O6 127.041
C1 C3 C5 107.233 C1 C3 O7 127.041
C2 C1 C3 104.152 C2 C1 H8 111.221
C2 C1 H9 111.221 C2 C4 C5 110.690
C2 C4 H10 121.943 C3 C1 H8 111.221
C3 C1 H9 111.221 C3 C5 C4 110.690
C3 C5 H11 121.943 C4 C2 O6 125.726
C4 C5 H11 127.367 C5 C3 O7 125.726
C5 C4 H10 127.367 H8 C1 H9 107.838
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability