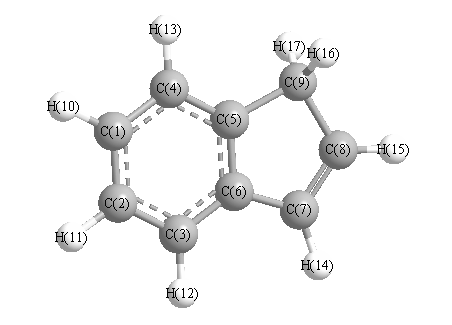
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3160 |
3142 |
12.77 |
|
|
|
| 2 |
A' |
3135 |
3117 |
5.14 |
|
|
|
| 3 |
A' |
3133 |
3114 |
20.60 |
|
|
|
| 4 |
A' |
3120 |
3101 |
26.57 |
|
|
|
| 5 |
A' |
3108 |
3090 |
2.42 |
|
|
|
| 6 |
A' |
3101 |
3083 |
5.34 |
|
|
|
| 7 |
A' |
2957 |
2940 |
10.47 |
|
|
|
| 8 |
A' |
1620 |
1611 |
2.08 |
|
|
|
| 9 |
A' |
1600 |
1591 |
0.64 |
|
|
|
| 10 |
A' |
1564 |
1555 |
0.48 |
|
|
|
| 11 |
A' |
1450 |
1442 |
8.30 |
|
|
|
| 12 |
A' |
1447 |
1438 |
4.58 |
|
|
|
| 13 |
A' |
1391 |
1383 |
1.93 |
|
|
|
| 14 |
A' |
1345 |
1337 |
11.39 |
|
|
|
| 15 |
A' |
1317 |
1309 |
7.10 |
|
|
|
| 16 |
A' |
1266 |
1259 |
0.03 |
|
|
|
| 17 |
A' |
1215 |
1208 |
1.45 |
|
|
|
| 18 |
A' |
1189 |
1182 |
4.35 |
|
|
|
| 19 |
A' |
1143 |
1137 |
0.55 |
|
|
|
| 20 |
A' |
1131 |
1124 |
0.00 |
|
|
|
| 21 |
A' |
1091 |
1085 |
0.84 |
|
|
|
| 22 |
A' |
1042 |
1036 |
1.70 |
|
|
|
| 23 |
A' |
1019 |
1013 |
4.28 |
|
|
|
| 24 |
A' |
946 |
940 |
10.30 |
|
|
|
| 25 |
A' |
846 |
841 |
3.01 |
|
|
|
| 26 |
A' |
821 |
816 |
1.57 |
|
|
|
| 27 |
A' |
724 |
720 |
2.40 |
|
|
|
| 28 |
A' |
583 |
580 |
1.54 |
|
|
|
| 29 |
A' |
528 |
525 |
0.13 |
|
|
|
| 30 |
A' |
377 |
375 |
1.32 |
|
|
|
| 31 |
A" |
2990 |
2973 |
5.95 |
|
|
|
| 32 |
A" |
1088 |
1081 |
1.37 |
|
|
|
| 33 |
A" |
957 |
952 |
0.02 |
|
|
|
| 34 |
A" |
923 |
918 |
1.83 |
|
|
|
| 35 |
A" |
919 |
914 |
0.50 |
|
|
|
| 36 |
A" |
894 |
888 |
2.71 |
|
|
|
| 37 |
A" |
840 |
835 |
0.06 |
|
|
|
| 38 |
A" |
760 |
756 |
34.30 |
|
|
|
| 39 |
A" |
709 |
705 |
17.87 |
|
|
|
| 40 |
A" |
686 |
682 |
10.09 |
|
|
|
| 41 |
A" |
547 |
544 |
3.64 |
|
|
|
| 42 |
A" |
411 |
409 |
6.81 |
|
|
|
| 43 |
A" |
381 |
379 |
5.04 |
|
|
|
| 44 |
A" |
206 |
204 |
3.51 |
|
|
|
| 45 |
A" |
190 |
189 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29934.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29761.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.027 |
|
|
|
| 2 |
C |
0.033 |
|
|
|
| 3 |
C |
-0.024 |
|
|
|
| 4 |
C |
-0.018 |
|
|
|
| 5 |
C |
-0.078 |
|
|
|
| 6 |
C |
0.076 |
|
|
|
| 7 |
C |
0.038 |
|
|
|
| 8 |
C |
-0.063 |
|
|
|
| 9 |
C |
0.064 |
|
|
|
| 10 |
H |
-0.024 |
|
|
|
| 11 |
H |
-0.023 |
|
|
|
| 12 |
H |
-0.040 |
|
|
|
| 13 |
H |
-0.043 |
|
|
|
| 14 |
H |
-0.034 |
|
|
|
| 15 |
H |
-0.020 |
|
|
|
| 16 |
H |
0.063 |
|
|
|
| 17 |
H |
0.063 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.450 |
0.620 |
0.000 |
0.766 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.245 |
0.682 |
0.000 |
| y |
0.682 |
-46.830 |
0.000 |
| z |
0.000 |
0.000 |
-56.344 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.342 |
0.682 |
0.000 |
| y |
0.682 |
5.464 |
0.000 |
| z |
0.000 |
0.000 |
-8.806 |
|
| Polar |
|---|
| 3z2-r2 | -17.613 |
|---|
| x2-y2 | -1.415 |
|---|
| xy | 0.682 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
19.476 |
1.749 |
0.000 |
| y |
1.749 |
16.362 |
0.000 |
| z |
0.000 |
0.000 |
6.005 |
<r2> (average value of r
2) Å
2
| <r2> |
296.911 |
| (<r2>)1/2 |
17.231 |