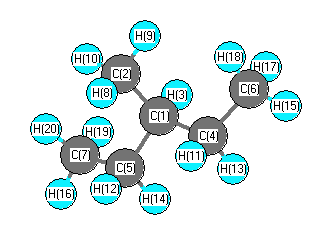
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3051 |
3033 |
61.73 |
|
|
|
| 2 |
A' |
3043 |
3025 |
40.35 |
|
|
|
| 3 |
A' |
3040 |
3022 |
40.11 |
|
|
|
| 4 |
A' |
2986 |
2969 |
48.83 |
|
|
|
| 5 |
A' |
2966 |
2949 |
14.16 |
|
|
|
| 6 |
A' |
2961 |
2944 |
22.88 |
|
|
|
| 7 |
A' |
2944 |
2927 |
70.01 |
|
|
|
| 8 |
A' |
2910 |
2893 |
14.12 |
|
|
|
| 9 |
A' |
1433 |
1424 |
3.48 |
|
|
|
| 10 |
A' |
1429 |
1421 |
9.35 |
|
|
|
| 11 |
A' |
1417 |
1409 |
10.63 |
|
|
|
| 12 |
A' |
1408 |
1399 |
2.53 |
|
|
|
| 13 |
A' |
1348 |
1340 |
1.93 |
|
|
|
| 14 |
A' |
1344 |
1336 |
5.24 |
|
|
|
| 15 |
A' |
1319 |
1312 |
1.14 |
|
|
|
| 16 |
A' |
1302 |
1295 |
0.08 |
|
|
|
| 17 |
A' |
1249 |
1241 |
1.14 |
|
|
|
| 18 |
A' |
1147 |
1141 |
0.18 |
|
|
|
| 19 |
A' |
1143 |
1137 |
0.69 |
|
|
|
| 20 |
A' |
1057 |
1051 |
0.08 |
|
|
|
| 21 |
A' |
949 |
944 |
3.31 |
|
|
|
| 22 |
A' |
923 |
918 |
1.93 |
|
|
|
| 23 |
A' |
798 |
793 |
0.91 |
|
|
|
| 24 |
A' |
751 |
747 |
4.93 |
|
|
|
| 25 |
A' |
429 |
426 |
0.21 |
|
|
|
| 26 |
A' |
378 |
376 |
0.15 |
|
|
|
| 27 |
A' |
215 |
214 |
0.03 |
|
|
|
| 28 |
A' |
199 |
198 |
0.01 |
|
|
|
| 29 |
A' |
80 |
79 |
0.00 |
|
|
|
| 30 |
A" |
3056 |
3039 |
15.12 |
|
|
|
| 31 |
A" |
3049 |
3031 |
29.59 |
|
|
|
| 32 |
A" |
3042 |
3024 |
5.87 |
|
|
|
| 33 |
A" |
2980 |
2962 |
0.01 |
|
|
|
| 34 |
A" |
2965 |
2948 |
42.61 |
|
|
|
| 35 |
A" |
2939 |
2921 |
0.56 |
|
|
|
| 36 |
A" |
1428 |
1419 |
3.54 |
|
|
|
| 37 |
A" |
1422 |
1413 |
2.58 |
|
|
|
| 38 |
A" |
1420 |
1412 |
0.12 |
|
|
|
| 39 |
A" |
1398 |
1390 |
0.14 |
|
|
|
| 40 |
A" |
1342 |
1334 |
2.06 |
|
|
|
| 41 |
A" |
1326 |
1318 |
0.30 |
|
|
|
| 42 |
A" |
1255 |
1248 |
1.14 |
|
|
|
| 43 |
A" |
1220 |
1213 |
4.00 |
|
|
|
| 44 |
A" |
1132 |
1126 |
5.36 |
|
|
|
| 45 |
A" |
1020 |
1014 |
3.76 |
|
|
|
| 46 |
A" |
1015 |
1009 |
0.01 |
|
|
|
| 47 |
A" |
953 |
948 |
6.13 |
|
|
|
| 48 |
A" |
866 |
861 |
0.13 |
|
|
|
| 49 |
A" |
745 |
740 |
0.25 |
|
|
|
| 50 |
A" |
438 |
435 |
0.26 |
|
|
|
| 51 |
A" |
299 |
297 |
0.04 |
|
|
|
| 52 |
A" |
263 |
261 |
0.00 |
|
|
|
| 53 |
A" |
213 |
212 |
0.14 |
|
|
|
| 54 |
A" |
82 |
82 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 40042.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 39810.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.136 |
|
|
|
| 2 |
C |
-0.007 |
|
|
|
| 3 |
H |
-0.003 |
|
|
|
| 4 |
C |
-0.034 |
|
|
|
| 5 |
C |
-0.034 |
|
|
|
| 6 |
C |
-0.045 |
|
|
|
| 7 |
C |
-0.045 |
|
|
|
| 8 |
H |
0.025 |
|
|
|
| 9 |
H |
0.025 |
|
|
|
| 10 |
H |
0.025 |
|
|
|
| 11 |
H |
0.013 |
|
|
|
| 12 |
H |
0.013 |
|
|
|
| 13 |
H |
0.011 |
|
|
|
| 14 |
H |
0.011 |
|
|
|
| 15 |
H |
0.027 |
|
|
|
| 16 |
H |
0.027 |
|
|
|
| 17 |
H |
0.029 |
|
|
|
| 18 |
H |
0.034 |
|
|
|
| 19 |
H |
0.029 |
|
|
|
| 20 |
H |
0.034 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.060 |
0.018 |
0.000 |
0.062 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.744 |
0.117 |
0.000 |
| y |
0.117 |
-41.971 |
0.000 |
| z |
0.000 |
0.000 |
-42.253 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.368 |
0.117 |
0.000 |
| y |
0.117 |
0.027 |
0.000 |
| z |
0.000 |
0.000 |
-0.396 |
|
| Polar |
|---|
| 3z2-r2 | -0.791 |
|---|
| x2-y2 | 0.228 |
|---|
| xy | 0.117 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.872 |
-0.375 |
0.000 |
| y |
-0.375 |
9.863 |
0.000 |
| z |
0.000 |
0.000 |
12.308 |
<r2> (average value of r
2) Å
2
| <r2> |
241.014 |
| (<r2>)1/2 |
15.525 |