return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (2-methylbutyraldehyde)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-270.999255
Energy at 298.15K-271.010063
HF Energy-270.038715
Nuclear repulsion energy238.541015
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3163 3034 18.77      
2 A 3152 3024 23.85      
3 A 3146 3019 33.28      
4 A 3137 3009 33.42      
5 A 3085 2960 23.90      
6 A 3070 2945 14.59      
7 A 3057 2933 19.99      
8 A 3042 2918 34.87      
9 A 3019 2896 19.41      
10 A 2918 2800 120.95      
11 A 1822 1748 136.81      
12 A 1507 1446 2.21      
13 A 1501 1440 7.33      
14 A 1495 1434 4.55      
15 A 1492 1432 7.16      
16 A 1483 1423 0.90      
17 A 1439 1380 5.86      
18 A 1424 1366 2.45      
19 A 1414 1357 4.10      
20 A 1389 1332 2.41      
21 A 1376 1320 2.20      
22 A 1315 1261 0.89      
23 A 1284 1231 0.48      
24 A 1204 1155 1.67      
25 A 1185 1136 3.88      
26 A 1130 1084 1.76      
27 A 1064 1021 0.50      
28 A 1028 986 9.03      
29 A 978 938 3.58      
30 A 945 907 6.22      
31 A 924 887 9.16      
32 A 804 772 6.41      
33 A 785 753 1.74      
34 A 659 632 8.25      
35 A 408 391 0.74      
36 A 392 376 0.47      
37 A 301 289 4.59      
38 A 262 251 0.33      
39 A 239 230 2.64      
40 A 208 199 0.44      
41 A 96 92 2.60      
42 A 78 75 2.87      

Unscaled Zero Point Vibrational Energy (zpe) 31209.0 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 29941.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.23350 0.06915 0.05781

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.382 -0.705 0.208
C2 -0.083 0.070 0.403
C3 1.063 -0.690 -0.302
C4 2.460 -0.135 0.009
C5 -0.235 1.526 -0.049
O6 -2.407 -0.242 -0.251
H7 -1.329 -1.782 0.523
H8 0.120 0.036 1.496
H9 1.024 -1.756 0.000
H10 0.886 -0.668 -1.396
H11 3.241 -0.754 -0.467
H12 2.580 0.898 -0.360
H13 2.647 -0.130 1.100
H14 -0.399 1.576 -1.141
H15 -1.106 1.997 0.438
H16 0.663 2.118 0.197

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.52532.49763.88902.52201.21461.12372.11312.63352.77824.67264.31244.16632.82632.72523.4860
C21.52531.54522.58141.53202.43422.23581.11262.17292.17273.53372.89202.82472.17962.18122.1891
C32.49761.54521.53482.58073.49932.75572.15611.10851.10892.18552.19772.18762.82443.53102.8796
C43.88902.58141.53483.16614.87504.16312.77772.16552.17621.10471.10431.10643.52494.17632.8875
C52.52201.53202.58073.16612.80813.53142.17603.51542.80834.17822.90133.51691.10541.10311.1034
O61.21462.43423.49934.87502.80812.03313.08453.75853.51275.67575.11765.23252.85112.67893.8983
H71.12372.23582.75574.16313.53142.03312.52062.41003.13514.78784.82194.34393.86143.78644.3917
H82.11311.11262.15612.77772.17603.08452.52062.50293.07383.77103.20092.56333.09782.54302.5134
H92.63352.17291.10852.16553.51543.75852.41002.50291.77552.47783.09832.54653.79854.33663.8953
H102.77822.17271.10892.17622.80833.51273.13513.07381.77552.53332.52953.10132.59883.79893.2171
H114.67263.53372.18551.10474.17825.67574.78783.77102.47782.53331.78281.78774.37465.22323.9158
H124.31242.89202.19771.10432.90135.11764.82193.20093.09832.52951.78281.78733.15403.92852.3394
H134.16632.82472.18761.10643.51695.23254.34392.56332.54653.10131.78771.78734.14874.36423.1310
H142.82632.17962.82443.52491.10542.85113.86143.09783.79852.59884.37463.15404.14871.78001.7926
H152.72522.18123.53104.17631.10312.67893.78642.54304.33663.79895.22323.92854.36421.78001.7898
H163.48602.18912.87962.88751.10343.89834.39172.51343.89533.21713.91582.33943.13101.79261.7898

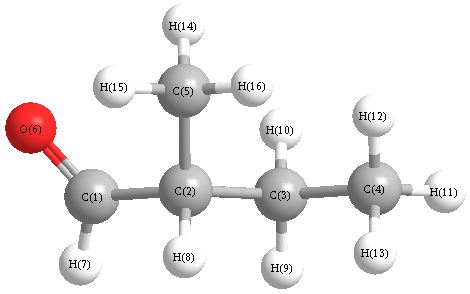
picture of 2-methylbutyraldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 108.864 C1 C2 C5 111.160
C1 C2 H8 105.389 C2 C1 O6 124.971
C2 C1 H7 114.281 C2 C3 C4 113.879
C2 C3 H9 108.817 C2 C3 H10 108.781
C2 C5 H14 110.418 C2 C5 H15 110.681
C2 C5 H16 111.289 C3 C2 C5 113.990
C3 C2 H8 107.306 C3 C4 H11 110.729
C3 C4 H12 111.727 C3 C4 H13 110.796
C4 C3 H9 108.946 C4 C3 H10 109.753
C5 C2 H8 109.710 O6 C1 H7 120.747
H9 C3 H10 106.393 H11 C4 H12 107.624
H11 C4 H13 107.905 H12 C4 H13 107.905
H14 C5 H15 107.412 H14 C5 H16 108.506
H15 C5 H16 108.415
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability