return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (2-methylbutyraldehyde)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-270.990216
Energy at 298.15K-271.001047
HF Energy-270.039357
Nuclear repulsion energy238.771132
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3174 3037 17.84      
2 A 3164 3027 22.53      
3 A 3158 3022 31.18      
4 A 3149 3013 31.59      
5 A 3098 2964 22.63      
6 A 3079 2946 13.85      
7 A 3067 2934 19.24      
8 A 3053 2921 33.29      
9 A 3030 2898 18.85      
10 A 2938 2810 110.53      
11 A 1868 1787 127.55      
12 A 1510 1444 2.29      
13 A 1504 1439 7.71      
14 A 1498 1433 4.96      
15 A 1495 1430 7.35      
16 A 1486 1422 0.96      
17 A 1446 1384 6.32      
18 A 1428 1366 3.69      
19 A 1421 1359 3.79      
20 A 1393 1333 2.02      
21 A 1381 1322 2.23      
22 A 1319 1261 0.79      
23 A 1288 1232 0.49      
24 A 1207 1154 1.72      
25 A 1189 1137 4.00      
26 A 1135 1085 1.66      
27 A 1068 1022 0.36      
28 A 1031 987 9.45      
29 A 981 938 3.83      
30 A 949 908 5.82      
31 A 929 889 8.94      
32 A 807 772 6.35      
33 A 787 752 1.76      
34 A 664 635 8.91      
35 A 409 391 0.73      
36 A 394 377 0.48      
37 A 303 290 4.56      
38 A 262 251 0.37      
39 A 240 230 2.64      
40 A 209 200 0.42      
41 A 98 93 2.83      
42 A 79 75 2.67      

Unscaled Zero Point Vibrational Energy (zpe) 31341.8 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29984.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.23364 0.06931 0.05792

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.383 -0.704 0.208
C2 -0.085 0.070 0.402
C3 1.061 -0.690 -0.301
C4 2.458 -0.135 0.010
C5 -0.234 1.526 -0.049
O6 -2.402 -0.242 -0.251
H7 -1.334 -1.780 0.523
H8 0.119 0.037 1.495
H9 1.022 -1.755 0.001
H10 0.885 -0.669 -1.395
H11 3.239 -0.755 -0.465
H12 2.580 0.897 -0.362
H13 2.644 -0.129 1.100
H14 -0.400 1.576 -1.140
H15 -1.103 1.998 0.439
H16 0.665 2.116 0.196

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.52392.49693.88782.52151.20931.12252.11272.63322.77754.67104.31154.16492.82552.72603.4849
C21.52391.54502.58101.53192.42782.23551.11212.17222.17223.53272.89252.82382.17942.18082.1887
C32.49691.54501.53452.57973.49292.75792.15491.10791.10842.18472.19722.18722.82453.52992.8772
C43.88782.58101.53453.16424.86834.16512.77582.16462.17561.10421.10381.10583.52444.17342.8840
C52.52151.53192.57973.16422.80453.53082.17453.51422.80774.17632.90033.51371.10491.10271.1029
O61.20932.42783.49294.86832.80452.02633.07963.75253.50695.66865.11155.22542.84712.67933.8941
H71.12252.23552.75794.16513.53082.02632.52142.41373.13684.78954.82354.34593.86073.78584.3909
H82.11271.11212.15492.77582.17453.07962.52142.50193.07243.76833.20002.56113.09632.54102.5117
H92.63322.17221.10792.16463.51423.75252.41372.50191.77452.47613.09702.54643.79824.33553.8925
H102.77752.17221.10842.17562.80773.50693.13683.07241.77452.53302.52853.10052.59963.79863.2148
H114.67103.53272.18471.10424.17635.66864.78953.76832.47612.53301.78201.78684.37465.22033.9122
H124.31152.89252.19721.10382.90035.11154.82353.20003.09702.52851.78201.78653.15393.92642.3372
H134.16492.82382.18721.10583.51375.22544.34592.56112.54643.10051.78681.78654.14674.35953.1264
H142.82552.17942.82453.52441.10492.84713.86073.09633.79822.59964.37463.15394.14671.77951.7918
H152.72602.18083.52994.17341.10272.67933.78582.54104.33553.79865.22033.92644.35951.77951.7887
H163.48492.18872.87722.88401.10293.89414.39092.51173.89253.21483.91222.33723.12641.79181.7887

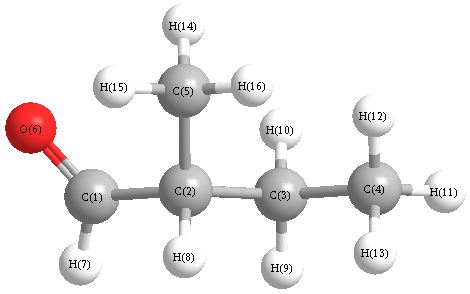
picture of 2-methylbutyraldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 108.904 C1 C2 C5 111.206
C1 C2 H8 105.476 C2 C1 O6 124.920
C2 C1 H7 114.439 C2 C3 C4 113.885
C2 C3 H9 108.814 C2 C3 H10 108.789
C2 C5 H14 110.437 C2 C5 H15 110.681
C2 C5 H16 111.291 C3 C2 C5 113.942
C3 C2 H8 107.263 C3 C4 H11 110.717
C3 C4 H12 111.736 C3 C4 H13 110.817
C4 C3 H9 108.935 C4 C3 H10 109.760
C5 C2 H8 109.635 O6 C1 H7 120.640
H9 C3 H10 106.385 H11 C4 H12 107.616
H11 C4 H13 107.896 H12 C4 H13 107.902
H14 C5 H15 107.431 H14 C5 H16 108.498
H15 C5 H16 108.383
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability