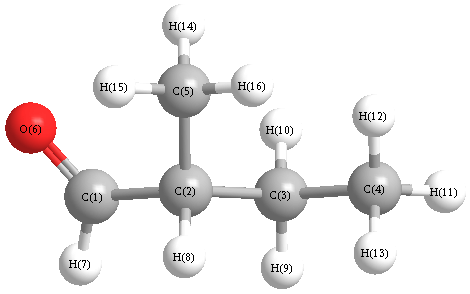
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3078 |
3060 |
13.36 |
|
|
|
| 2 |
A |
3063 |
3046 |
17.69 |
|
|
|
| 3 |
A |
3056 |
3039 |
24.64 |
|
|
|
| 4 |
A |
3048 |
3031 |
30.30 |
|
|
|
| 5 |
A |
2990 |
2973 |
19.92 |
|
|
|
| 6 |
A |
2984 |
2967 |
14.82 |
|
|
|
| 7 |
A |
2970 |
2952 |
24.36 |
|
|
|
| 8 |
A |
2949 |
2932 |
28.24 |
|
|
|
| 9 |
A |
2898 |
2881 |
17.64 |
|
|
|
| 10 |
A |
2742 |
2726 |
149.46 |
|
|
|
| 11 |
A |
1767 |
1756 |
164.47 |
|
|
|
| 12 |
A |
1430 |
1422 |
3.65 |
|
|
|
| 13 |
A |
1422 |
1414 |
9.09 |
|
|
|
| 14 |
A |
1419 |
1410 |
5.12 |
|
|
|
| 15 |
A |
1412 |
1404 |
7.90 |
|
|
|
| 16 |
A |
1404 |
1395 |
2.05 |
|
|
|
| 17 |
A |
1362 |
1354 |
5.96 |
|
|
|
| 18 |
A |
1349 |
1341 |
1.70 |
|
|
|
| 19 |
A |
1336 |
1328 |
4.99 |
|
|
|
| 20 |
A |
1310 |
1302 |
6.31 |
|
|
|
| 21 |
A |
1301 |
1294 |
1.01 |
|
|
|
| 22 |
A |
1249 |
1242 |
2.47 |
|
|
|
| 23 |
A |
1211 |
1204 |
0.54 |
|
|
|
| 24 |
A |
1147 |
1140 |
3.03 |
|
|
|
| 25 |
A |
1130 |
1124 |
2.64 |
|
|
|
| 26 |
A |
1079 |
1073 |
0.73 |
|
|
|
| 27 |
A |
1022 |
1016 |
0.48 |
|
|
|
| 28 |
A |
984 |
978 |
13.39 |
|
|
|
| 29 |
A |
939 |
933 |
6.75 |
|
|
|
| 30 |
A |
906 |
901 |
13.48 |
|
|
|
| 31 |
A |
881 |
876 |
13.14 |
|
|
|
| 32 |
A |
774 |
770 |
6.63 |
|
|
|
| 33 |
A |
758 |
754 |
3.22 |
|
|
|
| 34 |
A |
632 |
628 |
4.91 |
|
|
|
| 35 |
A |
395 |
393 |
1.16 |
|
|
|
| 36 |
A |
380 |
378 |
0.49 |
|
|
|
| 37 |
A |
277 |
275 |
4.77 |
|
|
|
| 38 |
A |
261 |
259 |
0.67 |
|
|
|
| 39 |
A |
222 |
220 |
1.54 |
|
|
|
| 40 |
A |
181 |
180 |
0.51 |
|
|
|
| 41 |
A |
92 |
92 |
1.31 |
|
|
|
| 42 |
A |
75 |
75 |
2.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29942.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29768.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.160 |
|
|
|
| 2 |
C |
-0.179 |
|
|
|
| 3 |
C |
-0.051 |
|
|
|
| 4 |
C |
-0.046 |
|
|
|
| 5 |
C |
0.004 |
|
|
|
| 6 |
O |
-0.164 |
|
|
|
| 7 |
H |
-0.017 |
|
|
|
| 8 |
H |
0.032 |
|
|
|
| 9 |
H |
0.021 |
|
|
|
| 10 |
H |
0.030 |
|
|
|
| 11 |
H |
0.034 |
|
|
|
| 12 |
H |
0.040 |
|
|
|
| 13 |
H |
0.033 |
|
|
|
| 14 |
H |
0.038 |
|
|
|
| 15 |
H |
0.039 |
|
|
|
| 16 |
H |
0.028 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.368 |
-0.319 |
0.619 |
2.469 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.141 |
-0.188 |
-1.370 |
| y |
-0.188 |
-37.540 |
-0.176 |
| z |
-1.370 |
-0.176 |
-37.524 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.610 |
-0.188 |
-1.370 |
| y |
-0.188 |
3.793 |
-0.176 |
| z |
-1.370 |
-0.176 |
3.817 |
|
| Polar |
|---|
| 3z2-r2 | 7.634 |
|---|
| x2-y2 | -7.602 |
|---|
| xy | -0.188 |
|---|
| xz | -1.370 |
|---|
| yz | -0.176 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.472 |
-0.240 |
0.222 |
| y |
-0.240 |
9.082 |
-0.242 |
| z |
0.222 |
-0.242 |
7.420 |
<r2> (average value of r
2) Å
2
| <r2> |
206.790 |
| (<r2>)1/2 |
14.380 |