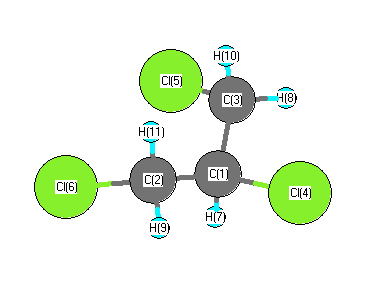
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3096 |
3078 |
2.56 |
|
|
|
| 2 |
A' |
3044 |
3026 |
2.59 |
|
|
|
| 3 |
A' |
3019 |
3002 |
11.68 |
|
|
|
| 4 |
A' |
1412 |
1404 |
4.78 |
|
|
|
| 5 |
A' |
1258 |
1251 |
14.96 |
|
|
|
| 6 |
A' |
1213 |
1206 |
1.83 |
|
|
|
| 7 |
A' |
1133 |
1127 |
16.70 |
|
|
|
| 8 |
A' |
999 |
993 |
1.37 |
|
|
|
| 9 |
A' |
851 |
846 |
13.14 |
|
|
|
| 10 |
A' |
771 |
767 |
5.05 |
|
|
|
| 11 |
A' |
688 |
684 |
110.39 |
|
|
|
| 12 |
A' |
380 |
378 |
0.05 |
|
|
|
| 13 |
A' |
326 |
324 |
0.61 |
|
|
|
| 14 |
A' |
160 |
159 |
6.47 |
|
|
|
| 15 |
A' |
108 |
107 |
1.94 |
|
|
|
| 16 |
A" |
3093 |
3075 |
0.53 |
|
|
|
| 17 |
A" |
3020 |
3002 |
0.32 |
|
|
|
| 18 |
A" |
1392 |
1384 |
6.42 |
|
|
|
| 19 |
A" |
1327 |
1320 |
0.00 |
|
|
|
| 20 |
A" |
1215 |
1208 |
2.37 |
|
|
|
| 21 |
A" |
1105 |
1099 |
0.00 |
|
|
|
| 22 |
A" |
1089 |
1082 |
1.16 |
|
|
|
| 23 |
A" |
829 |
824 |
4.67 |
|
|
|
| 24 |
A" |
720 |
716 |
16.77 |
|
|
|
| 25 |
A" |
242 |
241 |
12.70 |
|
|
|
| 26 |
A" |
240 |
239 |
0.18 |
|
|
|
| 27 |
A" |
31 |
31 |
0.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16380.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 16285.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.139 |
|
|
|
| 2 |
C |
-0.020 |
|
|
|
| 3 |
C |
-0.020 |
|
|
|
| 4 |
Cl |
-0.086 |
|
|
|
| 5 |
Cl |
-0.096 |
|
|
|
| 6 |
Cl |
-0.096 |
|
|
|
| 7 |
H |
0.088 |
|
|
|
| 8 |
H |
0.097 |
|
|
|
| 9 |
H |
0.097 |
|
|
|
| 10 |
H |
0.086 |
|
|
|
| 11 |
H |
0.086 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.183 |
1.511 |
0.000 |
1.522 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-55.429 |
-1.936 |
0.000 |
| y |
-1.936 |
-61.244 |
0.000 |
| z |
0.000 |
0.000 |
-57.063 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.724 |
-1.936 |
0.000 |
| y |
-1.936 |
-4.998 |
0.000 |
| z |
0.000 |
0.000 |
1.273 |
|
| Polar |
|---|
| 3z2-r2 | 2.547 |
|---|
| x2-y2 | 5.815 |
|---|
| xy | -1.936 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.791 |
1.068 |
0.000 |
| y |
1.068 |
13.098 |
0.000 |
| z |
0.000 |
0.000 |
8.665 |
<r2> (average value of r
2) Å
2
| <r2> |
367.174 |
| (<r2>)1/2 |
19.162 |