return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (3-Pentanone)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-271.000752
Energy at 298.15K 
HF Energy-270.053117
Nuclear repulsion energy237.995133
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3167 3030 28.60      
2 A1 3082 2948 0.99      
3 A1 3066 2933 31.58      
4 A1 1863 1782 95.63      
5 A1 1500 1435 5.70      
6 A1 1470 1407 9.16      
7 A1 1428 1366 8.41      
8 A1 1372 1312 2.49      
9 A1 1134 1085 0.88      
10 A1 1039 994 2.01      
11 A1 802 767 1.12      
12 A1 413 395 0.43      
13 A1 197 189 0.41      
14 A2 3174 3036 0.00      
15 A2 3098 2964 0.00      
16 A2 1495 1430 0.00      
17 A2 1271 1216 0.00      
18 A2 1007 964 0.00      
19 A2 721 690 0.00      
20 A2 233 223 0.00      
21 A2 30 28 0.00      
22 B1 3174 3037 42.93      
23 B1 3107 2973 22.17      
24 B1 1495 1431 10.78      
25 B1 1312 1255 1.00      
26 B1 1150 1100 0.17      
27 B1 824 788 6.78      
28 B1 475 454 0.01      
29 B1 212 203 0.22      
30 B1 69 66 0.18      
31 B2 3167 3030 15.81      
32 B2 3081 2948 38.92      
33 B2 3058 2925 11.31      
34 B2 1502 1437 12.72      
35 B2 1458 1395 0.92      
36 B2 1435 1373 14.74      
37 B2 1415 1354 15.72      
38 B2 1162 1112 54.95      
39 B2 1034 989 13.84      
40 B2 991 948 11.13      
41 B2 636 608 7.43      
42 B2 316 302 12.12      

Unscaled Zero Point Vibrational Energy (zpe) 31316.6 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29960.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.29554 0.06475 0.05531

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 1.280
C2 0.000 0.000 0.066
C3 0.000 1.298 -0.739
C4 0.000 -1.298 -0.739
C5 0.000 2.551 0.139
C6 0.000 -2.551 0.139
H7 0.881 1.281 -1.411
H8 -0.881 1.281 -1.411
H9 -0.881 -1.281 -1.411
H10 0.881 -1.281 -1.411
H11 0.000 3.463 -0.481
H12 -0.889 2.570 0.793
H13 0.889 2.570 0.793
H14 0.000 -3.463 -0.481
H15 0.889 -2.570 0.793
H16 -0.889 -2.570 0.793

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
O11.21422.40072.40072.79422.79423.10763.10763.10763.10763.88552.76232.76233.88552.76232.7623
C21.21421.52781.52782.55192.55192.14422.14422.14422.14423.50642.81442.81443.50642.81442.8144
C32.40071.52782.59701.52993.94841.10801.10802.80752.80752.18032.18002.18004.76904.25434.2543
C42.40071.52782.59703.94841.52992.80752.80751.10801.10804.76904.25434.25432.18032.18002.1800
C52.79422.55191.52993.94845.10172.18902.18904.22654.22651.10361.10301.10306.04625.23795.2379
C62.79422.55193.94841.52995.10174.22654.22652.18902.18906.04625.23795.23791.10361.10301.1030
H73.10762.14421.10802.80752.18904.22651.76263.11002.56232.53043.10602.55234.91454.43654.7766
H83.10762.14421.10802.80752.18904.22651.76262.56233.11002.53042.55233.10604.91454.77664.4365
H93.10762.14422.80751.10804.22652.18903.11002.56231.76264.91454.43654.77662.53043.10602.5523
H103.10762.14422.80751.10804.22652.18902.56233.11001.76264.91454.77664.43652.53042.55233.1060
H113.88553.50642.18034.76901.10366.04622.53042.53044.91454.91451.79191.79196.92696.22986.2298
H122.76232.81442.18004.25431.10305.23793.10602.55234.43654.77661.79191.77746.22985.43805.1393
H132.76232.81442.18004.25431.10305.23792.55233.10604.77664.43651.79191.77746.22985.13935.4380
H143.88553.50644.76902.18036.04621.10364.91454.91452.53042.53046.92696.22986.22981.79191.7919
H152.76232.81444.25432.18005.23791.10304.43654.77663.10602.55236.22985.43805.13931.79191.7774
H162.76232.81444.25432.18005.23791.10304.77664.43652.55233.10606.22985.13935.43801.79191.7774

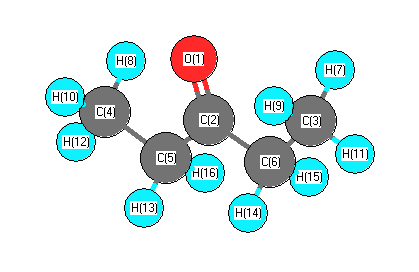
picture of 3-Pentanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 121.800 O1 C2 C4 121.800
C2 C3 C5 113.142 C2 C3 H7 107.816
C2 C3 H8 107.816 C2 C4 C6 113.142
C2 C4 H9 107.816 C2 C4 H10 107.816
C3 C2 C4 116.401 C3 C5 H11 110.725
C3 C5 H12 110.736 C3 C5 H13 110.736
C4 C6 H14 110.725 C4 C6 H15 110.736
C4 C6 H16 110.736 C5 C3 H7 111.152
C5 C3 H8 111.152 C6 C4 H9 111.152
C6 C4 H10 111.152 H7 C3 H8 105.388
H9 C4 H10 105.388 H11 C5 H12 108.594
H11 C5 H13 108.594 H12 C5 H13 107.352
H14 C6 H15 108.594 H14 C6 H16 108.594
H15 C6 H16 107.352
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability