return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(CH3)CONH(CH3) (Urea, N,N'-dimethyl-)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-303.053109
Energy at 298.15K-303.063391
HF Energy-302.066939
Nuclear repulsion energy247.380357
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3620 3473 14.99      
2 A 3185 3056 13.50      
3 A 3136 3009 9.89      
4 A 3049 2926 5.70      
5 A 1830 1756 229.34      
6 A 1528 1466 13.37      
7 A 1496 1435 0.23      
8 A 1482 1422 2.01      
9 A 1453 1394 2.03      
10 A 1218 1169 0.47      
11 A 1188 1139 0.66      
12 A 1144 1097 8.25      
13 A 935 897 0.00      
14 A 600 576 143.13      
15 A 464 445 20.83      
16 A 226 217 2.70      
17 A 206 197 0.50      
18 A 117 113 0.65      
19 B 3618 3471 9.97      
20 B 3185 3056 8.39      
21 B 3136 3009 56.74      
22 B 3048 2925 107.22      
23 B 1583 1519 342.02      
24 B 1509 1448 0.27      
25 B 1488 1428 55.17      
26 B 1456 1397 6.95      
27 B 1289 1236 213.46      
28 B 1162 1115 15.13      
29 B 1150 1103 16.27      
30 B 1069 1026 1.47      
31 B 785 753 49.69      
32 B 748 717 14.38      
33 B 519 498 72.96      
34 B 324 311 38.84      
35 B 130 124 3.30      
36 B 124 119 2.87      

Unscaled Zero Point Vibrational Energy (zpe) 26599.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 25519.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.32199 0.07250 0.06088

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.096
O2 0.000 0.000 1.317
N3 0.000 1.163 -0.677
N4 0.000 -1.163 -0.677
C5 -0.294 2.412 0.021
C6 0.294 -2.412 0.021
H7 -0.495 1.052 -1.559
H8 0.495 -1.052 -1.559
H9 -0.132 3.253 -0.674
H10 0.132 -3.253 -0.674
H11 0.397 2.515 0.872
H12 -0.397 -2.515 0.872
H13 -1.328 2.459 0.416
H14 1.328 -2.459 0.416

Atom - Atom Distances (Å)
  C1 O2 N3 N4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.22151.39621.39622.43132.43132.02252.02253.34553.34552.66202.66202.81282.8128
O21.22152.30852.30852.75422.75423.10253.10253.81643.81642.58502.58502.93642.9364
N31.39622.30852.32571.46123.65461.01742.43512.09444.41792.09424.01052.15344.0093
N41.39622.30852.32573.65461.46122.43511.01744.41792.09444.01052.09424.00932.1534
C52.43132.75421.46123.65464.86042.09473.88881.10295.72381.10065.00151.10795.1492
C62.43132.75423.65461.46124.86043.88882.09475.72381.10295.00151.10065.14921.1079
H72.02253.10251.01742.43512.09473.88882.32562.39984.43982.97414.31802.56364.4216
H82.02253.10252.43511.01743.88882.09472.32564.43982.39984.31802.97414.42162.5636
H93.34553.81642.09444.41791.10295.72382.39984.43986.51141.79275.97771.80255.9953
H103.34553.81644.41792.09445.72381.10294.43982.39986.51145.97771.79275.99531.8025
H112.66202.58502.09424.01051.10065.00152.97414.31801.79275.97775.09261.78525.0809
H122.66202.58504.01052.09425.00151.10064.31802.97415.97771.79275.09265.08091.7852
H132.81282.93642.15344.00931.10795.14922.56364.42161.80255.99531.78525.08095.5891
H142.81282.93644.00932.15345.14921.10794.42162.56365.99531.80255.08091.78525.5891

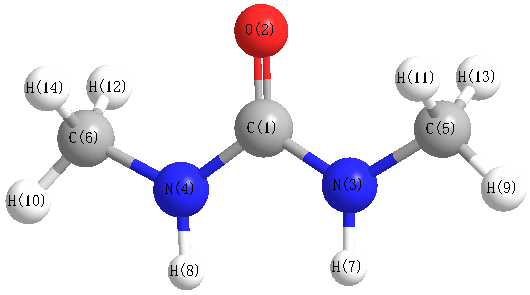
picture of Urea, N,N'-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C5 116.598 C1 N3 H7 112.912
C1 N4 C6 116.598 C1 N4 H8 112.912
O2 C1 N3 123.606 O2 C1 N4 123.606
N3 C1 N4 112.789 N3 C5 H9 108.729
N3 C5 H11 108.851 N3 C5 H13 113.178
N4 C6 H10 108.729 N4 C6 H12 108.851
N4 C6 H14 113.178 C5 N3 H7 114.176
C6 N4 H8 114.176 H9 C5 H11 108.897
H9 C5 H13 109.238 H10 C6 H12 108.897
H10 C6 H14 109.238 H11 C5 H13 107.870
H12 C6 H14 107.870
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability