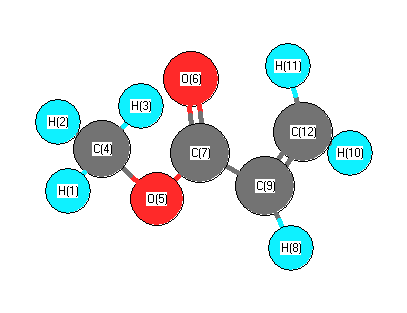
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3192 |
3173 |
1.83 |
|
|
|
| 2 |
A |
3126 |
3108 |
4.29 |
|
|
|
| 3 |
A |
3104 |
3086 |
13.00 |
|
|
|
| 4 |
A |
3083 |
3065 |
3.47 |
|
|
|
| 5 |
A |
3056 |
3039 |
17.93 |
|
|
|
| 6 |
A |
2975 |
2958 |
30.87 |
|
|
|
| 7 |
A |
1746 |
1736 |
148.16 |
|
|
|
| 8 |
A |
1654 |
1644 |
19.95 |
|
|
|
| 9 |
A |
1418 |
1410 |
5.25 |
|
|
|
| 10 |
A |
1395 |
1387 |
7.92 |
|
|
|
| 11 |
A |
1394 |
1386 |
33.94 |
|
|
|
| 12 |
A |
1360 |
1352 |
57.52 |
|
|
|
| 13 |
A |
1274 |
1267 |
67.20 |
|
|
|
| 14 |
A |
1168 |
1162 |
295.05 |
|
|
|
| 15 |
A |
1146 |
1139 |
13.74 |
|
|
|
| 16 |
A |
1115 |
1109 |
0.24 |
|
|
|
| 17 |
A |
1033 |
1027 |
3.40 |
|
|
|
| 18 |
A |
1005 |
999 |
15.87 |
|
|
|
| 19 |
A |
991 |
986 |
20.77 |
|
|
|
| 20 |
A |
949 |
944 |
13.46 |
|
|
|
| 21 |
A |
841 |
836 |
8.37 |
|
|
|
| 22 |
A |
796 |
792 |
19.45 |
|
|
|
| 23 |
A |
643 |
640 |
4.44 |
|
|
|
| 24 |
A |
515 |
512 |
0.00 |
|
|
|
| 25 |
A |
461 |
458 |
1.30 |
|
|
|
| 26 |
A |
329 |
327 |
15.39 |
|
|
|
| 27 |
A |
213 |
211 |
2.84 |
|
|
|
| 28 |
A |
204 |
203 |
3.17 |
|
|
|
| 29 |
A |
109 |
109 |
0.21 |
|
|
|
| 30 |
A |
103 |
102 |
2.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20199.4 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20082.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.045 |
|
|
|
| 2 |
H |
0.051 |
|
|
|
| 3 |
H |
0.051 |
|
|
|
| 4 |
C |
0.067 |
|
|
|
| 5 |
O |
-0.233 |
|
|
|
| 6 |
O |
-0.225 |
|
|
|
| 7 |
C |
0.191 |
|
|
|
| 8 |
H |
0.006 |
|
|
|
| 9 |
C |
-0.094 |
|
|
|
| 10 |
H |
0.040 |
|
|
|
| 11 |
H |
0.041 |
|
|
|
| 12 |
C |
0.059 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.198 |
-1.401 |
-0.000 |
1.415 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.703 |
0.690 |
-0.000 |
| y |
0.690 |
-37.973 |
-0.000 |
| z |
-0.000 |
-0.000 |
-35.918 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.242 |
0.690 |
-0.000 |
| y |
0.690 |
-6.663 |
-0.000 |
| z |
-0.000 |
-0.000 |
-3.580 |
|
| Polar |
|---|
| 3z2-r2 | -7.159 |
|---|
| x2-y2 | 11.270 |
|---|
| xy | 0.690 |
|---|
| xz | -0.000 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.718 |
-0.451 |
-0.000 |
| y |
-0.451 |
7.175 |
0.000 |
| z |
-0.000 |
0.000 |
4.195 |
<r2> (average value of r
2) Å
2
| <r2> |
178.878 |
| (<r2>)1/2 |
13.375 |