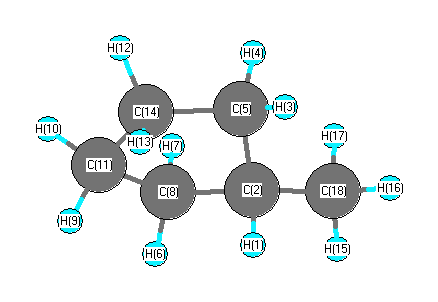
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3043 |
3025 |
25.45 |
|
|
|
| 2 |
A |
3042 |
3025 |
83.09 |
|
|
|
| 3 |
A |
3035 |
3018 |
20.57 |
|
|
|
| 4 |
A |
3024 |
3007 |
11.19 |
|
|
|
| 5 |
A |
3020 |
3003 |
5.99 |
|
|
|
| 6 |
A |
3013 |
2996 |
43.03 |
|
|
|
| 7 |
A |
2992 |
2974 |
46.49 |
|
|
|
| 8 |
A |
2979 |
2961 |
19.36 |
|
|
|
| 9 |
A |
2959 |
2942 |
70.77 |
|
|
|
| 10 |
A |
2954 |
2937 |
17.78 |
|
|
|
| 11 |
A |
2950 |
2933 |
20.51 |
|
|
|
| 12 |
A |
2925 |
2908 |
10.21 |
|
|
|
| 13 |
A |
1440 |
1432 |
1.41 |
|
|
|
| 14 |
A |
1421 |
1413 |
9.67 |
|
|
|
| 15 |
A |
1417 |
1409 |
1.99 |
|
|
|
| 16 |
A |
1414 |
1406 |
1.30 |
|
|
|
| 17 |
A |
1413 |
1405 |
2.54 |
|
|
|
| 18 |
A |
1402 |
1394 |
0.04 |
|
|
|
| 19 |
A |
1348 |
1340 |
2.43 |
|
|
|
| 20 |
A |
1331 |
1323 |
0.55 |
|
|
|
| 21 |
A |
1291 |
1284 |
2.49 |
|
|
|
| 22 |
A |
1280 |
1273 |
0.24 |
|
|
|
| 23 |
A |
1272 |
1264 |
0.32 |
|
|
|
| 24 |
A |
1263 |
1256 |
1.94 |
|
|
|
| 25 |
A |
1239 |
1232 |
0.02 |
|
|
|
| 26 |
A |
1209 |
1202 |
0.09 |
|
|
|
| 27 |
A |
1197 |
1190 |
0.86 |
|
|
|
| 28 |
A |
1167 |
1160 |
0.23 |
|
|
|
| 29 |
A |
1153 |
1147 |
0.16 |
|
|
|
| 30 |
A |
1125 |
1118 |
0.79 |
|
|
|
| 31 |
A |
1083 |
1077 |
0.05 |
|
|
|
| 32 |
A |
1031 |
1025 |
0.07 |
|
|
|
| 33 |
A |
987 |
982 |
0.99 |
|
|
|
| 34 |
A |
974 |
968 |
2.49 |
|
|
|
| 35 |
A |
943 |
937 |
0.51 |
|
|
|
| 36 |
A |
902 |
897 |
0.02 |
|
|
|
| 37 |
A |
895 |
890 |
0.03 |
|
|
|
| 38 |
A |
879 |
874 |
0.80 |
|
|
|
| 39 |
A |
838 |
834 |
1.24 |
|
|
|
| 40 |
A |
791 |
787 |
0.18 |
|
|
|
| 41 |
A |
730 |
725 |
0.93 |
|
|
|
| 42 |
A |
609 |
605 |
0.38 |
|
|
|
| 43 |
A |
518 |
515 |
0.80 |
|
|
|
| 44 |
A |
418 |
416 |
0.31 |
|
|
|
| 45 |
A |
295 |
293 |
0.05 |
|
|
|
| 46 |
A |
229 |
228 |
0.00 |
|
|
|
| 47 |
A |
179 |
178 |
0.02 |
|
|
|
| 48 |
A |
24 |
24 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35822.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 35614.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.000 |
|
|
|
| 2 |
C |
-0.154 |
|
|
|
| 3 |
H |
0.008 |
|
|
|
| 4 |
H |
0.014 |
|
|
|
| 5 |
C |
0.010 |
|
|
|
| 6 |
H |
0.008 |
|
|
|
| 7 |
H |
0.014 |
|
|
|
| 8 |
C |
0.010 |
|
|
|
| 9 |
H |
0.020 |
|
|
|
| 10 |
H |
0.014 |
|
|
|
| 11 |
C |
-0.028 |
|
|
|
| 12 |
H |
0.014 |
|
|
|
| 13 |
H |
0.020 |
|
|
|
| 14 |
C |
-0.028 |
|
|
|
| 15 |
H |
0.025 |
|
|
|
| 16 |
H |
0.025 |
|
|
|
| 17 |
H |
0.025 |
|
|
|
| 18 |
C |
0.005 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.010 |
-0.000 |
-0.087 |
0.087 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.828 |
-0.001 |
-0.213 |
| y |
-0.001 |
-40.492 |
-0.001 |
| z |
-0.213 |
-0.001 |
-39.521 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.822 |
-0.001 |
-0.213 |
| y |
-0.001 |
-0.317 |
-0.001 |
| z |
-0.213 |
-0.001 |
1.139 |
|
| Polar |
|---|
| 3z2-r2 | 2.278 |
|---|
| x2-y2 | -0.336 |
|---|
| xy | -0.001 |
|---|
| xz | -0.213 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.041 |
0.000 |
0.023 |
| y |
0.000 |
9.946 |
0.000 |
| z |
0.023 |
0.000 |
8.971 |
<r2> (average value of r
2) Å
2
| <r2> |
180.856 |
| (<r2>)1/2 |
13.448 |