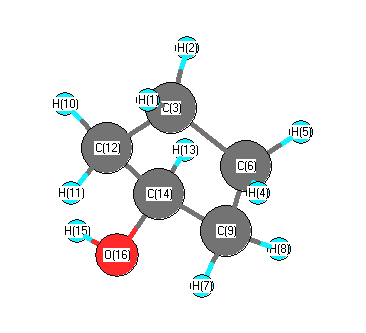
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3661 |
3639 |
12.16 |
|
|
|
| 2 |
A |
3050 |
3033 |
51.86 |
|
|
|
| 3 |
A |
3036 |
3018 |
17.51 |
|
|
|
| 4 |
A |
3024 |
3006 |
14.85 |
|
|
|
| 5 |
A |
3021 |
3003 |
35.23 |
|
|
|
| 6 |
A |
2996 |
2979 |
34.84 |
|
|
|
| 7 |
A |
2984 |
2967 |
25.74 |
|
|
|
| 8 |
A |
2970 |
2953 |
27.26 |
|
|
|
| 9 |
A |
2963 |
2946 |
27.69 |
|
|
|
| 10 |
A |
2868 |
2851 |
60.30 |
|
|
|
| 11 |
A |
1441 |
1433 |
3.11 |
|
|
|
| 12 |
A |
1415 |
1407 |
5.25 |
|
|
|
| 13 |
A |
1413 |
1404 |
1.55 |
|
|
|
| 14 |
A |
1404 |
1396 |
1.25 |
|
|
|
| 15 |
A |
1374 |
1366 |
3.45 |
|
|
|
| 16 |
A |
1321 |
1314 |
33.17 |
|
|
|
| 17 |
A |
1285 |
1278 |
1.45 |
|
|
|
| 18 |
A |
1269 |
1262 |
3.08 |
|
|
|
| 19 |
A |
1262 |
1254 |
5.60 |
|
|
|
| 20 |
A |
1239 |
1232 |
17.20 |
|
|
|
| 21 |
A |
1227 |
1220 |
0.87 |
|
|
|
| 22 |
A |
1212 |
1205 |
13.30 |
|
|
|
| 23 |
A |
1165 |
1158 |
4.90 |
|
|
|
| 24 |
A |
1150 |
1143 |
9.61 |
|
|
|
| 25 |
A |
1143 |
1136 |
2.38 |
|
|
|
| 26 |
A |
1081 |
1074 |
51.46 |
|
|
|
| 27 |
A |
1052 |
1046 |
29.45 |
|
|
|
| 28 |
A |
1025 |
1019 |
11.38 |
|
|
|
| 29 |
A |
963 |
957 |
0.58 |
|
|
|
| 30 |
A |
942 |
936 |
10.58 |
|
|
|
| 31 |
A |
923 |
918 |
2.08 |
|
|
|
| 32 |
A |
885 |
879 |
0.09 |
|
|
|
| 33 |
A |
864 |
859 |
0.05 |
|
|
|
| 34 |
A |
797 |
793 |
2.35 |
|
|
|
| 35 |
A |
750 |
745 |
0.71 |
|
|
|
| 36 |
A |
590 |
586 |
0.87 |
|
|
|
| 37 |
A |
526 |
523 |
4.07 |
|
|
|
| 38 |
A |
458 |
456 |
6.35 |
|
|
|
| 39 |
A |
354 |
352 |
16.42 |
|
|
|
| 40 |
A |
297 |
295 |
90.11 |
|
|
|
| 41 |
A |
198 |
197 |
0.59 |
|
|
|
| 42 |
A |
43 |
42 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30819.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 30640.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.013 |
|
|
|
| 2 |
H |
0.022 |
|
|
|
| 3 |
C |
-0.041 |
|
|
|
| 4 |
H |
0.024 |
|
|
|
| 5 |
H |
0.015 |
|
|
|
| 6 |
C |
-0.033 |
|
|
|
| 7 |
H |
0.027 |
|
|
|
| 8 |
H |
0.017 |
|
|
|
| 9 |
C |
0.012 |
|
|
|
| 10 |
H |
0.004 |
|
|
|
| 11 |
H |
0.020 |
|
|
|
| 12 |
C |
-0.007 |
|
|
|
| 13 |
H |
-0.006 |
|
|
|
| 14 |
C |
0.028 |
|
|
|
| 15 |
H |
0.135 |
|
|
|
| 16 |
O |
-0.231 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.321 |
-1.105 |
0.683 |
1.338 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.254 |
3.495 |
-1.617 |
| y |
3.495 |
-37.817 |
-0.156 |
| z |
-1.617 |
-0.156 |
-37.965 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.363 |
3.495 |
-1.617 |
| y |
3.495 |
0.293 |
-0.156 |
| z |
-1.617 |
-0.156 |
0.070 |
|
| Polar |
|---|
| 3z2-r2 | 0.140 |
|---|
| x2-y2 | -0.438 |
|---|
| xy | 3.495 |
|---|
| xz | -1.617 |
|---|
| yz | -0.156 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.400 |
0.236 |
-0.009 |
| y |
0.236 |
8.916 |
-0.075 |
| z |
-0.009 |
-0.075 |
7.891 |
<r2> (average value of r
2) Å
2
| <r2> |
167.203 |
| (<r2>)1/2 |
12.931 |