return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O2 (γ–Butyrolactone)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-305.703709
Energy at 298.15K 
HF Energy-304.722211
Nuclear repulsion energy239.329714
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3156 3008        
2 A 3141 2994        
3 A 3133 2986        
4 A 3072 2928        
5 A 3067 2923        
6 A 3045 2902        
7 A 1875 1787        
8 A 1514 1443        
9 A 1483 1414        
10 A 1453 1385        
11 A 1394 1329        
12 A 1342 1279        
13 A 1303 1242        
14 A 1254 1196        
15 A 1215 1158        
16 A 1190 1134        
17 A 1176 1121        
18 A 1089 1038        
19 A 1078 1027        
20 A 1005 958        
21 A 946 901        
22 A 895 853        
23 A 876 835        
24 A 807 769        
25 A 677 645        
26 A 629 599        
27 A 523 499        
28 A 486 463        
29 A 229 218        
30 A 152 145        

Unscaled Zero Point Vibrational Energy (zpe) 21602.1 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 20589.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.23959 0.11841 0.08483

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.889 -0.002 0.006
C2 -0.034 1.210 0.193
C3 -1.398 0.666 -0.256
C4 -1.268 -0.814 0.148
O5 0.124 -1.144 -0.051
O6 2.091 -0.025 -0.081
H7 0.347 2.074 -0.375
H8 -0.034 1.474 1.269
H9 -1.500 0.749 -1.354
H10 -2.265 1.160 0.217
H11 -1.870 -1.504 -0.467
H12 -1.524 -0.966 1.216

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12
C11.53482.39692.30901.37601.20612.17872.15012.84963.36743.17652.8663
C21.53481.53522.37092.37242.47361.10181.10822.18032.23123.34222.8290
C32.39691.53521.53972.37433.56172.24562.19971.10651.10372.23092.2014
C42.30902.37091.53971.44453.45863.35002.83112.18002.21251.10261.1089
O51.37602.37242.37431.44452.26393.24222.93632.81433.33002.06832.0864
O61.20612.47363.56173.45862.26392.74442.92983.88824.52444.24613.9549
H72.17871.10182.24563.35003.24222.74441.79112.47542.83014.21023.9084
H82.15011.10822.19972.83112.93632.92981.79113.09152.48673.90532.8597
H92.84962.18031.10652.18002.81433.88822.47543.09151.79532.44953.0899
H103.36742.23121.10372.21253.33004.52442.83012.48671.79532.77852.4632
H113.17653.34222.23091.10262.06834.24614.21023.90532.44952.77851.7998
H122.86632.82902.20141.10892.08643.95493.90842.85973.08992.46321.7998

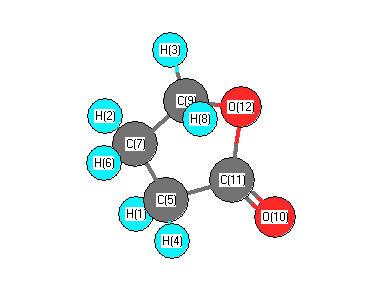
picture of γ–Butyrolactone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 102.655 C1 C2 H7 110.364
C1 C2 H8 107.787 C1 O5 C4 109.875
C2 C1 O5 109.053 C2 C1 O6 128.572
C2 C3 C4 100.897 C2 C3 H9 110.193
C2 C3 H10 114.462 C3 C2 H7 115.795
C3 C2 H8 111.620 C3 C4 O5 105.382
C3 C4 H11 114.180 C3 C4 H12 111.395
C4 C3 H9 109.853 C4 C3 H10 112.598
O5 C1 O6 122.374 O5 C4 H11 107.832
O5 C4 H12 108.885 H7 C2 H8 108.281
H9 C3 H10 108.635 H11 C4 H12 108.949
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability