return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O2 (γ–Butyrolactone)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-305.673661
Energy at 298.15K 
HF Energy-304.723483
Nuclear repulsion energy239.873183
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3173 3044 8.81      
2 A 3158 3030 28.52      
3 A 3150 3022 15.84      
4 A 3089 2964 17.91      
5 A 3085 2960 8.74      
6 A 3066 2942 46.42      
7 A 1903 1825 358.99      
8 A 1527 1465 0.43      
9 A 1496 1435 6.40      
10 A 1465 1405 7.72      
11 A 1412 1355 25.58      
12 A 1357 1302 5.68      
13 A 1321 1268 22.34      
14 A 1268 1217 11.47      
15 A 1230 1180 33.26      
16 A 1204 1155 22.38      
17 A 1199 1150 148.41      
18 A 1106 1061 49.46      
19 A 1096 1052 33.53      
20 A 1017 976 13.93      
21 A 956 917 3.74      
22 A 905 868 12.81      
23 A 888 852 14.74      
24 A 819 786 5.21      
25 A 686 658 5.03      
26 A 638 612 3.20      
27 A 532 511 3.26      
28 A 493 473 4.01      
29 A 229 220 2.54      
30 A 153 146 0.28      

Unscaled Zero Point Vibrational Energy (zpe) 21810.4 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 20924.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.24072 0.11895 0.08516

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.887 -0.002 0.006
C2 -0.033 1.208 0.189
C3 -1.398 0.666 -0.250
C4 -1.265 -0.815 0.144
O5 0.125 -1.140 -0.050
O6 2.086 -0.025 -0.080
H7 0.347 2.068 -0.383
H8 -0.025 1.478 1.262
H9 -1.510 0.755 -1.345
H10 -2.259 1.156 0.232
H11 -1.862 -1.501 -0.477
H12 -1.525 -0.976 1.208

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12
C11.53132.39412.30411.37001.20242.17482.14522.85393.36053.16742.8655
C21.53131.53312.36922.36522.46631.10031.10652.17792.22783.33582.8350
C32.39411.53311.53782.37023.55582.24202.19751.10501.10212.22732.1993
C42.30412.36921.53781.44023.44993.34432.83592.17752.20931.10101.1071
O51.37002.36522.37021.44022.25593.23242.93162.81783.32172.06362.0809
O61.20242.46633.55583.44992.25592.73842.91863.89134.51394.23303.9500
H72.17481.10032.24203.34433.23242.73841.78742.46932.82874.19763.9118
H82.14521.10652.19752.83592.93162.91861.78743.08662.48133.90742.8765
H92.85392.17791.10502.17752.81783.89132.46933.08661.79172.44233.0847
H103.36052.22781.10212.20933.32174.51392.82872.48131.79172.77832.4573
H113.16743.33582.22731.10102.06364.23304.19763.90742.44232.77831.7964
H122.86552.83502.19931.10712.08093.95003.91182.87653.08472.45731.7964

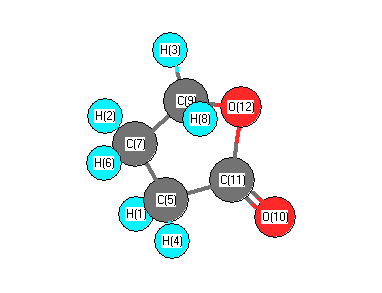
picture of γ–Butyrolactone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 102.750 C1 C2 H7 110.391
C1 C2 H8 107.741 C1 O5 C4 110.127
C2 C1 O5 109.089 C2 C1 O6 128.490
C2 C3 C4 100.976 C2 C3 H9 110.235
C2 C3 H10 114.441 C3 C2 H7 115.750
C3 C2 H8 111.692 C3 C4 O5 105.435
C3 C4 H11 114.120 C3 C4 H12 111.475
C4 C3 H9 109.880 C4 C3 H10 112.583
O5 C1 O6 122.421 O5 C4 H11 107.853
O5 C4 H12 108.856 H7 C2 H8 108.187
H9 C3 H10 108.539 H11 C4 H12 108.885
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability