return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O2 (γ–Butyrolactone)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-305.662214
Energy at 298.15K 
HF Energy-304.724383
Nuclear repulsion energy240.322063
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3183 3045 8.31      
2 A 3168 3031 24.89      
3 A 3160 3024 16.91      
4 A 3100 2966 17.39      
5 A 3095 2961 7.49      
6 A 3074 2941 45.31      
7 A 1947 1862 376.92      
8 A 1532 1466 0.66      
9 A 1500 1435 7.45      
10 A 1467 1404 8.66      
11 A 1422 1361 32.17      
12 A 1362 1303 7.55      
13 A 1328 1270 27.24      
14 A 1274 1219 14.92      
15 A 1237 1183 60.60      
16 A 1213 1161 149.01      
17 A 1209 1156 2.83      
18 A 1122 1074 75.64      
19 A 1104 1056 6.64      
20 A 1023 978 11.05      
21 A 961 919 3.36      
22 A 910 871 13.90      
23 A 893 855 11.98      
24 A 826 791 4.91      
25 A 694 664 5.33      
26 A 643 615 3.30      
27 A 536 513 3.20      
28 A 500 478 4.31      
29 A 228 219 2.60      
30 A 152 146 0.27      

Unscaled Zero Point Vibrational Energy (zpe) 21931.5 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 20981.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.24129 0.11950 0.08549

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.886 -0.003 0.006
C2 -0.032 1.208 0.188
C3 -1.397 0.665 -0.248
C4 -1.260 -0.816 0.143
O5 0.126 -1.136 -0.049
O6 2.080 -0.025 -0.080
H7 0.348 2.066 -0.385
H8 -0.023 1.479 1.260
H9 -1.513 0.755 -1.343
H10 -2.258 1.153 0.237
H11 -1.856 -1.501 -0.479
H12 -1.523 -0.979 1.206

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 O6 H7 H8 H9 H10 H11 H12
C11.52982.39212.29851.36521.19742.17362.14382.85423.35713.16162.8627
C21.52981.53302.36772.36112.45941.09991.10612.17772.22723.33322.8361
C32.39211.53301.53772.36723.54882.24212.19691.10451.10152.22582.1984
C42.29852.36771.53771.43563.43933.34212.83622.17752.20941.10071.1068
O51.36522.36112.36721.43562.24773.22752.92842.81743.31742.06082.0784
O61.19742.45943.54883.43932.24772.73262.91183.88734.50564.22243.9425
H72.17361.09992.24213.34213.22752.73261.78642.46892.82964.19393.9123
H82.14381.10612.19692.83622.92842.91181.78643.08542.47953.90692.8802
H92.85422.17771.10452.17752.81743.88732.46893.08541.79072.44043.0830
H103.35712.22721.10152.20943.31744.50562.82962.47951.79072.77802.4551
H113.16163.33322.22581.10072.06084.22244.19393.90692.44042.77801.7947
H122.86272.83612.19841.10682.07843.94253.91232.88023.08302.45511.7947

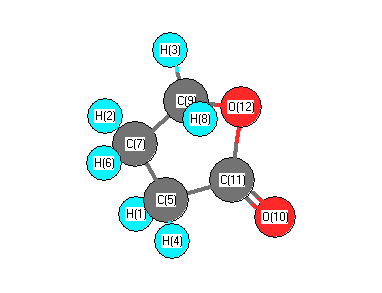
picture of γ–Butyrolactone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 102.705 C1 C2 H7 110.425
C1 C2 H8 107.754 C1 O5 C4 110.279
C2 C1 O5 109.157 C2 C1 O6 128.379
C2 C3 C4 100.897 C2 C3 H9 110.255
C2 C3 H10 114.437 C3 C2 H7 115.787
C3 C2 H8 111.683 C3 C4 O5 105.479
C3 C4 H11 114.024 C3 C4 H12 111.425
C4 C3 H9 109.909 C4 C3 H10 112.629
O5 C1 O6 122.464 O5 C4 H11 107.955
O5 C4 H12 108.985 H7 C2 H8 108.158
H9 C3 H10 108.527 H11 C4 H12 108.781
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability