Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -342.046087 |
| Energy at 298.15K | |
| HF Energy | -342.046087 |
| Nuclear repulsion energy | 243.359763 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
2972 |
2955 |
59.98 |
|
|
|
| 2 |
A1 |
1889 |
1878 |
526.78 |
|
|
|
| 3 |
A1 |
1467 |
1459 |
1.94 |
|
|
|
| 4 |
A1 |
1329 |
1322 |
1.27 |
|
|
|
| 5 |
A1 |
1153 |
1146 |
201.81 |
|
|
|
| 6 |
A1 |
932 |
927 |
9.07 |
|
|
|
| 7 |
A1 |
804 |
800 |
30.59 |
|
|
|
| 8 |
A1 |
700 |
696 |
0.67 |
|
|
|
| 9 |
A2 |
2999 |
2981 |
0.00 |
|
|
|
| 10 |
A2 |
1192 |
1185 |
0.00 |
|
|
|
| 11 |
A2 |
1114 |
1108 |
0.00 |
|
|
|
| 12 |
A2 |
164i |
163i |
0.00 |
|
|
|
| 13 |
B1 |
3021 |
3003 |
48.01 |
|
|
|
| 14 |
B1 |
1209 |
1201 |
0.82 |
|
|
|
| 15 |
B1 |
825 |
820 |
0.16 |
|
|
|
| 16 |
B1 |
717 |
713 |
12.91 |
|
|
|
| 17 |
B1 |
147 |
146 |
1.31 |
|
|
|
| 18 |
B2 |
2965 |
2947 |
31.83 |
|
|
|
| 19 |
B2 |
1455 |
1447 |
4.11 |
|
|
|
| 20 |
B2 |
1342 |
1335 |
19.61 |
|
|
|
| 21 |
B2 |
1140 |
1134 |
1.28 |
|
|
|
| 22 |
B2 |
1003 |
997 |
274.32 |
|
|
|
| 23 |
B2 |
727 |
723 |
3.86 |
|
|
|
| 24 |
B2 |
486 |
483 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15711.5 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15620.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.870 |
| O2 |
0.000 |
0.000 |
2.067 |
| O3 |
0.000 |
1.144 |
0.059 |
| O4 |
0.000 |
-1.144 |
0.059 |
| C5 |
0.000 |
0.777 |
-1.295 |
| C6 |
0.000 |
-0.777 |
-1.295 |
| H7 |
-0.900 |
1.198 |
-1.791 |
| H8 |
0.900 |
1.198 |
-1.791 |
| H9 |
0.900 |
-1.198 |
-1.791 |
| H10 |
-0.900 |
-1.198 |
-1.791 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1972 | 1.4024 | 1.4024 | 2.3003 | 2.3003 | 3.0539 | 3.0539 | 3.0539 | 3.0539 |
O2 | 1.1972 | | 2.3114 | 2.3114 | 3.4508 | 3.4508 | 4.1389 | 4.1389 | 4.1389 | 4.1389 | O3 | 1.4024 | 2.3114 | | 2.2879 | 1.4024 | 2.3504 | 2.0577 | 2.0577 | 3.1172 | 3.1172 | O4 | 1.4024 | 2.3114 | 2.2879 | | 2.3504 | 1.4024 | 3.1172 | 3.1172 | 2.0577 | 2.0577 | C5 | 2.3003 | 3.4508 | 1.4024 | 2.3504 | | 1.5549 | 1.1102 | 1.1102 | 2.2269 | 2.2269 | C6 | 2.3003 | 3.4508 | 2.3504 | 1.4024 | 1.5549 | | 2.2269 | 2.2269 | 1.1102 | 1.1102 | H7 | 3.0539 | 4.1389 | 2.0577 | 3.1172 | 1.1102 | 2.2269 | | 1.7995 | 2.9968 | 2.3964 | H8 | 3.0539 | 4.1389 | 2.0577 | 3.1172 | 1.1102 | 2.2269 | 1.7995 | | 2.3964 | 2.9968 | H9 | 3.0539 | 4.1389 | 3.1172 | 2.0577 | 2.2269 | 1.1102 | 2.9968 | 2.3964 | | 1.7995 | H10 | 3.0539 | 4.1389 | 3.1172 | 2.0577 | 2.2269 | 1.1102 | 2.3964 | 2.9968 | 1.7995 | |
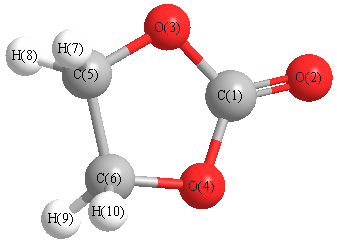 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
110.192 |
|
C1 |
O4 |
C6 |
110.192 |
| O2 |
C1 |
O3 |
125.341 |
|
O2 |
C1 |
O4 |
125.341 |
| O3 |
C1 |
O4 |
109.317 |
|
O3 |
C5 |
C6 |
105.150 |
| O3 |
C5 |
H7 |
109.406 |
|
O3 |
C5 |
H8 |
109.406 |
| O4 |
C6 |
C5 |
105.150 |
|
O4 |
C6 |
H9 |
109.406 |
| O4 |
C6 |
H10 |
109.406 |
|
C5 |
C6 |
H9 |
112.269 |
| C5 |
C6 |
H10 |
112.269 |
|
C6 |
C5 |
H7 |
112.269 |
| C6 |
C5 |
H8 |
112.269 |
|
H7 |
C5 |
H8 |
108.269 |
| H9 |
C6 |
H10 |
108.269 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.270 |
|
|
|
| 2 |
O |
-0.178 |
|
|
|
| 3 |
O |
-0.219 |
|
|
|
| 4 |
O |
-0.219 |
|
|
|
| 5 |
C |
0.078 |
|
|
|
| 6 |
C |
0.078 |
|
|
|
| 7 |
H |
0.048 |
|
|
|
| 8 |
H |
0.048 |
|
|
|
| 9 |
H |
0.048 |
|
|
|
| 10 |
H |
0.048 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.428 |
4.428 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.075 |
0.000 |
0.000 |
| y |
0.000 |
-35.720 |
0.000 |
| z |
0.000 |
0.000 |
-35.297 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.434 |
0.000 |
0.000 |
| y |
0.000 |
-2.034 |
0.000 |
| z |
0.000 |
0.000 |
-1.400 |
|
| Polar |
|---|
| 3z2-r2 | -2.800 |
|---|
| x2-y2 | 3.645 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.358 |
0.000 |
0.000 |
| y |
0.000 |
5.409 |
0.000 |
| z |
0.000 |
0.000 |
7.719 |
<r2> (average value of r
2) Å
2
| <r2> |
130.371 |
| (<r2>)1/2 |
11.418 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -342.046781 |
| Energy at 298.15K | -342.053046 |
| HF Energy | -342.046781 |
| Nuclear repulsion energy | 243.894004 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3034 |
3017 |
19.83 |
|
|
|
| 2 |
A |
2947 |
2929 |
31.53 |
|
|
|
| 3 |
A |
1891 |
1880 |
513.32 |
|
|
|
| 4 |
A |
1454 |
1446 |
2.23 |
|
|
|
| 5 |
A |
1337 |
1329 |
6.49 |
|
|
|
| 6 |
A |
1199 |
1193 |
14.66 |
|
|
|
| 7 |
A |
1151 |
1145 |
133.67 |
|
|
|
| 8 |
A |
1100 |
1093 |
46.14 |
|
|
|
| 9 |
A |
934 |
929 |
4.28 |
|
|
|
| 10 |
A |
799 |
794 |
33.61 |
|
|
|
| 11 |
A |
697 |
693 |
0.85 |
|
|
|
| 12 |
A |
136 |
135 |
0.36 |
|
|
|
| 13 |
B |
3043 |
3026 |
27.45 |
|
|
|
| 14 |
B |
2952 |
2935 |
57.67 |
|
|
|
| 15 |
B |
1447 |
1439 |
5.38 |
|
|
|
| 16 |
B |
1337 |
1329 |
16.84 |
|
|
|
| 17 |
B |
1211 |
1204 |
3.43 |
|
|
|
| 18 |
B |
1117 |
1111 |
2.91 |
|
|
|
| 19 |
B |
996 |
990 |
248.74 |
|
|
|
| 20 |
B |
863 |
858 |
11.41 |
|
|
|
| 21 |
B |
725 |
721 |
14.03 |
|
|
|
| 22 |
B |
664 |
660 |
1.40 |
|
|
|
| 23 |
B |
486 |
483 |
0.30 |
|
|
|
| 24 |
B |
155 |
154 |
1.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15837.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 15745.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.865 |
| O2 |
0.000 |
0.000 |
2.062 |
| O3 |
0.000 |
1.144 |
0.049 |
| O4 |
0.000 |
-1.144 |
0.049 |
| C5 |
0.194 |
0.745 |
-1.285 |
| C6 |
-0.194 |
-0.745 |
-1.285 |
| H7 |
-0.444 |
1.359 |
-1.951 |
| H8 |
1.260 |
0.897 |
-1.570 |
| H9 |
0.444 |
-1.359 |
-1.951 |
| H10 |
-1.260 |
-0.897 |
-1.570 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1969 | 1.4059 | 1.4059 | 2.2846 | 2.2846 | 3.1583 | 2.8852 | 3.1583 | 2.8852 |
O2 | 1.1969 | | 2.3161 | 2.3161 | 3.4352 | 3.4352 | 4.2602 | 3.9482 | 4.2602 | 3.9482 | O3 | 1.4059 | 2.3161 | | 2.2886 | 1.4059 | 2.3212 | 2.0592 | 2.0661 | 3.2345 | 2.8941 | O4 | 1.4059 | 2.3161 | 2.2886 | | 2.3212 | 1.4059 | 3.2345 | 2.8941 | 2.0592 | 2.0661 | C5 | 2.2846 | 3.4352 | 1.4059 | 2.3212 | | 1.5402 | 1.1073 | 1.1134 | 2.2212 | 2.2118 | C6 | 2.2846 | 3.4352 | 2.3212 | 1.4059 | 1.5402 | | 2.2212 | 2.2118 | 1.1073 | 1.1134 | H7 | 3.1583 | 4.2602 | 2.0592 | 3.2345 | 1.1073 | 2.2212 | | 1.8053 | 2.8596 | 2.4295 | H8 | 2.8852 | 3.9482 | 2.0661 | 2.8941 | 1.1134 | 2.2118 | 1.8053 | | 2.4295 | 3.0930 | H9 | 3.1583 | 4.2602 | 3.2345 | 2.0592 | 2.2212 | 1.1073 | 2.8596 | 2.4295 | | 1.8053 | H10 | 2.8852 | 3.9482 | 2.8941 | 2.0661 | 2.2118 | 1.1134 | 2.4295 | 3.0930 | 1.8053 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
108.677 |
|
C1 |
O4 |
C6 |
108.677 |
| O2 |
C1 |
O3 |
125.518 |
|
O2 |
C1 |
O4 |
125.518 |
| O3 |
C1 |
O4 |
108.964 |
|
O3 |
C5 |
C6 |
103.881 |
| O3 |
C5 |
H7 |
109.467 |
|
O3 |
C5 |
H8 |
109.642 |
| O4 |
C6 |
C5 |
103.881 |
|
O4 |
C6 |
H9 |
109.467 |
| O4 |
C6 |
H10 |
109.642 |
|
C5 |
C6 |
H9 |
113.045 |
| C5 |
C6 |
H10 |
111.917 |
|
C6 |
C5 |
H7 |
113.045 |
| C6 |
C5 |
H8 |
111.917 |
|
H7 |
C5 |
H8 |
108.763 |
| H9 |
C6 |
H10 |
108.763 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.268 |
|
|
|
| 2 |
O |
-0.175 |
|
|
|
| 3 |
O |
-0.221 |
|
|
|
| 4 |
O |
-0.221 |
|
|
|
| 5 |
C |
0.081 |
|
|
|
| 6 |
C |
0.081 |
|
|
|
| 7 |
H |
0.047 |
|
|
|
| 8 |
H |
0.046 |
|
|
|
| 9 |
H |
0.047 |
|
|
|
| 10 |
H |
0.046 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.381 |
4.381 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.164 |
0.133 |
0.000 |
| y |
0.133 |
-35.722 |
0.000 |
| z |
0.000 |
0.000 |
-35.119 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.256 |
0.133 |
0.000 |
| y |
0.133 |
-2.080 |
0.000 |
| z |
0.000 |
0.000 |
-1.177 |
|
| Polar |
|---|
| 3z2-r2 | -2.353 |
|---|
| x2-y2 | 3.557 |
|---|
| xy | 0.133 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.468 |
0.181 |
0.000 |
| y |
0.181 |
5.350 |
0.000 |
| z |
0.000 |
0.000 |
7.681 |
<r2> (average value of r
2) Å
2
| <r2> |
129.313 |
| (<r2>)1/2 |
11.372 |
