Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -342.422306 |
| Energy at 298.15K | -342.427133 |
| Nuclear repulsion energy | 269.842135 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3313 |
3179 |
0.24 |
|
|
|
| 2 |
A' |
3291 |
3157 |
0.01 |
|
|
|
| 3 |
A' |
3278 |
3145 |
1.75 |
|
|
|
| 4 |
A' |
2960 |
2840 |
112.34 |
|
|
|
| 5 |
A' |
1810 |
1736 |
251.06 |
|
|
|
| 6 |
A' |
1645 |
1578 |
6.02 |
|
|
|
| 7 |
A' |
1550 |
1487 |
39.61 |
|
|
|
| 8 |
A' |
1447 |
1389 |
27.04 |
|
|
|
| 9 |
A' |
1410 |
1353 |
4.97 |
|
|
|
| 10 |
A' |
1323 |
1270 |
31.63 |
|
|
|
| 11 |
A' |
1248 |
1198 |
5.92 |
|
|
|
| 12 |
A' |
1201 |
1152 |
17.48 |
|
|
|
| 13 |
A' |
1109 |
1064 |
5.15 |
|
|
|
| 14 |
A' |
1033 |
991 |
36.51 |
|
|
|
| 15 |
A' |
953 |
914 |
19.28 |
|
|
|
| 16 |
A' |
896 |
859 |
7.97 |
|
|
|
| 17 |
A' |
765 |
734 |
62.14 |
|
|
|
| 18 |
A' |
498 |
478 |
1.03 |
|
|
|
| 19 |
A' |
197 |
189 |
5.03 |
|
|
|
| 20 |
A" |
1016 |
975 |
0.20 |
|
|
|
| 21 |
A" |
877 |
842 |
1.16 |
|
|
|
| 22 |
A" |
846 |
812 |
4.76 |
|
|
|
| 23 |
A" |
781 |
749 |
55.40 |
|
|
|
| 24 |
A" |
648 |
621 |
2.12 |
|
|
|
| 25 |
A" |
599 |
574 |
6.75 |
|
|
|
| 26 |
A" |
284 |
273 |
11.97 |
|
|
|
| 27 |
A" |
131 |
126 |
1.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17554.3 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 16841.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.505 |
-0.905 |
0.000 |
| C2 |
-0.564 |
-1.745 |
0.000 |
| C3 |
-1.750 |
-1.051 |
0.000 |
| C4 |
0.000 |
0.367 |
0.000 |
| C5 |
-1.377 |
0.342 |
0.000 |
| C6 |
0.936 |
1.507 |
0.000 |
| O7 |
2.149 |
1.412 |
0.000 |
| H8 |
-0.331 |
-2.809 |
0.000 |
| H9 |
-2.753 |
-1.477 |
0.000 |
| H10 |
-2.036 |
1.211 |
0.000 |
| H11 |
0.413 |
2.497 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3597 | 2.2594 | 1.3684 | 2.2576 | 2.4501 | 2.8411 | 2.0791 | 3.3077 | 3.3073 | 3.4031 |
C2 | 1.3597 | | 1.3734 | 2.1854 | 2.2394 | 3.5811 | 4.1627 | 1.0893 | 2.2047 | 3.3021 | 4.3526 | C3 | 2.2594 | 1.3734 | | 2.2516 | 1.4420 | 3.7087 | 4.6112 | 2.2588 | 1.0899 | 2.2800 | 4.1546 | C4 | 1.3684 | 2.1854 | 2.2516 | | 1.3768 | 1.4753 | 2.3897 | 3.1925 | 3.3130 | 2.2045 | 2.1698 | C5 | 2.2576 | 2.2394 | 1.4420 | 1.3768 | | 2.5895 | 3.6841 | 3.3198 | 2.2809 | 1.0911 | 2.8008 | C6 | 2.4501 | 3.5811 | 3.7087 | 1.4753 | 2.5895 | | 1.2161 | 4.4977 | 4.7445 | 2.9872 | 1.1197 | O7 | 2.8411 | 4.1627 | 4.6112 | 2.3897 | 3.6841 | 1.2161 | | 4.8956 | 5.6896 | 4.1898 | 2.0466 | H8 | 2.0791 | 1.0893 | 2.2588 | 3.1925 | 3.3198 | 4.4977 | 4.8956 | | 2.7637 | 4.3666 | 5.3573 | H9 | 3.3077 | 2.2047 | 1.0899 | 3.3130 | 2.2809 | 4.7445 | 5.6896 | 2.7637 | | 2.7818 | 5.0804 | H10 | 3.3073 | 3.3021 | 2.2800 | 2.2045 | 1.0911 | 2.9872 | 4.1898 | 4.3666 | 2.7818 | | 2.7661 | H11 | 3.4031 | 4.3526 | 4.1546 | 2.1698 | 2.8008 | 1.1197 | 2.0466 | 5.3573 | 5.0804 | 2.7661 | |
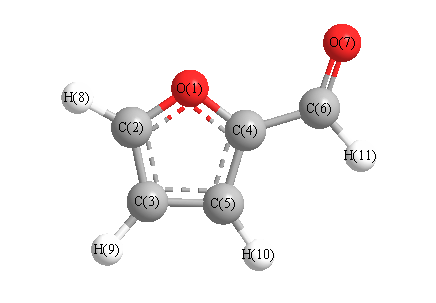 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.520 |
|
O1 |
C2 |
H8 |
115.765 |
| O1 |
C4 |
C5 |
110.650 |
|
O1 |
C4 |
C6 |
118.941 |
| C2 |
O1 |
C4 |
106.463 |
|
C2 |
C3 |
C5 |
105.361 |
| C2 |
C3 |
H9 |
126.639 |
|
C3 |
C2 |
H8 |
132.715 |
| C3 |
C5 |
C4 |
106.006 |
|
C3 |
C5 |
H10 |
127.806 |
| C4 |
C5 |
H10 |
126.188 |
|
C4 |
C6 |
O7 |
124.941 |
| C4 |
C6 |
H11 |
112.747 |
|
C5 |
C3 |
H9 |
128.000 |
| C5 |
C4 |
C6 |
130.409 |
|
O7 |
C6 |
H11 |
122.312 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -342.424107 |
| Energy at 298.15K | -342.428923 |
| Nuclear repulsion energy | 269.045742 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3313 |
3179 |
0.36 |
|
|
|
| 2 |
A' |
3300 |
3166 |
0.30 |
|
|
|
| 3 |
A' |
3283 |
3150 |
0.94 |
|
|
|
| 4 |
A' |
2983 |
2862 |
106.61 |
|
|
|
| 5 |
A' |
1800 |
1727 |
202.97 |
|
|
|
| 6 |
A' |
1656 |
1589 |
49.64 |
|
|
|
| 7 |
A' |
1545 |
1482 |
73.04 |
|
|
|
| 8 |
A' |
1458 |
1398 |
5.84 |
|
|
|
| 9 |
A' |
1407 |
1350 |
10.56 |
|
|
|
| 10 |
A' |
1272 |
1221 |
16.57 |
|
|
|
| 11 |
A' |
1261 |
1210 |
17.82 |
|
|
|
| 12 |
A' |
1187 |
1138 |
14.32 |
|
|
|
| 13 |
A' |
1113 |
1067 |
15.61 |
|
|
|
| 14 |
A' |
1026 |
984 |
35.40 |
|
|
|
| 15 |
A' |
968 |
929 |
10.69 |
|
|
|
| 16 |
A' |
895 |
858 |
5.41 |
|
|
|
| 17 |
A' |
758 |
728 |
70.45 |
|
|
|
| 18 |
A' |
494 |
474 |
0.88 |
|
|
|
| 19 |
A' |
201 |
193 |
6.37 |
|
|
|
| 20 |
A" |
1023 |
982 |
0.35 |
|
|
|
| 21 |
A" |
884 |
848 |
2.05 |
|
|
|
| 22 |
A" |
853 |
819 |
5.49 |
|
|
|
| 23 |
A" |
780 |
748 |
52.42 |
|
|
|
| 24 |
A" |
639 |
613 |
1.85 |
|
|
|
| 25 |
A" |
600 |
576 |
8.73 |
|
|
|
| 26 |
A" |
249 |
239 |
12.26 |
|
|
|
| 27 |
A" |
153 |
146 |
3.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17549.0 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 16836.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.270 |
-0.235 |
0.000 |
| C2 |
1.131 |
-1.589 |
0.000 |
| C3 |
-0.193 |
-1.954 |
0.000 |
| C4 |
0.000 |
0.286 |
0.000 |
| C5 |
-0.938 |
-0.720 |
0.000 |
| C6 |
-0.132 |
1.756 |
0.000 |
| O7 |
-1.209 |
2.326 |
0.000 |
| H8 |
2.058 |
-2.161 |
0.000 |
| H9 |
-0.584 |
-2.972 |
0.000 |
| H10 |
-2.018 |
-0.578 |
0.000 |
| H11 |
0.840 |
2.307 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3612 | 2.2577 | 1.3729 | 2.2605 | 2.4350 | 3.5641 | 2.0815 | 3.3058 | 3.3065 | 2.5779 |
C2 | 1.3612 | | 1.3739 | 2.1897 | 2.2441 | 3.5753 | 4.5609 | 1.0892 | 2.2032 | 3.3079 | 3.9067 | C3 | 2.2577 | 1.3739 | | 2.2482 | 1.4414 | 3.7101 | 4.3987 | 2.2605 | 1.0901 | 2.2856 | 4.3845 | C4 | 1.3729 | 2.1897 | 2.2482 | | 1.3749 | 1.4756 | 2.3712 | 3.1976 | 3.3095 | 2.1955 | 2.1888 | C5 | 2.2605 | 2.2441 | 1.4414 | 1.3749 | | 2.6030 | 3.0577 | 3.3244 | 2.2796 | 1.0899 | 3.5103 | C6 | 2.4350 | 3.5753 | 3.7101 | 1.4756 | 2.6030 | | 1.2181 | 4.4879 | 4.7488 | 3.0006 | 1.1181 | O7 | 3.5641 | 4.5609 | 4.3987 | 2.3712 | 3.0577 | 1.2181 | | 5.5504 | 5.3344 | 3.0149 | 2.0491 | H8 | 2.0815 | 1.0892 | 2.2605 | 3.1976 | 3.3244 | 4.4879 | 5.5504 | | 2.7631 | 4.3729 | 4.6314 | H9 | 3.3058 | 2.2032 | 1.0901 | 3.3095 | 2.2796 | 4.7488 | 5.3344 | 2.7631 | | 2.7904 | 5.4674 | H10 | 3.3065 | 3.3079 | 2.2856 | 2.1955 | 1.0899 | 3.0006 | 3.0149 | 4.3729 | 2.7904 | | 4.0616 | H11 | 2.5779 | 3.9067 | 4.3845 | 2.1888 | 3.5103 | 1.1181 | 2.0491 | 4.6314 | 5.4674 | 4.0616 | |
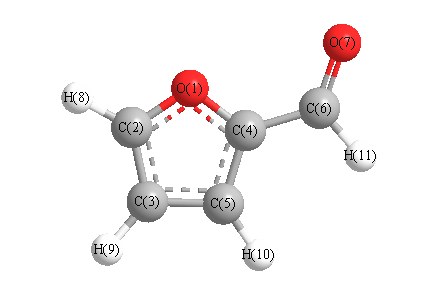 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.270 |
|
O1 |
C2 |
H8 |
115.861 |
| O1 |
C4 |
C5 |
110.703 |
|
O1 |
C4 |
C6 |
117.435 |
| C2 |
O1 |
C4 |
106.431 |
|
C2 |
C3 |
C5 |
105.686 |
| C2 |
C3 |
H9 |
126.414 |
|
C3 |
C2 |
H8 |
132.869 |
| C3 |
C5 |
C4 |
105.910 |
|
C3 |
C5 |
H10 |
128.558 |
| C4 |
C5 |
H10 |
125.532 |
|
C4 |
C6 |
O7 |
123.064 |
| C4 |
C6 |
H11 |
114.401 |
|
C5 |
C3 |
H9 |
127.900 |
| C5 |
C4 |
C6 |
131.862 |
|
O7 |
C6 |
H11 |
122.536 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
