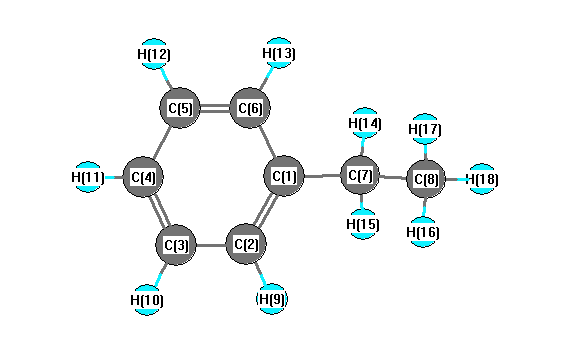
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3218 |
3080 |
15.92 |
|
|
|
| 2 |
A' |
3196 |
3058 |
4.64 |
|
|
|
| 3 |
A' |
3179 |
3043 |
9.93 |
|
|
|
| 4 |
A' |
3130 |
2995 |
39.95 |
|
|
|
| 5 |
A' |
3058 |
2927 |
28.95 |
|
|
|
| 6 |
A' |
3054 |
2923 |
29.45 |
|
|
|
| 7 |
A' |
1675 |
1603 |
6.20 |
|
|
|
| 8 |
A' |
1542 |
1475 |
12.99 |
|
|
|
| 9 |
A' |
1507 |
1442 |
3.25 |
|
|
|
| 10 |
A' |
1487 |
1423 |
1.11 |
|
|
|
| 11 |
A' |
1403 |
1343 |
2.59 |
|
|
|
| 12 |
A' |
1353 |
1295 |
1.42 |
|
|
|
| 13 |
A' |
1238 |
1185 |
0.26 |
|
|
|
| 14 |
A' |
1204 |
1152 |
0.21 |
|
|
|
| 15 |
A' |
1088 |
1041 |
7.18 |
|
|
|
| 16 |
A' |
1062 |
1017 |
2.86 |
|
|
|
| 17 |
A' |
1023 |
979 |
0.18 |
|
|
|
| 18 |
A' |
1014 |
970 |
0.09 |
|
|
|
| 19 |
A' |
987 |
945 |
2.63 |
|
|
|
| 20 |
A' |
935 |
895 |
1.65 |
|
|
|
| 21 |
A' |
790 |
756 |
9.15 |
|
|
|
| 22 |
A' |
774 |
741 |
18.91 |
|
|
|
| 23 |
A' |
722 |
691 |
38.47 |
|
|
|
| 24 |
A' |
563 |
539 |
7.60 |
|
|
|
| 25 |
A' |
499 |
477 |
4.59 |
|
|
|
| 26 |
A' |
307 |
293 |
0.18 |
|
|
|
| 27 |
A' |
134 |
128 |
0.69 |
|
|
|
| 28 |
A" |
3204 |
3067 |
33.28 |
|
|
|
| 29 |
A" |
3181 |
3044 |
6.07 |
|
|
|
| 30 |
A" |
3134 |
3000 |
30.82 |
|
|
|
| 31 |
A" |
3095 |
2962 |
6.49 |
|
|
|
| 32 |
A" |
1656 |
1585 |
0.74 |
|
|
|
| 33 |
A" |
1497 |
1433 |
7.50 |
|
|
|
| 34 |
A" |
1494 |
1430 |
7.42 |
|
|
|
| 35 |
A" |
1366 |
1308 |
0.23 |
|
|
|
| 36 |
A" |
1361 |
1303 |
0.36 |
|
|
|
| 37 |
A" |
1273 |
1218 |
0.01 |
|
|
|
| 38 |
A" |
1180 |
1129 |
0.02 |
|
|
|
| 39 |
A" |
1123 |
1075 |
3.95 |
|
|
|
| 40 |
A" |
1058 |
1013 |
2.30 |
|
|
|
| 41 |
A" |
993 |
951 |
0.00 |
|
|
|
| 42 |
A" |
866 |
829 |
0.02 |
|
|
|
| 43 |
A" |
786 |
752 |
0.69 |
|
|
|
| 44 |
A" |
627 |
600 |
0.04 |
|
|
|
| 45 |
A" |
413 |
395 |
0.00 |
|
|
|
| 46 |
A" |
348 |
333 |
0.08 |
|
|
|
| 47 |
A" |
217 |
208 |
0.01 |
|
|
|
| 48 |
A" |
19 |
18 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34514.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 33033.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.123 |
|
|
|
| 2 |
C |
-0.165 |
|
|
|
| 3 |
C |
-0.130 |
|
|
|
| 4 |
C |
-0.141 |
|
|
|
| 5 |
C |
-0.130 |
|
|
|
| 6 |
C |
-0.165 |
|
|
|
| 7 |
C |
-0.261 |
|
|
|
| 8 |
C |
-0.342 |
|
|
|
| 9 |
H |
0.121 |
|
|
|
| 10 |
H |
0.126 |
|
|
|
| 11 |
H |
0.128 |
|
|
|
| 12 |
H |
0.126 |
|
|
|
| 13 |
H |
0.121 |
|
|
|
| 14 |
H |
0.117 |
|
|
|
| 15 |
H |
0.117 |
|
|
|
| 16 |
H |
0.119 |
|
|
|
| 17 |
H |
0.119 |
|
|
|
| 18 |
H |
0.117 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.052 |
0.360 |
0.000 |
0.364 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-52.372 |
-0.097 |
0.000 |
| y |
-0.097 |
-44.879 |
0.000 |
| z |
0.000 |
0.000 |
-44.929 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.468 |
-0.097 |
0.000 |
| y |
-0.097 |
3.772 |
0.000 |
| z |
0.000 |
0.000 |
3.696 |
|
| Polar |
|---|
| 3z2-r2 | 7.392 |
|---|
| x2-y2 | -7.493 |
|---|
| xy | -0.097 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.488 |
0.812 |
0.000 |
| y |
0.812 |
16.869 |
0.000 |
| z |
0.000 |
0.000 |
13.808 |
<r2> (average value of r
2) Å
2
| <r2> |
287.545 |
| (<r2>)1/2 |
16.957 |