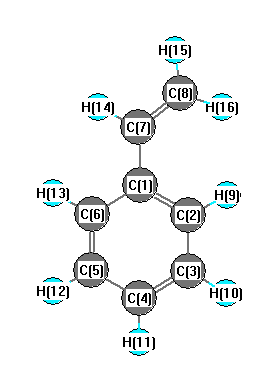
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3257 |
3117 |
11.74 |
|
|
|
| 2 |
A' |
3220 |
3082 |
13.11 |
|
|
|
| 3 |
A' |
3210 |
3072 |
29.25 |
|
|
|
| 4 |
A' |
3202 |
3064 |
9.27 |
|
|
|
| 5 |
A' |
3192 |
3055 |
0.65 |
|
|
|
| 6 |
A' |
3185 |
3048 |
7.52 |
|
|
|
| 7 |
A' |
3173 |
3036 |
1.79 |
|
|
|
| 8 |
A' |
3158 |
3023 |
13.65 |
|
|
|
| 9 |
A' |
1719 |
1645 |
12.00 |
|
|
|
| 10 |
A' |
1675 |
1603 |
1.82 |
|
|
|
| 11 |
A' |
1650 |
1579 |
2.62 |
|
|
|
| 12 |
A' |
1543 |
1477 |
11.21 |
|
|
|
| 13 |
A' |
1497 |
1432 |
5.32 |
|
|
|
| 14 |
A' |
1463 |
1400 |
6.25 |
|
|
|
| 15 |
A' |
1371 |
1312 |
0.52 |
|
|
|
| 16 |
A' |
1360 |
1301 |
2.57 |
|
|
|
| 17 |
A' |
1327 |
1270 |
1.90 |
|
|
|
| 18 |
A' |
1241 |
1187 |
1.88 |
|
|
|
| 19 |
A' |
1204 |
1152 |
0.26 |
|
|
|
| 20 |
A' |
1181 |
1131 |
0.19 |
|
|
|
| 21 |
A' |
1121 |
1073 |
4.43 |
|
|
|
| 22 |
A' |
1069 |
1023 |
0.68 |
|
|
|
| 23 |
A' |
1045 |
1000 |
5.50 |
|
|
|
| 24 |
A' |
1022 |
978 |
0.02 |
|
|
|
| 25 |
A' |
795 |
761 |
0.06 |
|
|
|
| 26 |
A' |
628 |
601 |
0.04 |
|
|
|
| 27 |
A' |
559 |
535 |
5.48 |
|
|
|
| 28 |
A' |
448 |
429 |
0.08 |
|
|
|
| 29 |
A' |
246 |
236 |
0.57 |
|
|
|
| 30 |
A" |
1039 |
994 |
13.70 |
|
|
|
| 31 |
A" |
1014 |
971 |
0.39 |
|
|
|
| 32 |
A" |
995 |
952 |
0.01 |
|
|
|
| 33 |
A" |
944 |
904 |
28.50 |
|
|
|
| 34 |
A" |
943 |
903 |
15.30 |
|
|
|
| 35 |
A" |
862 |
825 |
0.39 |
|
|
|
| 36 |
A" |
808 |
774 |
31.33 |
|
|
|
| 37 |
A" |
718 |
687 |
40.39 |
|
|
|
| 38 |
A" |
663 |
635 |
0.05 |
|
|
|
| 39 |
A" |
446 |
427 |
8.41 |
|
|
|
| 40 |
A" |
411 |
394 |
0.29 |
|
|
|
| 41 |
A" |
206 |
197 |
1.91 |
|
|
|
| 42 |
A" |
17 |
16 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29412.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 28150.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.122 |
|
|
|
| 2 |
C |
-0.120 |
|
|
|
| 3 |
C |
-0.140 |
|
|
|
| 4 |
C |
-0.133 |
|
|
|
| 5 |
C |
-0.139 |
|
|
|
| 6 |
C |
-0.148 |
|
|
|
| 7 |
C |
-0.098 |
|
|
|
| 8 |
C |
-0.337 |
|
|
|
| 9 |
H |
0.116 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
| 11 |
H |
0.130 |
|
|
|
| 12 |
H |
0.128 |
|
|
|
| 13 |
H |
0.114 |
|
|
|
| 14 |
H |
0.119 |
|
|
|
| 15 |
H |
0.135 |
|
|
|
| 16 |
H |
0.124 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.076 |
-0.174 |
0.000 |
0.190 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.122 |
0.207 |
0.000 |
| y |
0.207 |
-42.490 |
0.000 |
| z |
0.000 |
0.000 |
-52.306 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.276 |
0.207 |
0.000 |
| y |
0.207 |
4.723 |
0.000 |
| z |
0.000 |
0.000 |
-10.000 |
|
| Polar |
|---|
| 3z2-r2 | -19.999 |
|---|
| x2-y2 | 0.369 |
|---|
| xy | 0.207 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
15.283 |
-2.724 |
0.000 |
| y |
-2.724 |
18.544 |
0.000 |
| z |
0.000 |
0.000 |
6.547 |
<r2> (average value of r
2) Å
2
| <r2> |
273.567 |
| (<r2>)1/2 |
16.540 |