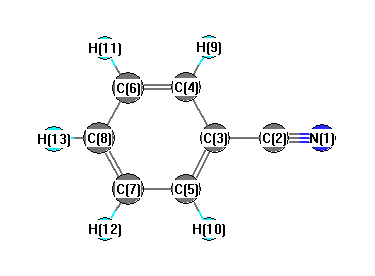
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3229 |
3098 |
2.37 |
|
|
|
| 2 |
A1 |
3217 |
3086 |
7.44 |
|
|
|
| 3 |
A1 |
3198 |
3068 |
0.04 |
|
|
|
| 4 |
A1 |
2367 |
2271 |
29.98 |
|
|
|
| 5 |
A1 |
1669 |
1601 |
0.84 |
|
|
|
| 6 |
A1 |
1538 |
1475 |
10.10 |
|
|
|
| 7 |
A1 |
1234 |
1184 |
0.67 |
|
|
|
| 8 |
A1 |
1206 |
1157 |
0.97 |
|
|
|
| 9 |
A1 |
1061 |
1018 |
3.43 |
|
|
|
| 10 |
A1 |
1026 |
984 |
0.33 |
|
|
|
| 11 |
A1 |
779 |
747 |
1.87 |
|
|
|
| 12 |
A1 |
467 |
448 |
0.00 |
|
|
|
| 13 |
A2 |
1007 |
965 |
0.00 |
|
|
|
| 14 |
A2 |
870 |
835 |
0.00 |
|
|
|
| 15 |
A2 |
410 |
394 |
0.00 |
|
|
|
| 16 |
B1 |
1030 |
988 |
0.06 |
|
|
|
| 17 |
B1 |
960 |
921 |
3.25 |
|
|
|
| 18 |
B1 |
788 |
756 |
37.29 |
|
|
|
| 19 |
B1 |
714 |
685 |
35.78 |
|
|
|
| 20 |
B1 |
575 |
552 |
17.80 |
|
|
|
| 21 |
B1 |
393 |
377 |
0.55 |
|
|
|
| 22 |
B1 |
147 |
141 |
1.84 |
|
|
|
| 23 |
B2 |
3226 |
3095 |
5.50 |
|
|
|
| 24 |
B2 |
3209 |
3078 |
3.51 |
|
|
|
| 25 |
B2 |
1644 |
1577 |
1.00 |
|
|
|
| 26 |
B2 |
1490 |
1430 |
8.22 |
|
|
|
| 27 |
B2 |
1368 |
1312 |
1.80 |
|
|
|
| 28 |
B2 |
1342 |
1288 |
0.60 |
|
|
|
| 29 |
B2 |
1189 |
1141 |
0.18 |
|
|
|
| 30 |
B2 |
1113 |
1068 |
4.22 |
|
|
|
| 31 |
B2 |
640 |
614 |
0.18 |
|
|
|
| 32 |
B2 |
572 |
548 |
0.24 |
|
|
|
| 33 |
B2 |
166 |
159 |
4.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21922.4 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 21027.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.124 |
|
|
|
| 2 |
C |
-0.059 |
|
|
|
| 3 |
C |
0.078 |
|
|
|
| 4 |
C |
-0.094 |
|
|
|
| 5 |
C |
-0.094 |
|
|
|
| 6 |
C |
-0.089 |
|
|
|
| 7 |
C |
-0.089 |
|
|
|
| 8 |
C |
-0.090 |
|
|
|
| 9 |
H |
0.114 |
|
|
|
| 10 |
H |
0.114 |
|
|
|
| 11 |
H |
0.111 |
|
|
|
| 12 |
H |
0.111 |
|
|
|
| 13 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.641 |
4.641 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.437 |
0.000 |
0.000 |
| y |
0.000 |
-39.052 |
0.000 |
| z |
0.000 |
0.000 |
-55.658 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.082 |
0.000 |
0.000 |
| y |
0.000 |
12.996 |
0.000 |
| z |
0.000 |
0.000 |
-11.914 |
|
| Polar |
|---|
| 3z2-r2 | -23.827 |
|---|
| x2-y2 | -9.385 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.931 |
0.000 |
0.000 |
| y |
0.000 |
11.985 |
0.000 |
| z |
0.000 |
0.000 |
17.229 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |