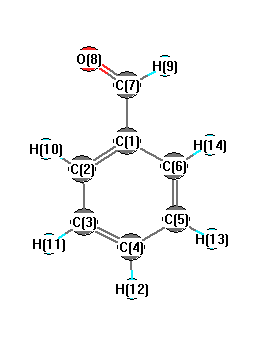
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3223 |
3085 |
10.62 |
280.29 |
0.12 |
0.22 |
| 2 |
A' |
3217 |
3079 |
11.12 |
44.79 |
0.22 |
0.37 |
| 3 |
A' |
3208 |
3070 |
9.21 |
84.91 |
0.75 |
0.86 |
| 4 |
A' |
3196 |
3058 |
1.16 |
78.20 |
0.72 |
0.84 |
| 5 |
A' |
3184 |
3047 |
4.51 |
49.63 |
0.41 |
0.59 |
| 6 |
A' |
2908 |
2783 |
119.35 |
144.25 |
0.28 |
0.44 |
| 7 |
A' |
1817 |
1739 |
260.02 |
82.68 |
0.37 |
0.54 |
| 8 |
A' |
1672 |
1600 |
27.76 |
77.24 |
0.53 |
0.69 |
| 9 |
A' |
1657 |
1586 |
10.96 |
8.49 |
0.53 |
0.69 |
| 10 |
A' |
1536 |
1470 |
0.77 |
1.44 |
0.38 |
0.56 |
| 11 |
A' |
1500 |
1435 |
13.51 |
1.63 |
0.28 |
0.43 |
| 12 |
A' |
1422 |
1361 |
7.41 |
2.27 |
0.51 |
0.68 |
| 13 |
A' |
1377 |
1318 |
5.64 |
0.99 |
0.75 |
0.85 |
| 14 |
A' |
1341 |
1284 |
16.28 |
1.51 |
0.43 |
0.60 |
| 15 |
A' |
1239 |
1186 |
59.63 |
27.36 |
0.24 |
0.39 |
| 16 |
A' |
1189 |
1138 |
17.72 |
7.64 |
0.15 |
0.26 |
| 17 |
A' |
1184 |
1133 |
2.12 |
6.60 |
0.70 |
0.83 |
| 18 |
A' |
1107 |
1059 |
5.21 |
0.79 |
0.15 |
0.26 |
| 19 |
A' |
1055 |
1010 |
2.17 |
14.57 |
0.08 |
0.15 |
| 20 |
A' |
1023 |
979 |
0.21 |
25.33 |
0.07 |
0.13 |
| 21 |
A' |
846 |
810 |
33.73 |
9.99 |
0.13 |
0.23 |
| 22 |
A' |
660 |
632 |
22.57 |
3.08 |
0.31 |
0.47 |
| 23 |
A' |
621 |
594 |
0.60 |
5.02 |
0.75 |
0.86 |
| 24 |
A' |
440 |
421 |
0.30 |
5.22 |
0.31 |
0.47 |
| 25 |
A' |
211 |
201 |
7.99 |
0.57 |
0.49 |
0.66 |
| 26 |
A" |
1045 |
1000 |
0.91 |
3.22 |
0.75 |
0.86 |
| 27 |
A" |
1028 |
984 |
0.06 |
0.29 |
0.75 |
0.86 |
| 28 |
A" |
1010 |
967 |
0.02 |
0.13 |
0.75 |
0.86 |
| 29 |
A" |
955 |
914 |
1.32 |
0.20 |
0.75 |
0.86 |
| 30 |
A" |
877 |
839 |
0.02 |
0.49 |
0.75 |
0.86 |
| 31 |
A" |
771 |
738 |
43.79 |
0.11 |
0.75 |
0.86 |
| 32 |
A" |
713 |
683 |
28.65 |
0.06 |
0.75 |
0.86 |
| 33 |
A" |
464 |
444 |
6.31 |
0.19 |
0.75 |
0.86 |
| 34 |
A" |
416 |
398 |
0.08 |
0.00 |
0.75 |
0.86 |
| 35 |
A" |
238 |
227 |
7.42 |
0.94 |
0.75 |
0.86 |
| 36 |
A" |
119 |
114 |
4.26 |
1.85 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 24232.1 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 23192.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.054 |
|
|
|
| 2 |
C |
-0.118 |
|
|
|
| 3 |
C |
-0.133 |
|
|
|
| 4 |
C |
-0.111 |
|
|
|
| 5 |
C |
-0.133 |
|
|
|
| 6 |
C |
-0.134 |
|
|
|
| 7 |
C |
0.104 |
|
|
|
| 8 |
O |
-0.269 |
|
|
|
| 9 |
H |
0.064 |
|
|
|
| 10 |
H |
0.145 |
|
|
|
| 11 |
H |
0.136 |
|
|
|
| 12 |
H |
0.136 |
|
|
|
| 13 |
H |
0.134 |
|
|
|
| 14 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.239 |
-2.429 |
0.000 |
3.303 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.182 |
6.058 |
0.000 |
| y |
6.058 |
-46.488 |
0.000 |
| z |
0.000 |
0.000 |
-48.517 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.320 |
6.058 |
0.000 |
| y |
6.058 |
-0.139 |
0.000 |
| z |
0.000 |
0.000 |
-3.182 |
|
| Polar |
|---|
| 3z2-r2 | -6.363 |
|---|
| x2-y2 | 2.306 |
|---|
| xy | 6.058 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.302 |
-1.083 |
0.000 |
| y |
-1.083 |
15.731 |
0.000 |
| z |
0.000 |
0.000 |
5.822 |
<r2> (average value of r
2) Å
2
| <r2> |
258.460 |
| (<r2>)1/2 |
16.077 |