return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5OCH3 (Anisole)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-346.182902
Energy at 298.15K-346.191994
HF Energy-344.702808
Nuclear repulsion energy348.039411
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3269 3104 2.48      
2 A' 3255 3090 9.52      
3 A' 3244 3080 9.67      
4 A' 3224 3061 6.58      
5 A' 3212 3049 1.40      
6 A' 3199 3037 17.14      
7 A' 3068 2913 42.54      
8 A' 1669 1584 71.31      
9 A' 1647 1563 14.73      
10 A' 1545 1467 43.48      
11 A' 1540 1462 77.28      
12 A' 1501 1425 7.86      
13 A' 1491 1415 9.63      
14 A' 1476 1402 7.12      
15 A' 1346 1278 0.41      
16 A' 1311 1244 222.58      
17 A' 1221 1160 6.11      
18 A' 1210 1149 10.90      
19 A' 1194 1133 2.29      
20 A' 1114 1058 13.37      
21 A' 1099 1043 38.99      
22 A' 1054 1001 1.69      
23 A' 1023 972 0.95      
24 A' 807 766 15.59      
25 A' 621 589 0.40      
26 A' 560 531 5.17      
27 A' 450 427 0.77      
28 A' 269 255 2.31      
29 A" 3133 2975 29.73      
30 A" 1527 1450 7.58      
31 A" 1198 1138 1.20      
32 A" 985 935 0.02      
33 A" 976 927 0.02      
34 A" 917 871 6.65      
35 A" 848 806 0.08      
36 A" 786 746 60.63      
37 A" 713 677 22.25      
38 A" 530 503 8.69      
39 A" 429 408 0.01      
40 A" 297 282 0.22      
41 A" 221 210 0.34      
42 A" 98 93 4.12      

Unscaled Zero Point Vibrational Energy (zpe) 29637.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 28137.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.16966 0.05320 0.04081

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.540 0.000
C2 0.910 -0.509 0.000
C3 0.444 -1.819 0.000
C4 -0.913 -2.090 0.000
C5 -1.817 -1.031 0.000
C6 -1.367 0.275 0.000
O7 0.355 1.845 0.000
C8 1.736 2.106 0.000
H9 1.968 -0.322 0.000
H10 1.156 -2.627 0.000
H11 -1.266 -3.106 0.000
H12 -2.876 -1.226 0.000
H13 -2.051 1.106 0.000
H14 1.840 3.183 0.000
H15 2.221 1.700 0.886
H16 2.221 1.700 -0.886

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 C8 H9 H10 H11 H12 H13 H14 H15 H16
C11.38852.40122.78432.40281.39281.35182.33802.14833.37213.85983.37542.12723.22002.65782.6578
C21.38851.39112.41322.77662.40782.41812.74251.07462.13283.38813.85343.37193.80712.71742.7174
C32.40121.39111.38382.39442.76843.66544.13312.13691.07712.13973.37283.84435.19364.04134.0413
C42.78432.41321.38381.39182.40754.13394.96273.38052.13831.07552.14483.39185.94844.99754.9975
C52.40282.77662.39441.39181.38133.60424.74053.85113.37472.14641.07682.14975.58004.95544.9554
C61.39282.40782.76842.40751.38132.33033.60373.38783.84553.38192.12801.07614.32953.96153.9615
O71.35182.41813.66544.13393.60422.33031.40612.70114.54335.20924.45742.51641.99922.07122.0712
C82.33802.74254.13314.96274.74053.60371.40612.43904.76896.01505.69023.91691.08171.08881.0888
H92.14831.07462.13693.38053.85113.38782.70112.43902.44424.26734.92794.26453.50702.22202.2220
H103.37212.13281.07712.13833.37473.84554.54334.76892.44422.46934.26934.92135.85034.54374.5437
H113.85983.38812.13971.07552.14643.38195.20926.01504.26732.46932.47554.28397.01416.00376.0037
H123.37543.85343.37282.14481.07682.12804.45745.69024.92794.26932.47552.47326.45615.94405.9440
H132.12723.37193.84433.39182.14971.07612.51643.91694.26454.92134.28392.47324.41064.40324.4032
H143.22003.80715.19365.94845.58004.32951.99921.08173.50705.85037.01416.45614.41061.76901.7690
H152.65782.71744.04134.99754.95543.96152.07121.08882.22204.54376.00375.94404.40321.76901.7725
H162.65782.71744.04134.99754.95543.96152.07121.08882.22204.54376.00375.94404.40321.76901.7725

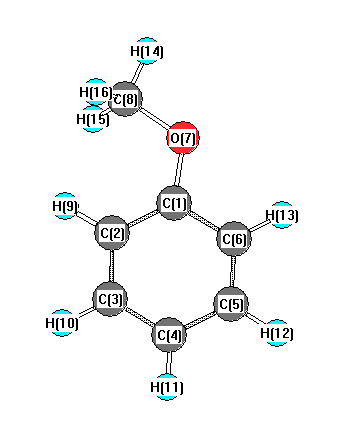
picture of Anisole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.503 C1 C2 H9 120.903
C1 C6 C5 120.023 C1 C6 H13 118.432
C1 O7 C8 115.929 C2 C1 C6 119.922
C2 C1 O7 123.862 C2 C3 C4 120.836
C2 C3 H10 119.013 C3 C2 H9 119.593
C3 C4 C5 119.231 C3 C4 H11 120.412
C4 C3 H10 120.151 C4 C5 C6 120.485
C4 C5 H12 120.104 C5 C4 H11 120.357
C5 C6 H13 121.545 C6 C1 O7 116.216
C6 C5 H12 119.411 O7 C8 H14 106.225
O7 C8 H15 111.606 O7 C8 H16 111.606
H14 C8 H15 109.182 H14 C8 H16 109.182
H15 C8 H16 108.965
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability