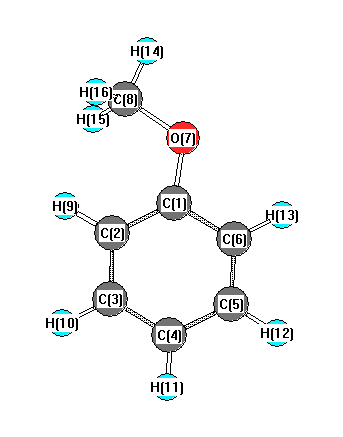
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3238 |
3099 |
5.48 |
|
|
|
| 2 |
A' |
3225 |
3087 |
10.96 |
|
|
|
| 3 |
A' |
3218 |
3080 |
18.50 |
|
|
|
| 4 |
A' |
3202 |
3064 |
11.02 |
|
|
|
| 5 |
A' |
3194 |
3057 |
1.96 |
|
|
|
| 6 |
A' |
3163 |
3027 |
21.48 |
|
|
|
| 7 |
A' |
3026 |
2896 |
48.94 |
|
|
|
| 8 |
A' |
1677 |
1605 |
82.69 |
|
|
|
| 9 |
A' |
1658 |
1587 |
17.27 |
|
|
|
| 10 |
A' |
1549 |
1482 |
105.09 |
|
|
|
| 11 |
A' |
1510 |
1445 |
27.97 |
|
|
|
| 12 |
A' |
1498 |
1434 |
9.68 |
|
|
|
| 13 |
A' |
1473 |
1409 |
2.21 |
|
|
|
| 14 |
A' |
1381 |
1321 |
22.48 |
|
|
|
| 15 |
A' |
1349 |
1291 |
15.31 |
|
|
|
| 16 |
A' |
1306 |
1250 |
213.58 |
|
|
|
| 17 |
A' |
1207 |
1156 |
12.62 |
|
|
|
| 18 |
A' |
1203 |
1152 |
1.51 |
|
|
|
| 19 |
A' |
1177 |
1127 |
2.09 |
|
|
|
| 20 |
A' |
1116 |
1068 |
9.93 |
|
|
|
| 21 |
A' |
1103 |
1055 |
49.20 |
|
|
|
| 22 |
A' |
1059 |
1013 |
1.39 |
|
|
|
| 23 |
A' |
1016 |
973 |
0.50 |
|
|
|
| 24 |
A' |
810 |
775 |
15.68 |
|
|
|
| 25 |
A' |
620 |
593 |
0.45 |
|
|
|
| 26 |
A' |
559 |
535 |
5.22 |
|
|
|
| 27 |
A' |
447 |
428 |
0.77 |
|
|
|
| 28 |
A' |
261 |
250 |
2.44 |
|
|
|
| 29 |
A" |
3087 |
2955 |
37.01 |
|
|
|
| 30 |
A" |
1488 |
1424 |
7.81 |
|
|
|
| 31 |
A" |
1178 |
1128 |
1.11 |
|
|
|
| 32 |
A" |
1004 |
961 |
0.18 |
|
|
|
| 33 |
A" |
985 |
943 |
0.03 |
|
|
|
| 34 |
A" |
911 |
872 |
7.70 |
|
|
|
| 35 |
A" |
840 |
804 |
0.07 |
|
|
|
| 36 |
A" |
777 |
744 |
54.28 |
|
|
|
| 37 |
A" |
715 |
684 |
25.75 |
|
|
|
| 38 |
A" |
524 |
501 |
9.05 |
|
|
|
| 39 |
A" |
425 |
407 |
0.02 |
|
|
|
| 40 |
A" |
268 |
256 |
0.23 |
|
|
|
| 41 |
A" |
206 |
197 |
0.38 |
|
|
|
| 42 |
A" |
100 |
95 |
3.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29375.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 28114.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.234 |
|
|
|
| 2 |
C |
-0.221 |
|
|
|
| 3 |
C |
-0.138 |
|
|
|
| 4 |
C |
-0.150 |
|
|
|
| 5 |
C |
-0.124 |
|
|
|
| 6 |
C |
-0.165 |
|
|
|
| 7 |
O |
-0.219 |
|
|
|
| 8 |
C |
-0.175 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.127 |
|
|
|
| 11 |
H |
0.127 |
|
|
|
| 12 |
H |
0.128 |
|
|
|
| 13 |
H |
0.130 |
|
|
|
| 14 |
H |
0.129 |
|
|
|
| 15 |
H |
0.097 |
|
|
|
| 16 |
H |
0.097 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.259 |
0.150 |
0.000 |
1.268 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.627 |
2.900 |
0.000 |
| y |
2.900 |
-43.890 |
0.000 |
| z |
0.000 |
0.000 |
-50.628 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.632 |
2.900 |
0.000 |
| y |
2.900 |
1.738 |
0.000 |
| z |
0.000 |
0.000 |
-8.369 |
|
| Polar |
|---|
| 3z2-r2 | -16.738 |
|---|
| x2-y2 | 3.263 |
|---|
| xy | 2.900 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.923 |
1.565 |
0.000 |
| y |
1.565 |
15.049 |
0.000 |
| z |
0.000 |
0.000 |
6.877 |
<r2> (average value of r
2) Å
2
| <r2> |
269.914 |
| (<r2>)1/2 |
16.429 |