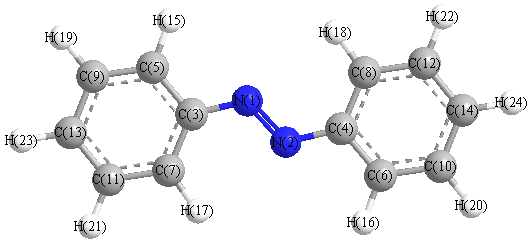
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3240 |
3101 |
0.00 |
|
|
|
| 2 |
Ag |
3225 |
3087 |
0.00 |
|
|
|
| 3 |
Ag |
3215 |
3077 |
0.00 |
|
|
|
| 4 |
Ag |
3205 |
3067 |
0.00 |
|
|
|
| 5 |
Ag |
3194 |
3057 |
0.00 |
|
|
|
| 6 |
Ag |
1676 |
1604 |
0.00 |
|
|
|
| 7 |
Ag |
1659 |
1588 |
0.00 |
|
|
|
| 8 |
Ag |
1592 |
1524 |
0.00 |
|
|
|
| 9 |
Ag |
1518 |
1453 |
0.00 |
|
|
|
| 10 |
Ag |
1495 |
1431 |
0.00 |
|
|
|
| 11 |
Ag |
1378 |
1319 |
0.00 |
|
|
|
| 12 |
Ag |
1340 |
1282 |
0.00 |
|
|
|
| 13 |
Ag |
1218 |
1166 |
0.00 |
|
|
|
| 14 |
Ag |
1180 |
1130 |
0.00 |
|
|
|
| 15 |
Ag |
1168 |
1118 |
0.00 |
|
|
|
| 16 |
Ag |
1100 |
1053 |
0.00 |
|
|
|
| 17 |
Ag |
1052 |
1007 |
0.00 |
|
|
|
| 18 |
Ag |
1023 |
979 |
0.00 |
|
|
|
| 19 |
Ag |
944 |
904 |
0.00 |
|
|
|
| 20 |
Ag |
678 |
649 |
0.00 |
|
|
|
| 21 |
Ag |
616 |
590 |
0.00 |
|
|
|
| 22 |
Ag |
303 |
290 |
0.00 |
|
|
|
| 23 |
Ag |
219 |
209 |
0.00 |
|
|
|
| 24 |
Au |
1025 |
981 |
0.36 |
|
|
|
| 25 |
Au |
1009 |
966 |
0.00 |
|
|
|
| 26 |
Au |
969 |
928 |
8.33 |
|
|
|
| 27 |
Au |
869 |
832 |
0.09 |
|
|
|
| 28 |
Au |
810 |
775 |
56.40 |
|
|
|
| 29 |
Au |
713 |
682 |
79.59 |
|
|
|
| 30 |
Au |
564 |
540 |
14.89 |
|
|
|
| 31 |
Au |
417 |
399 |
0.00 |
|
|
|
| 32 |
Au |
309 |
296 |
0.59 |
|
|
|
| 33 |
Au |
63 |
61 |
1.31 |
|
|
|
| 34 |
Au |
19 |
18 |
0.04 |
|
|
|
| 35 |
Bg |
1025 |
981 |
0.00 |
|
|
|
| 36 |
Bg |
1010 |
967 |
0.00 |
|
|
|
| 37 |
Bg |
963 |
922 |
0.00 |
|
|
|
| 38 |
Bg |
870 |
833 |
0.00 |
|
|
|
| 39 |
Bg |
788 |
754 |
0.00 |
|
|
|
| 40 |
Bg |
708 |
678 |
0.00 |
|
|
|
| 41 |
Bg |
489 |
468 |
0.00 |
|
|
|
| 42 |
Bg |
420 |
402 |
0.00 |
|
|
|
| 43 |
Bg |
252 |
241 |
0.00 |
|
|
|
| 44 |
Bg |
94 |
90 |
0.00 |
|
|
|
| 45 |
Bu |
3240 |
3101 |
9.14 |
|
|
|
| 46 |
Bu |
3225 |
3087 |
33.49 |
|
|
|
| 47 |
Bu |
3215 |
3077 |
33.36 |
|
|
|
| 48 |
Bu |
3205 |
3067 |
17.02 |
|
|
|
| 49 |
Bu |
3194 |
3057 |
2.35 |
|
|
|
| 50 |
Bu |
1668 |
1596 |
2.07 |
|
|
|
| 51 |
Bu |
1653 |
1582 |
4.54 |
|
|
|
| 52 |
Bu |
1530 |
1465 |
12.65 |
|
|
|
| 53 |
Bu |
1496 |
1432 |
13.19 |
|
|
|
| 54 |
Bu |
1375 |
1316 |
8.71 |
|
|
|
| 55 |
Bu |
1335 |
1277 |
1.98 |
|
|
|
| 56 |
Bu |
1271 |
1216 |
19.65 |
|
|
|
| 57 |
Bu |
1180 |
1130 |
0.41 |
|
|
|
| 58 |
Bu |
1174 |
1124 |
26.13 |
|
|
|
| 59 |
Bu |
1104 |
1057 |
13.59 |
|
|
|
| 60 |
Bu |
1051 |
1006 |
12.84 |
|
|
|
| 61 |
Bu |
1022 |
978 |
3.25 |
|
|
|
| 62 |
Bu |
844 |
808 |
0.45 |
|
|
|
| 63 |
Bu |
624 |
597 |
0.78 |
|
|
|
| 64 |
Bu |
541 |
517 |
4.17 |
|
|
|
| 65 |
Bu |
525 |
503 |
24.88 |
|
|
|
| 66 |
Bu |
66 |
63 |
1.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 42079.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 40273.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.116 |
|
|
|
| 2 |
N |
-0.116 |
|
|
|
| 3 |
C |
0.110 |
|
|
|
| 4 |
C |
0.110 |
|
|
|
| 5 |
C |
-0.122 |
|
|
|
| 6 |
C |
-0.122 |
|
|
|
| 7 |
C |
-0.139 |
|
|
|
| 8 |
C |
-0.139 |
|
|
|
| 9 |
C |
-0.138 |
|
|
|
| 10 |
C |
-0.138 |
|
|
|
| 11 |
C |
-0.132 |
|
|
|
| 12 |
C |
-0.132 |
|
|
|
| 13 |
C |
-0.116 |
|
|
|
| 14 |
C |
-0.116 |
|
|
|
| 15 |
H |
0.129 |
|
|
|
| 16 |
H |
0.129 |
|
|
|
| 17 |
H |
0.127 |
|
|
|
| 18 |
H |
0.127 |
|
|
|
| 19 |
H |
0.132 |
|
|
|
| 20 |
H |
0.132 |
|
|
|
| 21 |
H |
0.133 |
|
|
|
| 22 |
H |
0.133 |
|
|
|
| 23 |
H |
0.132 |
|
|
|
| 24 |
H |
0.132 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-66.593 |
4.751 |
0.000 |
| y |
4.751 |
-68.960 |
0.000 |
| z |
0.000 |
0.000 |
-85.899 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.836 |
4.751 |
0.000 |
| y |
4.751 |
7.286 |
0.000 |
| z |
0.000 |
0.000 |
-18.122 |
|
| Polar |
|---|
| 3z2-r2 | -36.244 |
|---|
| x2-y2 | 2.367 |
|---|
| xy | 4.751 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
34.862 |
10.433 |
0.000 |
| y |
10.433 |
30.607 |
0.000 |
| z |
0.000 |
0.000 |
10.143 |
<r2> (average value of r
2) Å
2
| <r2> |
1082.430 |
| (<r2>)1/2 |
32.900 |