return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C9H12 (n-propyl benzene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-349.544544
Energy at 298.15K-349.557269
HF Energy-347.924963
Nuclear repulsion energy414.285582
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3250 3085 11.02      
2 A' 3220 3057 4.01      
3 A' 3197 3035 6.66      
4 A' 3165 3005 25.11      
5 A' 3091 2934 46.74      
6 A' 3086 2930 14.16      
7 A' 3074 2918 12.89      
8 A' 1665 1581 4.77      
9 A' 1537 1459 12.63      
10 A' 1532 1454 6.52      
11 A' 1515 1439 0.47      
12 A' 1500 1424 0.64      
13 A' 1421 1350 2.33      
14 A' 1388 1317 0.97      
15 A' 1316 1249 0.48      
16 A' 1247 1184 0.37      
17 A' 1213 1152 0.08      
18 A' 1133 1076 6.65      
19 A' 1083 1028 0.21      
20 A' 1059 1006 3.28      
21 A' 1029 977 0.17      
22 A' 991 940 0.04      
23 A' 948 900 0.62      
24 A' 919 873 2.01      
25 A' 836 793 1.42      
26 A' 780 740 36.43      
27 A' 722 686 31.33      
28 A' 596 566 4.32      
29 A' 513 487 8.07      
30 A' 315 299 0.02      
31 A' 280 266 0.55      
32 A' 103 98 0.31      
33 A" 3232 3068 22.57      
34 A" 3202 3040 3.80      
35 A" 3158 2998 34.43      
36 A" 3127 2969 14.00      
37 A" 3104 2947 0.74      
38 A" 1641 1558 0.43      
39 A" 1527 1449 7.20      
40 A" 1492 1417 4.66      
41 A" 1480 1405 2.22      
42 A" 1359 1290 0.66      
43 A" 1336 1268 0.23      
44 A" 1261 1197 0.40      
45 A" 1196 1135 0.02      
46 A" 1134 1076 4.17      
47 A" 1076 1021 1.50      
48 A" 986 936 0.01      
49 A" 885 840 0.52      
50 A" 874 830 0.16      
51 A" 756 717 1.11      
52 A" 629 597 0.02      
53 A" 417 396 0.00      
54 A" 346 329 0.06      
55 A" 259 246 0.01      
56 A" 93 89 0.02      
57 A" 46 43 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 41167.3 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 39084.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.14647 0.02912 0.02604

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.393 -0.725 1.195
C2 0.393 -2.112 1.200
C3 0.393 -2.811 0.000
C4 0.393 -2.112 -1.200
C5 0.393 -0.725 -1.195
C6 0.396 -0.014 0.000
C7 0.347 1.478 0.000
C8 -1.082 1.997 0.000
C9 -1.137 3.512 0.000
H10 0.395 -0.187 2.130
H11 0.396 -2.645 2.136
H12 0.396 -3.887 0.000
H13 0.396 -2.645 -2.136
H14 0.395 -0.187 -2.130
H15 0.867 1.862 0.876
H16 0.867 1.862 -0.876
H17 -1.599 1.605 -0.872
H18 -1.599 1.605 0.872
H19 -2.162 3.868 0.000
H20 -0.641 3.916 0.878
H21 -0.641 3.916 -0.878

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21
C11.38722.40402.76762.39041.39052.50673.31854.66061.07872.13813.38053.84463.36882.64943.34813.69713.08185.38994.76545.1871
C21.38721.38832.39932.76762.41673.78564.52775.95082.13821.07702.14233.37753.84644.01554.50894.69844.22946.61276.12486.4595
C32.40401.38831.38832.40402.79654.28915.02906.50533.37962.14201.07632.14203.37964.77804.77804.92184.92187.15086.86246.8624
C42.76762.39931.38831.38722.41673.78564.52775.95083.84643.37752.14231.07702.13824.50894.01554.22944.69846.61276.45956.1248
C52.39042.76762.40401.38721.39052.50673.31854.66063.36883.84463.38052.13811.07873.34812.64943.08183.69715.38995.18714.7654
C61.39052.41672.79652.41671.39051.49312.49573.84512.13743.38853.87283.38852.13742.12402.12402.71302.71304.64924.15864.1586
C72.50673.78564.28913.78562.50671.49311.52042.51812.70444.64375.36544.64372.70441.08881.08882.13662.13663.46562.77342.7734
C83.31854.52775.02904.52773.31852.49571.52041.51593.38995.31946.06705.31943.38992.14092.14091.08761.08762.16042.15582.1558
C94.66065.95086.50535.95084.66063.84512.51811.51594.53556.69507.55636.69504.53552.73982.73982.14772.14771.08541.08641.0864
H101.07872.13823.37963.84643.36882.13742.70443.38994.53552.45804.26944.92344.26072.44843.66924.02552.96135.24634.41365.1924
H112.13811.07702.14203.37753.84463.38854.64375.31946.69502.45802.47044.27114.92344.70355.44155.57574.86177.31616.76087.2943
H123.38052.14231.07632.14233.38053.87285.36546.06707.55634.26942.47042.47044.26945.83475.83475.90765.90768.16607.92077.9207
H133.84463.37752.14201.07702.13813.38854.64375.31946.69504.92344.27112.47042.45805.44154.70354.86175.57577.31617.29436.7608
H143.36883.84643.37962.13821.07872.13742.70443.38994.53554.26074.92344.26942.45803.66922.44842.96134.02555.24635.19244.4136
H152.64944.01554.77804.50893.34812.12401.08882.14092.73982.44844.70355.83475.44153.66921.75283.03402.47923.73742.54813.0936
H163.34814.50894.77804.01552.64942.12401.08882.14092.73983.66925.44155.83474.70352.44841.75282.47923.03403.73743.09362.5481
H173.69714.69844.92184.22943.08182.71302.13661.08762.14774.02555.57575.90764.86172.96133.03402.47921.74502.49013.05372.5022
H183.08184.22944.92184.69843.69712.71302.13661.08762.14772.96134.86175.90765.57574.02552.47923.03401.74502.49012.50223.0537
H195.38996.61277.15086.61275.38994.64923.46562.16041.08545.24637.31618.16607.31615.24633.73743.73742.49012.49011.75741.7574
H204.76546.12486.86246.45955.18714.15862.77342.15581.08644.41366.76087.92077.29435.19242.54813.09363.05372.50221.75741.7558
H215.18716.45956.86246.12484.76544.15862.77342.15581.08645.19247.29437.92076.76084.41363.09362.54812.50223.05371.75741.7558

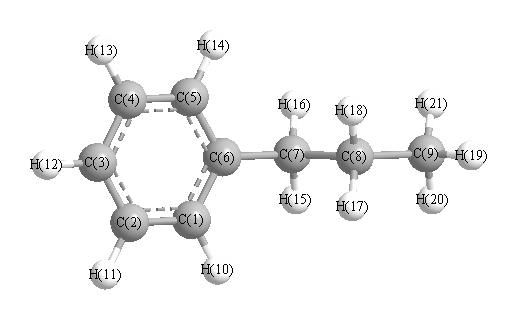
picture of n-propyl benzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.031 C1 C2 H11 119.846
C1 C6 C5 118.529 C1 C6 C7 120.713
C2 C1 C6 120.919 C2 C1 H10 119.716
C2 C3 C4 119.570 C2 C3 H12 120.215
C3 C2 H11 120.123 C3 C4 C5 120.031
C3 C4 H13 120.123 C4 C3 H12 120.215
C4 C5 C6 120.919 C4 C5 H14 119.716
C5 C4 H13 119.846 C5 C6 C7 120.713
C6 C1 H10 119.365 C6 C5 H14 119.365
C6 C7 C8 111.821 C6 C7 H15 109.702
C6 C7 H16 109.702 C7 C8 C9 112.064
C7 C8 H17 108.889 C7 C8 H18 108.889
C8 C7 H15 109.152 C8 C7 H16 109.152
C8 C9 H19 111.229 C8 C9 H20 110.796
C8 C9 H21 110.796 C9 C8 H17 110.078
C9 C8 H18 110.078 H15 C7 H16 107.198
H17 C8 H18 106.684 H19 C9 H20 108.036
H19 C9 H21 108.036 H20 C9 H21 107.816
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability