Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -5223.946811 |
| Energy at 298.15K | -5223.957079 |
| HF Energy | -5223.022354 |
| Nuclear repulsion energy | 422.780805 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3155 |
2995 |
0.00 |
|
|
|
| 2 |
Ag |
1498 |
1422 |
0.00 |
|
|
|
| 3 |
Ag |
1310 |
1244 |
0.00 |
|
|
|
| 4 |
Ag |
1102 |
1047 |
0.00 |
|
|
|
| 5 |
Ag |
720 |
684 |
0.00 |
|
|
|
| 6 |
Ag |
199 |
188 |
0.00 |
|
|
|
| 7 |
Au |
3225 |
3062 |
0.00 |
|
|
|
| 8 |
Au |
1134 |
1077 |
3.75 |
|
|
|
| 9 |
Au |
761 |
723 |
3.65 |
|
|
|
| 10 |
Au |
115 |
110 |
4.01 |
|
|
|
| 11 |
Bg |
3201 |
3039 |
0.00 |
|
|
|
| 12 |
Bg |
1310 |
1244 |
0.00 |
|
|
|
| 13 |
Bg |
964 |
916 |
0.00 |
|
|
|
| 14 |
Bu |
3164 |
3004 |
5.73 |
|
|
|
| 15 |
Bu |
1499 |
1423 |
7.75 |
|
|
|
| 16 |
Bu |
1230 |
1168 |
48.34 |
|
|
|
| 17 |
Bu |
653 |
620 |
47.47 |
|
|
|
| 18 |
Bu |
183 |
174 |
5.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12711.6 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 12068.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.489 |
0.568 |
0.000 |
| C2 |
-0.489 |
-0.568 |
0.000 |
| Br3 |
-0.489 |
2.232 |
0.000 |
| Br4 |
0.489 |
-2.232 |
0.000 |
| H5 |
1.109 |
0.561 |
0.885 |
| H6 |
1.109 |
0.561 |
-0.885 |
| H7 |
-1.109 |
-0.561 |
0.885 |
| H8 |
-1.109 |
-0.561 |
-0.885 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4990 | 1.9296 | 2.8007 | 1.0806 | 1.0806 | 2.1472 | 2.1472 |
C2 | 1.4990 | | 2.8007 | 1.9296 | 2.1472 | 2.1472 | 1.0806 | 1.0806 | Br3 | 1.9296 | 2.8007 | | 4.5704 | 2.4756 | 2.4756 | 2.9951 | 2.9951 | Br4 | 2.8007 | 1.9296 | 4.5704 | | 2.9951 | 2.9951 | 2.4756 | 2.4756 | H5 | 1.0806 | 2.1472 | 2.4756 | 2.9951 | | 1.7693 | 2.4858 | 3.0512 | H6 | 1.0806 | 2.1472 | 2.4756 | 2.9951 | 1.7693 | | 3.0512 | 2.4858 | H7 | 2.1472 | 1.0806 | 2.9951 | 2.4756 | 2.4858 | 3.0512 | | 1.7693 | H8 | 2.1472 | 1.0806 | 2.9951 | 2.4756 | 3.0512 | 2.4858 | 1.7693 | |
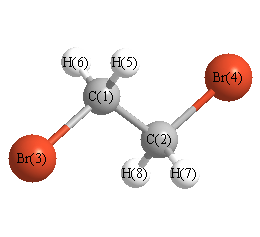 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.897 |
|
C1 |
C2 |
H7 |
111.662 |
| C1 |
C2 |
H8 |
111.662 |
|
C2 |
C1 |
Br3 |
108.897 |
| C2 |
C1 |
H5 |
111.662 |
|
C2 |
C1 |
H6 |
111.662 |
| Br3 |
C1 |
H5 |
107.257 |
|
Br3 |
C1 |
H6 |
107.257 |
| Br4 |
C2 |
H7 |
107.257 |
|
Br4 |
C2 |
H8 |
107.257 |
| H5 |
C1 |
H6 |
109.904 |
|
H7 |
C2 |
H8 |
109.904 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -5223.944065 |
| Energy at 298.15K | |
| HF Energy | -5223.017946 |
| Nuclear repulsion energy | 459.836076 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3187 |
3026 |
0.03 |
85.76 |
0.74 |
0.85 |
| 2 |
A |
3141 |
2982 |
13.64 |
218.32 |
0.01 |
0.03 |
| 3 |
A |
1485 |
1410 |
0.34 |
3.50 |
0.69 |
0.82 |
| 4 |
A |
1330 |
1263 |
17.35 |
1.66 |
0.43 |
0.60 |
| 5 |
A |
1218 |
1156 |
1.73 |
7.36 |
0.73 |
0.84 |
| 6 |
A |
1068 |
1014 |
0.42 |
1.74 |
0.72 |
0.84 |
| 7 |
A |
933 |
886 |
5.89 |
4.51 |
0.38 |
0.55 |
| 8 |
A |
603 |
572 |
5.12 |
16.79 |
0.06 |
0.11 |
| 9 |
A |
238 |
226 |
0.92 |
1.27 |
0.51 |
0.67 |
| 10 |
A |
88 |
84 |
0.16 |
0.93 |
0.70 |
0.83 |
| 11 |
B |
3200 |
3038 |
0.54 |
25.60 |
0.75 |
0.86 |
| 12 |
B |
3135 |
2977 |
2.10 |
40.78 |
0.75 |
0.86 |
| 13 |
B |
1481 |
1406 |
12.00 |
9.91 |
0.75 |
0.86 |
| 14 |
B |
1302 |
1236 |
49.78 |
2.36 |
0.75 |
0.86 |
| 15 |
B |
1147 |
1089 |
2.36 |
3.13 |
0.75 |
0.86 |
| 16 |
B |
864 |
821 |
15.04 |
1.28 |
0.75 |
0.86 |
| 17 |
B |
635 |
603 |
8.55 |
7.37 |
0.75 |
0.86 |
| 18 |
B |
365 |
346 |
4.23 |
1.72 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12710.1 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 12067.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.307 |
0.684 |
1.181 |
| C2 |
-0.307 |
-0.684 |
1.181 |
| Br3 |
-0.307 |
1.753 |
-0.294 |
| Br4 |
0.307 |
-1.753 |
-0.294 |
| H5 |
0.023 |
1.214 |
2.083 |
| H6 |
1.385 |
0.634 |
1.110 |
| H7 |
-0.023 |
-1.214 |
2.083 |
| H8 |
-1.385 |
-0.634 |
1.110 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5004 | 1.9223 | 2.8487 | 1.0833 | 1.0814 | 2.1274 | 2.1465 |
C2 | 1.5004 | | 2.8487 | 1.9223 | 2.1274 | 2.1465 | 1.0833 | 1.0814 | Br3 | 1.9223 | 2.8487 | | 3.5589 | 2.4589 | 2.4672 | 3.8118 | 2.9711 | Br4 | 2.8487 | 1.9223 | 3.5589 | | 3.8118 | 2.9711 | 2.4589 | 2.4672 | H5 | 1.0833 | 2.1274 | 2.4589 | 3.8118 | | 1.7712 | 2.4288 | 2.5190 | H6 | 1.0814 | 2.1465 | 2.4672 | 2.9711 | 1.7712 | | 2.5190 | 3.0466 | H7 | 2.1274 | 1.0833 | 3.8118 | 2.4589 | 2.4288 | 2.5190 | | 1.7712 | H8 | 2.1465 | 1.0814 | 2.9711 | 2.4672 | 2.5190 | 3.0466 | 1.7712 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.085 |
|
C1 |
C2 |
H7 |
109.803 |
| C1 |
C2 |
H8 |
111.453 |
|
C2 |
C1 |
Br3 |
112.085 |
| C2 |
C1 |
H5 |
109.803 |
|
C2 |
C1 |
H6 |
111.453 |
| Br3 |
C1 |
H5 |
106.422 |
|
Br3 |
C1 |
H6 |
107.100 |
| Br4 |
C2 |
H7 |
106.422 |
|
Br4 |
C2 |
H8 |
107.100 |
| H5 |
C1 |
H6 |
109.818 |
|
H7 |
C2 |
H8 |
109.818 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
