Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -5227.712526 |
| Energy at 298.15K | -5227.722685 |
| HF Energy | -5227.712526 |
| Nuclear repulsion energy | 419.742991 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3133 |
2999 |
0.00 |
|
|
|
| 2 |
Ag |
1488 |
1424 |
0.00 |
|
|
|
| 3 |
Ag |
1280 |
1225 |
0.00 |
|
|
|
| 4 |
Ag |
1087 |
1040 |
0.00 |
|
|
|
| 5 |
Ag |
678 |
649 |
0.00 |
|
|
|
| 6 |
Ag |
189 |
181 |
0.00 |
|
|
|
| 7 |
Au |
3217 |
3079 |
0.16 |
|
|
|
| 8 |
Au |
1104 |
1056 |
3.12 |
|
|
|
| 9 |
Au |
757 |
725 |
4.22 |
|
|
|
| 10 |
Au |
103 |
99 |
4.39 |
|
|
|
| 11 |
Bg |
3194 |
3057 |
0.00 |
|
|
|
| 12 |
Bg |
1287 |
1231 |
0.00 |
|
|
|
| 13 |
Bg |
946 |
905 |
0.00 |
|
|
|
| 14 |
Bu |
3142 |
3007 |
5.57 |
|
|
|
| 15 |
Bu |
1484 |
1420 |
8.03 |
|
|
|
| 16 |
Bu |
1205 |
1153 |
49.99 |
|
|
|
| 17 |
Bu |
612 |
585 |
69.05 |
|
|
|
| 18 |
Bu |
173 |
166 |
6.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12538.4 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 12000.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.490 |
0.567 |
0.000 |
| C2 |
-0.490 |
-0.567 |
0.000 |
| Br3 |
-0.490 |
2.250 |
0.000 |
| Br4 |
0.490 |
-2.250 |
0.000 |
| H5 |
1.111 |
0.572 |
0.887 |
| H6 |
1.111 |
0.572 |
-0.887 |
| H7 |
-1.111 |
-0.572 |
0.887 |
| H8 |
-1.111 |
-0.572 |
-0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4983 | 1.9485 | 2.8169 | 1.0826 | 1.0826 | 2.1553 | 2.1553 |
C2 | 1.4983 | | 2.8169 | 1.9485 | 2.1553 | 2.1553 | 1.0826 | 1.0826 | Br3 | 1.9485 | 2.8169 | | 4.6063 | 2.4834 | 2.4834 | 3.0226 | 3.0226 | Br4 | 2.8169 | 1.9485 | 4.6063 | | 3.0226 | 3.0226 | 2.4834 | 2.4834 | H5 | 1.0826 | 2.1553 | 2.4834 | 3.0226 | | 1.7744 | 2.4983 | 3.0643 | H6 | 1.0826 | 2.1553 | 2.4834 | 3.0226 | 1.7744 | | 3.0643 | 2.4983 | H7 | 2.1553 | 1.0826 | 3.0226 | 2.4834 | 2.4983 | 3.0643 | | 1.7744 | H8 | 2.1553 | 1.0826 | 3.0226 | 2.4834 | 3.0643 | 2.4983 | 1.7744 | |
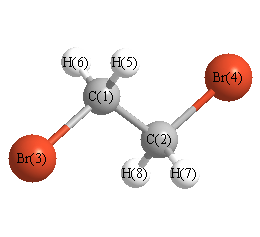 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.919 |
|
C1 |
C2 |
H7 |
112.250 |
| C1 |
C2 |
H8 |
112.250 |
|
C2 |
C1 |
Br3 |
108.919 |
| C2 |
C1 |
H5 |
112.250 |
|
C2 |
C1 |
H6 |
112.250 |
| Br3 |
C1 |
H5 |
106.509 |
|
Br3 |
C1 |
H6 |
106.509 |
| Br4 |
C2 |
H7 |
106.509 |
|
Br4 |
C2 |
H8 |
106.509 |
| H5 |
C1 |
H6 |
110.069 |
|
H7 |
C2 |
H8 |
110.069 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.209 |
|
|
|
| 2 |
C |
-0.209 |
|
|
|
| 3 |
Br |
-0.121 |
|
|
|
| 4 |
Br |
-0.121 |
|
|
|
| 5 |
H |
0.165 |
|
|
|
| 6 |
H |
0.165 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.165 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.674 |
1.340 |
0.000 |
| y |
1.340 |
-55.532 |
0.000 |
| z |
0.000 |
0.000 |
-49.259 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.722 |
1.340 |
0.000 |
| y |
1.340 |
-6.566 |
0.000 |
| z |
0.000 |
0.000 |
2.844 |
|
| Polar |
|---|
| 3z2-r2 | 5.688 |
|---|
| x2-y2 | 6.858 |
|---|
| xy | 1.340 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.720 |
-1.910 |
0.000 |
| y |
-1.910 |
13.590 |
0.000 |
| z |
0.000 |
0.000 |
6.573 |
<r2> (average value of r
2) Å
2
| <r2> |
419.383 |
| (<r2>)1/2 |
20.479 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -5227.709149 |
| Energy at 298.15K | |
| HF Energy | -5227.709149 |
| Nuclear repulsion energy | 455.159163 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3173 |
3036 |
0.01 |
91.44 |
0.75 |
0.86 |
| 2 |
A |
3113 |
2979 |
14.36 |
247.88 |
0.02 |
0.03 |
| 3 |
A |
1465 |
1402 |
0.80 |
3.20 |
0.74 |
0.85 |
| 4 |
A |
1308 |
1252 |
18.47 |
3.10 |
0.39 |
0.56 |
| 5 |
A |
1195 |
1144 |
1.47 |
8.78 |
0.75 |
0.86 |
| 6 |
A |
1047 |
1002 |
0.96 |
1.90 |
0.61 |
0.76 |
| 7 |
A |
912 |
873 |
7.56 |
5.52 |
0.37 |
0.54 |
| 8 |
A |
576 |
551 |
7.31 |
18.93 |
0.06 |
0.11 |
| 9 |
A |
224 |
215 |
1.13 |
1.39 |
0.44 |
0.62 |
| 10 |
A |
76 |
73 |
0.21 |
1.06 |
0.70 |
0.82 |
| 11 |
B |
3187 |
3050 |
0.87 |
28.64 |
0.75 |
0.86 |
| 12 |
B |
3105 |
2972 |
2.00 |
47.37 |
0.75 |
0.86 |
| 13 |
B |
1461 |
1398 |
12.48 |
10.67 |
0.75 |
0.86 |
| 14 |
B |
1276 |
1221 |
60.01 |
2.97 |
0.75 |
0.86 |
| 15 |
B |
1125 |
1077 |
1.78 |
3.17 |
0.75 |
0.86 |
| 16 |
B |
848 |
811 |
18.91 |
1.26 |
0.75 |
0.86 |
| 17 |
B |
607 |
581 |
13.95 |
8.70 |
0.75 |
0.86 |
| 18 |
B |
354 |
339 |
5.80 |
2.06 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12525.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 11988.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.306 |
0.684 |
1.183 |
| C2 |
-0.306 |
-0.684 |
1.183 |
| Br3 |
-0.306 |
1.780 |
-0.294 |
| Br4 |
0.306 |
-1.780 |
-0.294 |
| H5 |
0.019 |
1.216 |
2.085 |
| H6 |
1.387 |
0.646 |
1.110 |
| H7 |
-0.019 |
-1.216 |
2.085 |
| H8 |
-1.387 |
-0.646 |
1.110 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4995 | 1.9384 | 2.8730 | 1.0860 | 1.0833 | 2.1284 | 2.1544 |
C2 | 1.4995 | | 2.8730 | 1.9384 | 2.1284 | 2.1544 | 1.0860 | 1.0833 | Br3 | 1.9384 | 2.8730 | | 3.6130 | 2.4670 | 2.4748 | 3.8364 | 3.0041 | Br4 | 2.8730 | 1.9384 | 3.6130 | | 3.8364 | 3.0041 | 2.4670 | 2.4748 | H5 | 1.0860 | 2.1284 | 2.4670 | 3.8364 | | 1.7734 | 2.4314 | 2.5285 | H6 | 1.0833 | 2.1544 | 2.4748 | 3.0041 | 1.7734 | | 2.5285 | 3.0595 | H7 | 2.1284 | 1.0860 | 3.8364 | 2.4670 | 2.4314 | 2.5285 | | 1.7734 | H8 | 2.1544 | 1.0833 | 3.0041 | 2.4748 | 2.5285 | 3.0595 | 1.7734 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.755 |
|
C1 |
C2 |
H7 |
109.782 |
| C1 |
C2 |
H8 |
112.042 |
|
C2 |
C1 |
Br3 |
112.755 |
| C2 |
C1 |
H5 |
109.782 |
|
C2 |
C1 |
H6 |
112.042 |
| Br3 |
C1 |
H5 |
105.842 |
|
Br3 |
C1 |
H6 |
106.515 |
| Br4 |
C2 |
H7 |
105.842 |
|
Br4 |
C2 |
H8 |
106.515 |
| H5 |
C1 |
H6 |
109.664 |
|
H7 |
C2 |
H8 |
109.664 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.216 |
|
|
|
| 2 |
C |
-0.216 |
|
|
|
| 3 |
Br |
-0.100 |
|
|
|
| 4 |
Br |
-0.100 |
|
|
|
| 5 |
H |
0.154 |
|
|
|
| 6 |
H |
0.162 |
|
|
|
| 7 |
H |
0.154 |
|
|
|
| 8 |
H |
0.162 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.615 |
2.615 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.052 |
0.904 |
0.000 |
| y |
0.904 |
-53.500 |
0.000 |
| z |
0.000 |
0.000 |
-44.894 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.144 |
0.904 |
0.000 |
| y |
0.904 |
-6.526 |
0.000 |
| z |
0.000 |
0.000 |
6.382 |
|
| Polar |
|---|
| 3z2-r2 | 12.764 |
|---|
| x2-y2 | 4.447 |
|---|
| xy | 0.904 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.995 |
-0.737 |
0.000 |
| y |
-0.737 |
10.382 |
0.000 |
| z |
0.000 |
0.000 |
9.033 |
<r2> (average value of r
2) Å
2
| <r2> |
307.508 |
| (<r2>)1/2 |
17.536 |
