return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2Br (n-propyl bromide)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-2691.043656
Energy at 298.15K-2691.054177
HF Energy-2690.190400
Nuclear repulsion energy238.711627
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3172 3012 19.29      
2 A' 3137 2978 16.57      
3 A' 3110 2953 7.06      
4 A' 3092 2935 18.21      
5 A' 1531 1454 6.60      
6 A' 1514 1438 1.01      
7 A' 1499 1423 2.09      
8 A' 1424 1352 2.77      
9 A' 1379 1309 4.93      
10 A' 1269 1205 33.15      
11 A' 1140 1083 1.01      
12 A' 1075 1021 1.01      
13 A' 929 882 8.51      
14 A' 701 666 15.67      
15 A' 320 304 0.79      
16 A' 214 204 1.06      
17 A" 3189 3027 10.35      
18 A" 3164 3004 21.38      
19 A" 3138 2980 0.26      
20 A" 1527 1450 8.39      
21 A" 1341 1274 0.03      
22 A" 1260 1196 0.46      
23 A" 1078 1024 2.02      
24 A" 866 822 0.00      
25 A" 756 718 3.45      
26 A" 252 240 0.02      
27 A" 122 116 0.75      

Unscaled Zero Point Vibrational Energy (zpe) 21100.1 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 20032.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.85955 0.05576 0.05392

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.823 0.000
C2 1.496 0.644 0.000
C3 2.196 1.992 0.000
Br4 -0.919 -0.874 0.000
H5 -0.339 1.351 0.882
H6 -0.339 1.351 -0.882
H7 1.786 0.067 0.873
H8 1.786 0.067 -0.873
H9 3.273 1.870 0.000
H10 1.925 2.571 -0.878
H11 1.925 2.571 0.878

Atom - Atom Distances (Å)
  C1 C2 C3 Br4 H5 H6 H7 H8 H9 H10 H11
C11.50652.48731.93001.08251.08252.12672.12673.43672.74412.7441
C21.50651.51862.85252.15502.15501.08581.08582.15922.16042.1604
C32.48731.51864.23262.75882.75882.15272.15271.08461.08611.0861
Br41.93002.85254.23262.46322.46322.99412.99415.01064.55254.5525
H51.08252.15502.75882.46321.76422.48263.04023.75423.11582.5710
H61.08252.15502.75882.46321.76423.04022.48263.75422.57103.1158
H72.12671.08582.15272.99412.48263.04021.74562.49483.05832.5075
H82.12671.08582.15272.99413.04022.48261.74562.49482.50753.0583
H93.43672.15921.08465.01063.75423.75422.49482.49481.75541.7554
H102.74412.16041.08614.55253.11582.57103.05832.50751.75541.7561
H112.74412.16041.08614.55252.57103.11582.50753.05831.75541.7561

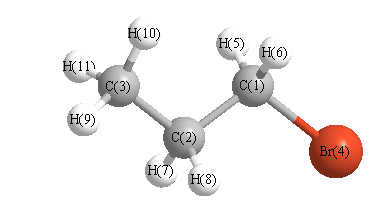
picture of n-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 110.616 C1 C2 H7 109.174
C1 C2 H8 109.174 C2 C1 Br4 111.615
C2 C1 H5 111.631 C2 C1 H6 111.631
C2 C3 H9 110.981 C2 C3 H10 110.987
C2 C3 H11 110.987 C3 C2 H7 110.395
C3 C2 H8 110.395 Br4 C1 H5 106.266
Br4 C1 H6 106.266 H5 C1 H6 109.148
H7 C2 H8 107.004 H9 C3 H10 107.929
H9 C3 H11 107.929 H10 C3 H11 107.890
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability