Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -158.138408 |
| Energy at 298.15K | -158.149133 |
| HF Energy | -157.355677 |
| Nuclear repulsion energy | 131.652479 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
123 |
116 |
0.01 |
|
|
|
|
Au |
238 |
226 |
0.00 |
|
|
|
|
Bu |
257 |
244 |
0.01 |
|
|
|
|
Bg |
278 |
264 |
0.00 |
|
|
|
|
Ag |
430 |
409 |
0.00 |
|
|
|
|
Au |
749 |
711 |
3.09 |
|
|
|
|
Bg |
818 |
777 |
0.00 |
|
|
|
|
Ag |
866 |
822 |
0.00 |
|
|
|
|
Au |
973 |
924 |
0.81 |
|
|
|
|
Bu |
992 |
942 |
5.59 |
|
|
|
|
Bu |
1055 |
1002 |
0.07 |
|
|
|
|
Ag |
1108 |
1051 |
0.00 |
|
|
|
|
Ag |
1191 |
1131 |
0.00 |
|
|
|
|
Bg |
1222 |
1160 |
0.00 |
|
|
|
|
Au |
1305 |
1239 |
0.22 |
|
|
|
|
Bu |
1330 |
1263 |
1.56 |
|
|
|
|
Bg |
1351 |
1282 |
0.00 |
|
|
|
|
Ag |
1409 |
1338 |
0.00 |
|
|
|
|
Ag |
1422 |
1350 |
0.00 |
|
|
|
|
Bu |
1424 |
1352 |
6.80 |
|
|
|
|
Ag |
1508 |
1431 |
0.00 |
|
|
|
|
Bu |
1516 |
1439 |
1.13 |
|
|
|
|
Bg |
1527 |
1449 |
0.00 |
|
|
|
|
Au |
1529 |
1451 |
14.83 |
|
|
|
|
Ag |
1531 |
1454 |
0.00 |
|
|
|
|
Bu |
1535 |
1457 |
9.29 |
|
|
|
|
Ag |
3076 |
2921 |
0.00 |
|
|
|
|
Bu |
3084 |
2928 |
18.64 |
|
|
|
|
Ag |
3087 |
2931 |
0.00 |
|
|
|
|
Bu |
3088 |
2932 |
84.09 |
|
|
|
|
Bg |
3097 |
2940 |
0.00 |
|
|
|
|
Au |
3120 |
2962 |
19.21 |
|
|
|
|
Bg |
3156 |
2997 |
0.00 |
|
|
|
|
Au |
3159 |
2999 |
77.71 |
|
|
|
|
Ag |
3164 |
3004 |
0.00 |
|
|
|
|
Bu |
3164 |
3004 |
56.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29440.3 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 27950.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.420 |
0.631 |
0.000 |
| C2 |
0.420 |
-0.631 |
0.000 |
| C3 |
0.420 |
1.893 |
0.000 |
| C4 |
-0.420 |
-1.893 |
0.000 |
| H5 |
-1.072 |
0.623 |
0.872 |
| H6 |
-1.072 |
0.623 |
-0.872 |
| H7 |
1.072 |
-0.623 |
0.872 |
| H8 |
1.072 |
-0.623 |
-0.872 |
| H9 |
-0.196 |
2.787 |
0.000 |
| H10 |
1.061 |
1.929 |
0.877 |
| H11 |
1.061 |
1.929 |
-0.877 |
| H12 |
0.196 |
-2.787 |
0.000 |
| H13 |
-1.061 |
-1.929 |
0.877 |
| H14 |
-1.061 |
-1.929 |
-0.877 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5165 | 1.5167 | 2.5245 | 1.0885 | 1.0885 | 2.1356 | 2.1356 | 2.1673 | 2.1555 | 2.1555 | 3.4729 | 2.7805 | 2.7805 |
C2 | 1.5165 | | 2.5245 | 1.5167 | 2.1356 | 2.1356 | 1.0885 | 1.0885 | 3.4729 | 2.7805 | 2.7805 | 2.1673 | 2.1555 | 2.1555 | C3 | 1.5167 | 2.5245 | | 3.8790 | 2.1450 | 2.1450 | 2.7419 | 2.7419 | 1.0853 | 1.0865 | 1.0865 | 4.6855 | 4.1917 | 4.1917 | C4 | 2.5245 | 1.5167 | 3.8790 | | 2.7419 | 2.7419 | 2.1450 | 2.1450 | 4.6855 | 4.1917 | 4.1917 | 1.0853 | 1.0865 | 1.0865 | H5 | 1.0885 | 2.1356 | 2.1450 | 2.7419 | | 1.7432 | 2.4804 | 3.0317 | 2.4917 | 2.5007 | 3.0515 | 3.7410 | 2.5518 | 3.0935 | H6 | 1.0885 | 2.1356 | 2.1450 | 2.7419 | 1.7432 | | 3.0317 | 2.4804 | 2.4917 | 3.0515 | 2.5007 | 3.7410 | 3.0935 | 2.5518 | H7 | 2.1356 | 1.0885 | 2.7419 | 2.1450 | 2.4804 | 3.0317 | | 1.7432 | 3.7410 | 2.5518 | 3.0935 | 2.4917 | 2.5007 | 3.0515 | H8 | 2.1356 | 1.0885 | 2.7419 | 2.1450 | 3.0317 | 2.4804 | 1.7432 | | 3.7410 | 3.0935 | 2.5518 | 2.4917 | 3.0515 | 2.5007 | H9 | 2.1673 | 3.4729 | 1.0853 | 4.6855 | 2.4917 | 2.4917 | 3.7410 | 3.7410 | | 1.7563 | 1.7563 | 5.5872 | 4.8736 | 4.8736 | H10 | 2.1555 | 2.7805 | 1.0865 | 4.1917 | 2.5007 | 3.0515 | 2.5518 | 3.0935 | 1.7563 | | 1.7540 | 4.8736 | 4.4022 | 4.7387 | H11 | 2.1555 | 2.7805 | 1.0865 | 4.1917 | 3.0515 | 2.5007 | 3.0935 | 2.5518 | 1.7563 | 1.7540 | | 4.8736 | 4.7387 | 4.4022 | H12 | 3.4729 | 2.1673 | 4.6855 | 1.0853 | 3.7410 | 3.7410 | 2.4917 | 2.4917 | 5.5872 | 4.8736 | 4.8736 | | 1.7563 | 1.7563 | H13 | 2.7805 | 2.1555 | 4.1917 | 1.0865 | 2.5518 | 3.0935 | 2.5007 | 3.0515 | 4.8736 | 4.4022 | 4.7387 | 1.7563 | | 1.7540 | H14 | 2.7805 | 2.1555 | 4.1917 | 1.0865 | 3.0935 | 2.5518 | 3.0515 | 2.5007 | 4.8736 | 4.7387 | 4.4022 | 1.7563 | 1.7540 | |
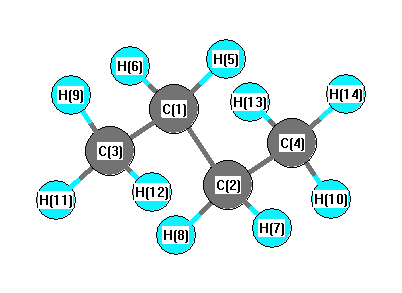 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
112.670 |
|
C1 |
C2 |
H7 |
109.026 |
| C1 |
C2 |
H8 |
109.026 |
|
C1 |
C3 |
H9 |
111.737 |
| C1 |
C3 |
H11 |
110.713 |
|
C1 |
C3 |
H12 |
30.918 |
| C2 |
C1 |
C3 |
112.670 |
|
C2 |
C1 |
H5 |
109.026 |
| C2 |
C1 |
H6 |
109.026 |
|
C2 |
C4 |
H10 |
17.301 |
| C2 |
C4 |
H13 |
110.713 |
|
C2 |
C4 |
H14 |
110.713 |
| C3 |
C1 |
H5 |
109.757 |
|
C3 |
C1 |
H6 |
109.757 |
| C4 |
C2 |
H7 |
109.757 |
|
C4 |
C2 |
H8 |
109.757 |
| H5 |
C1 |
H6 |
106.407 |
|
H7 |
C2 |
H8 |
106.407 |
| H9 |
C3 |
H11 |
107.937 |
|
H9 |
C3 |
H12 |
142.654 |
| H10 |
C4 |
H13 |
93.953 |
|
H10 |
C4 |
H14 |
113.998 |
| H11 |
C3 |
H12 |
93.476 |
|
H13 |
C4 |
H14 |
107.649 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -158.137478 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 133.594648 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3172 |
3012 |
25.88 |
|
|
|
| 2 |
A |
3159 |
2999 |
1.74 |
|
|
|
| 3 |
A |
3115 |
2957 |
24.39 |
|
|
|
| 4 |
A |
3091 |
2934 |
24.90 |
|
|
|
| 5 |
A |
3076 |
2921 |
10.34 |
|
|
|
| 6 |
A |
1540 |
1462 |
1.52 |
|
|
|
| 7 |
A |
1531 |
1453 |
4.70 |
|
|
|
| 8 |
A |
1511 |
1435 |
0.03 |
|
|
|
| 9 |
A |
1424 |
1351 |
4.73 |
|
|
|
| 10 |
A |
1394 |
1323 |
0.21 |
|
|
|
| 11 |
A |
1334 |
1266 |
0.30 |
|
|
|
| 12 |
A |
1210 |
1149 |
0.04 |
|
|
|
| 13 |
A |
1126 |
1069 |
0.14 |
|
|
|
| 14 |
A |
1007 |
956 |
0.13 |
|
|
|
| 15 |
A |
858 |
815 |
0.02 |
|
|
|
| 16 |
A |
813 |
772 |
1.62 |
|
|
|
| 17 |
A |
331 |
314 |
0.02 |
|
|
|
| 18 |
A |
281 |
266 |
0.02 |
|
|
|
| 19 |
A |
110 |
104 |
0.00 |
|
|
|
| 20 |
B |
3165 |
3005 |
39.67 |
|
|
|
| 21 |
B |
3158 |
2999 |
58.06 |
|
|
|
| 22 |
B |
3117 |
2959 |
22.92 |
|
|
|
| 23 |
B |
3089 |
2933 |
21.61 |
|
|
|
| 24 |
B |
3081 |
2925 |
25.18 |
|
|
|
| 25 |
B |
1532 |
1454 |
10.75 |
|
|
|
| 26 |
B |
1526 |
1448 |
9.33 |
|
|
|
| 27 |
B |
1507 |
1431 |
0.57 |
|
|
|
| 28 |
B |
1426 |
1354 |
6.61 |
|
|
|
| 29 |
B |
1384 |
1314 |
0.68 |
|
|
|
| 30 |
B |
1306 |
1240 |
0.30 |
|
|
|
| 31 |
B |
1170 |
1111 |
1.87 |
|
|
|
| 32 |
B |
998 |
947 |
1.44 |
|
|
|
| 33 |
B |
981 |
931 |
2.35 |
|
|
|
| 34 |
B |
758 |
720 |
3.39 |
|
|
|
| 35 |
B |
437 |
415 |
0.34 |
|
|
|
| 36 |
B |
225 |
213 |
0.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29470.2 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 27979.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
