Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -158.401120 |
| Energy at 298.15K | -158.411712 |
| HF Energy | -158.401120 |
| Nuclear repulsion energy | 131.323222 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
124 |
119 |
0.02 |
|
|
|
|
Au |
212 |
203 |
0.00 |
|
|
|
|
Bu |
245 |
235 |
0.01 |
|
|
|
|
Bg |
253 |
242 |
0.00 |
|
|
|
|
Ag |
418 |
400 |
0.00 |
|
|
|
|
Au |
732 |
701 |
3.91 |
|
|
|
|
Bg |
806 |
771 |
0.00 |
|
|
|
|
Ag |
851 |
815 |
0.00 |
|
|
|
|
Au |
959 |
918 |
1.06 |
|
|
|
|
Bu |
977 |
935 |
6.25 |
|
|
|
|
Bu |
1038 |
993 |
0.05 |
|
|
|
|
Ag |
1087 |
1041 |
0.00 |
|
|
|
|
Ag |
1174 |
1123 |
0.00 |
|
|
|
|
Bg |
1208 |
1156 |
0.00 |
|
|
|
|
Au |
1288 |
1232 |
0.18 |
|
|
|
|
Bu |
1318 |
1262 |
2.32 |
|
|
|
|
Bg |
1332 |
1275 |
0.00 |
|
|
|
|
Ag |
1396 |
1336 |
0.00 |
|
|
|
|
Ag |
1408 |
1348 |
0.00 |
|
|
|
|
Bu |
1410 |
1349 |
5.94 |
|
|
|
|
Ag |
1487 |
1423 |
0.00 |
|
|
|
|
Bu |
1491 |
1427 |
1.88 |
|
|
|
|
Bg |
1496 |
1432 |
0.00 |
|
|
|
|
Au |
1498 |
1433 |
14.74 |
|
|
|
|
Ag |
1505 |
1440 |
0.00 |
|
|
|
|
Bu |
1512 |
1447 |
8.78 |
|
|
|
|
Ag |
3035 |
2905 |
0.00 |
|
|
|
|
Bu |
3044 |
2914 |
35.21 |
|
|
|
|
Bu |
3050 |
2919 |
90.99 |
|
|
|
|
Ag |
3050 |
2919 |
0.00 |
|
|
|
|
Bg |
3058 |
2927 |
0.00 |
|
|
|
|
Au |
3082 |
2950 |
19.39 |
|
|
|
|
Bg |
3117 |
2983 |
0.00 |
|
|
|
|
Au |
3121 |
2987 |
107.86 |
|
|
|
|
Ag |
3125 |
2991 |
0.00 |
|
|
|
|
Bu |
3126 |
2992 |
73.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29015.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 27770.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.419 |
0.634 |
0.000 |
| C2 |
0.419 |
-0.634 |
0.000 |
| C3 |
0.419 |
1.899 |
0.000 |
| C4 |
-0.419 |
-1.899 |
0.000 |
| H5 |
-1.076 |
0.627 |
0.872 |
| H6 |
-1.076 |
0.627 |
-0.872 |
| H7 |
1.076 |
-0.627 |
0.872 |
| H8 |
1.076 |
-0.627 |
-0.872 |
| H9 |
-0.202 |
2.793 |
0.000 |
| H10 |
1.062 |
1.943 |
0.879 |
| H11 |
1.062 |
1.943 |
-0.879 |
| H12 |
0.202 |
-2.793 |
0.000 |
| H13 |
-1.062 |
-1.943 |
0.879 |
| H14 |
-1.062 |
-1.943 |
-0.879 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5192 | 1.5177 | 2.5328 | 1.0920 | 1.0920 | 2.1414 | 2.1414 | 2.1700 | 2.1629 | 2.1629 | 3.4823 | 2.7968 | 2.7968 |
C2 | 1.5192 | | 2.5328 | 1.5177 | 2.1414 | 2.1414 | 1.0920 | 1.0920 | 3.4823 | 2.7968 | 2.7968 | 2.1700 | 2.1629 | 2.1629 | C3 | 1.5177 | 2.5328 | | 3.8896 | 2.1477 | 2.1477 | 2.7525 | 2.7525 | 1.0884 | 1.0896 | 1.0896 | 4.6969 | 4.2099 | 4.2099 | C4 | 2.5328 | 1.5177 | 3.8896 | | 2.7525 | 2.7525 | 2.1477 | 2.1477 | 4.6969 | 4.2099 | 4.2099 | 1.0884 | 1.0896 | 1.0896 | H5 | 1.0920 | 2.1414 | 2.1477 | 2.7525 | | 1.7440 | 2.4913 | 3.0411 | 2.4925 | 2.5102 | 3.0602 | 3.7541 | 2.5701 | 3.1096 | H6 | 1.0920 | 2.1414 | 2.1477 | 2.7525 | 1.7440 | | 3.0411 | 2.4913 | 2.4925 | 3.0602 | 2.5102 | 3.7541 | 3.1096 | 2.5701 | H7 | 2.1414 | 1.0920 | 2.7525 | 2.1477 | 2.4913 | 3.0411 | | 1.7440 | 3.7541 | 2.5701 | 3.1096 | 2.4925 | 2.5102 | 3.0602 | H8 | 2.1414 | 1.0920 | 2.7525 | 2.1477 | 3.0411 | 2.4913 | 1.7440 | | 3.7541 | 3.1096 | 2.5701 | 2.4925 | 3.0602 | 2.5102 | H9 | 2.1700 | 3.4823 | 1.0884 | 4.6969 | 2.4925 | 2.4925 | 3.7541 | 3.7541 | | 1.7587 | 1.7587 | 5.6002 | 4.8923 | 4.8923 | H10 | 2.1629 | 2.7968 | 1.0896 | 4.2099 | 2.5102 | 3.0602 | 2.5701 | 3.1096 | 1.7587 | | 1.7570 | 4.8923 | 4.4278 | 4.7637 | H11 | 2.1629 | 2.7968 | 1.0896 | 4.2099 | 3.0602 | 2.5102 | 3.1096 | 2.5701 | 1.7587 | 1.7570 | | 4.8923 | 4.7637 | 4.4278 | H12 | 3.4823 | 2.1700 | 4.6969 | 1.0884 | 3.7541 | 3.7541 | 2.4925 | 2.4925 | 5.6002 | 4.8923 | 4.8923 | | 1.7587 | 1.7587 | H13 | 2.7968 | 2.1629 | 4.2099 | 1.0896 | 2.5701 | 3.1096 | 2.5102 | 3.0602 | 4.8923 | 4.4278 | 4.7637 | 1.7587 | | 1.7570 | H14 | 2.7968 | 2.1629 | 4.2099 | 1.0896 | 3.1096 | 2.5701 | 3.0602 | 2.5102 | 4.8923 | 4.7637 | 4.4278 | 1.7587 | 1.7570 | |
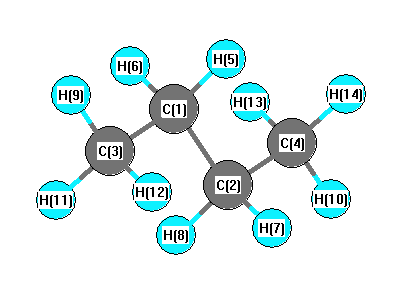 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
113.029 |
|
C1 |
C2 |
H7 |
109.098 |
| C1 |
C2 |
H8 |
109.098 |
|
C1 |
C3 |
H9 |
111.691 |
| C1 |
C3 |
H11 |
111.042 |
|
C1 |
C3 |
H12 |
30.862 |
| C2 |
C1 |
C3 |
113.029 |
|
C2 |
C1 |
H5 |
109.098 |
| C2 |
C1 |
H6 |
109.098 |
|
C2 |
C4 |
H10 |
17.238 |
| C2 |
C4 |
H13 |
111.042 |
|
C2 |
C4 |
H14 |
111.042 |
| C3 |
C1 |
H5 |
109.692 |
|
C3 |
C1 |
H6 |
109.692 |
| C4 |
C2 |
H7 |
109.692 |
|
C4 |
C2 |
H8 |
109.692 |
| H5 |
C1 |
H6 |
105.987 |
|
H7 |
C2 |
H8 |
105.987 |
| H9 |
C3 |
H11 |
107.705 |
|
H9 |
C3 |
H12 |
142.553 |
| H10 |
C4 |
H13 |
94.344 |
|
H10 |
C4 |
H14 |
114.346 |
| H11 |
C3 |
H12 |
93.845 |
|
H13 |
C4 |
H14 |
107.467 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.195 |
|
|
|
| 2 |
C |
-0.195 |
|
|
|
| 3 |
C |
-0.362 |
|
|
|
| 4 |
C |
-0.362 |
|
|
|
| 5 |
H |
0.108 |
|
|
|
| 6 |
H |
0.108 |
|
|
|
| 7 |
H |
0.108 |
|
|
|
| 8 |
H |
0.108 |
|
|
|
| 9 |
H |
0.118 |
|
|
|
| 10 |
H |
0.111 |
|
|
|
| 11 |
H |
0.111 |
|
|
|
| 12 |
H |
0.118 |
|
|
|
| 13 |
H |
0.111 |
|
|
|
| 14 |
H |
0.111 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.622 |
-0.427 |
0.000 |
| y |
-0.427 |
-29.012 |
0.000 |
| z |
0.000 |
0.000 |
-27.849 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.192 |
-0.427 |
0.000 |
| y |
-0.427 |
-0.776 |
0.000 |
| z |
0.000 |
0.000 |
0.968 |
|
| Polar |
|---|
| 3z2-r2 | 1.936 |
|---|
| x2-y2 | 0.389 |
|---|
| xy | -0.427 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.171 |
0.293 |
0.000 |
| y |
0.293 |
8.600 |
0.000 |
| z |
0.000 |
0.000 |
6.780 |
<r2> (average value of r
2) Å
2
| <r2> |
117.729 |
| (<r2>)1/2 |
10.850 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -158.399910 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 133.182451 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3133 |
2998 |
35.22 |
|
|
|
| 2 |
A |
3120 |
2987 |
3.32 |
|
|
|
| 3 |
A |
3076 |
2944 |
27.10 |
|
|
|
| 4 |
A |
3053 |
2922 |
29.25 |
|
|
|
| 5 |
A |
3038 |
2907 |
15.57 |
|
|
|
| 6 |
A |
1507 |
1443 |
0.00 |
|
|
|
| 7 |
A |
1502 |
1438 |
6.36 |
|
|
|
| 8 |
A |
1486 |
1422 |
0.00 |
|
|
|
| 9 |
A |
1408 |
1348 |
4.34 |
|
|
|
| 10 |
A |
1377 |
1318 |
0.36 |
|
|
|
| 11 |
A |
1312 |
1256 |
0.28 |
|
|
|
| 12 |
A |
1195 |
1143 |
0.05 |
|
|
|
| 13 |
A |
1105 |
1057 |
0.13 |
|
|
|
| 14 |
A |
988 |
946 |
0.17 |
|
|
|
| 15 |
A |
844 |
808 |
0.01 |
|
|
|
| 16 |
A |
795 |
761 |
1.90 |
|
|
|
| 17 |
A |
312 |
298 |
0.03 |
|
|
|
| 18 |
A |
240 |
230 |
0.02 |
|
|
|
| 19 |
A |
104 |
100 |
0.00 |
|
|
|
| 20 |
B |
3128 |
2994 |
48.24 |
|
|
|
| 21 |
B |
3121 |
2987 |
80.68 |
|
|
|
| 22 |
B |
3079 |
2947 |
27.45 |
|
|
|
| 23 |
B |
3052 |
2921 |
26.72 |
|
|
|
| 24 |
B |
3040 |
2909 |
29.78 |
|
|
|
| 25 |
B |
1504 |
1440 |
10.13 |
|
|
|
| 26 |
B |
1496 |
1432 |
10.11 |
|
|
|
| 27 |
B |
1482 |
1419 |
0.35 |
|
|
|
| 28 |
B |
1411 |
1351 |
7.10 |
|
|
|
| 29 |
B |
1369 |
1311 |
1.20 |
|
|
|
| 30 |
B |
1288 |
1233 |
0.33 |
|
|
|
| 31 |
B |
1155 |
1105 |
2.22 |
|
|
|
| 32 |
B |
982 |
940 |
1.93 |
|
|
|
| 33 |
B |
964 |
923 |
2.43 |
|
|
|
| 34 |
B |
743 |
711 |
4.11 |
|
|
|
| 35 |
B |
423 |
405 |
0.39 |
|
|
|
| 36 |
B |
207 |
198 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29019.3 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 27774.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
