return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2CH3 (1-Butene)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-156.910974
Energy at 298.15K-156.919003
HF Energy-156.164115
Nuclear repulsion energy118.026190
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3278 3112 11.58      
2 A 3202 3040 2.54      
3 A 3183 3022 19.80      
4 A 3170 3010 24.12      
5 A 3164 3004 23.85      
6 A 3121 2963 7.02      
7 A 3091 2935 25.91      
8 A 3083 2927 18.83      
9 A 1715 1628 7.84      
10 A 1533 1455 5.47      
11 A 1526 1449 6.57      
12 A 1505 1429 4.10      
13 A 1474 1399 2.31      
14 A 1419 1347 2.78      
15 A 1357 1289 1.03      
16 A 1328 1261 0.81      
17 A 1308 1242 0.19      
18 A 1215 1154 0.31      
19 A 1111 1055 3.71      
20 A 1061 1008 2.44      
21 A 1044 991 14.46      
22 A 996 946 2.19      
23 A 960 912 38.36      
24 A 882 837 2.04      
25 A 804 764 2.70      
26 A 670 636 9.94      
27 A 438 415 0.62      
28 A 321 305 0.54      
29 A 246 234 0.04      
30 A 106 101 0.07      

Unscaled Zero Point Vibrational Energy (zpe) 24155.3 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 22933.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.74828 0.14162 0.13866

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.695 -0.260 -0.296
C2 0.539 0.532 0.300
C3 -0.710 -0.277 0.365
C4 -1.832 0.008 -0.289
H5 -0.674 -1.167 0.976
H6 1.462 -0.565 -1.311
H7 2.606 0.328 -0.315
H8 1.886 -1.157 0.286
H9 0.812 0.858 1.302
H10 0.357 1.427 -0.288
H11 -1.902 0.886 -0.910
H12 -2.700 -0.623 -0.217

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.52242.49343.53682.83771.08531.08531.08622.14072.15353.82434.4108
C21.52241.48902.49842.19432.15672.16622.16041.08881.08712.74723.4778
C32.49341.48901.32971.08042.75843.43842.74202.11732.11412.09762.1027
C43.53682.49841.32972.07893.49664.44983.93863.20072.60881.07801.0762
H52.83772.19431.08042.07893.18733.82952.65182.53313.06503.04662.4134
H61.08532.15672.75843.49663.18731.76091.75503.04592.49783.68554.3046
H71.08532.16623.43844.44983.82951.76091.75692.47242.50364.58115.3923
H81.08622.16042.74203.93862.65181.75501.75692.49953.05764.46694.6448
H92.14071.08882.11733.20072.53313.04592.47242.49951.74973.50154.1036
H102.15351.08712.11412.60883.06502.49782.50363.05761.74972.40443.6822
H113.82432.74722.09761.07803.04663.68554.58114.46693.50152.40441.8433
H124.41083.47782.10271.07622.41344.30465.39234.64484.10363.68221.8433

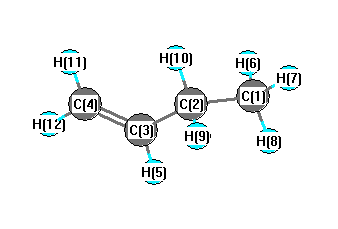
picture of 1-Butene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.783 C1 C2 H9 109.004
C1 C2 H10 110.115 C2 C1 H6 110.475
C2 C1 H7 111.240 C2 C1 H8 110.723
C2 C3 C4 124.744 C2 C3 H5 116.398
C3 C2 H9 109.461 C3 C2 H10 109.310
C3 C4 H11 120.840 C3 C4 H12 121.495
C4 C3 H5 118.845 H6 C1 H7 108.433
H6 C1 H8 107.843 H7 C1 H8 108.013
H9 C2 H10 107.048 H11 C4 H12 117.665
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability