Jump to
S1C2
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
Geometric Data calculated at MP4=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -155.755968 |
| Energy at 298.15K | -155.761328 |
| HF Energy | -154.972083 |
| Nuclear repulsion energy | 105.154458 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3246 |
3125 |
|
|
|
|
| 2 |
A |
3182 |
3064 |
|
|
|
|
| 3 |
A |
3165 |
3047 |
|
|
|
|
| 4 |
A |
1673 |
1611 |
|
|
|
|
| 5 |
A |
1478 |
1423 |
|
|
|
|
| 6 |
A |
1343 |
1293 |
|
|
|
|
| 7 |
A |
1068 |
1028 |
|
|
|
|
| 8 |
A |
1016 |
978 |
|
|
|
|
| 9 |
A |
947 |
912 |
|
|
|
|
| 10 |
A |
894 |
861 |
|
|
|
|
| 11 |
A |
757 |
729 |
|
|
|
|
| 12 |
A |
271 |
261 |
|
|
|
|
| 13 |
A |
170 |
163 |
|
|
|
|
| 14 |
B |
3244 |
3123 |
|
|
|
|
| 15 |
B |
3173 |
3055 |
|
|
|
|
| 16 |
B |
3157 |
3040 |
|
|
|
|
| 17 |
B |
1680 |
1618 |
|
|
|
|
| 18 |
B |
1446 |
1393 |
|
|
|
|
| 19 |
B |
1310 |
1261 |
|
|
|
|
| 20 |
B |
1106 |
1065 |
|
|
|
|
| 21 |
B |
1031 |
993 |
|
|
|
|
| 22 |
B |
948 |
912 |
|
|
|
|
| 23 |
B |
616 |
594 |
|
|
|
|
| 24 |
B |
476 |
458 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18697.3 cm
-1
Scaled (by 0.9629) Zero Point Vibrational Energy (zpe) 18003.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.108 |
0.725 |
0.557 |
| C2 |
-0.108 |
-0.725 |
0.557 |
| C3 |
-0.108 |
1.525 |
-0.494 |
| C4 |
0.108 |
-1.525 |
-0.494 |
| H5 |
0.451 |
1.163 |
1.483 |
| H6 |
-0.451 |
-1.163 |
1.483 |
| H7 |
0.090 |
2.584 |
-0.439 |
| H8 |
-0.499 |
1.134 |
-1.421 |
| H9 |
-0.090 |
-2.584 |
-0.439 |
| H10 |
0.499 |
-1.134 |
-1.421 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.4656 | 1.3384 | 2.4832 | 1.0809 | 2.1761 | 2.1092 | 2.1089 | 3.4609 | 2.7420 |
C2 | 1.4656 | | 2.4832 | 1.3384 | 2.1761 | 1.0809 | 3.4609 | 2.7420 | 2.1092 | 2.1089 | C3 | 1.3384 | 2.4832 | | 3.0583 | 2.0864 | 3.3548 | 1.0782 | 1.0797 | 4.1096 | 2.8808 | C4 | 2.4832 | 1.3384 | 3.0583 | | 3.3548 | 2.0864 | 4.1096 | 2.8808 | 1.0782 | 1.0797 | H5 | 1.0809 | 2.1761 | 2.0864 | 3.3548 | | 2.4949 | 2.4178 | 3.0559 | 4.2460 | 3.7033 | H6 | 2.1761 | 1.0809 | 3.3548 | 2.0864 | 2.4949 | | 4.2460 | 3.7033 | 2.4178 | 3.0559 | H7 | 2.1092 | 3.4609 | 1.0782 | 4.1096 | 2.4178 | 4.2460 | | 1.8474 | 5.1708 | 3.8668 | H8 | 2.1089 | 2.7420 | 1.0797 | 2.8808 | 3.0559 | 3.7033 | 1.8474 | | 3.8668 | 2.4773 | H9 | 3.4609 | 2.1092 | 4.1096 | 1.0782 | 4.2460 | 2.4178 | 5.1708 | 3.8668 | | 1.8474 | H10 | 2.7420 | 2.1089 | 2.8808 | 1.0797 | 3.7033 | 3.0559 | 3.8668 | 2.4773 | 1.8474 | |
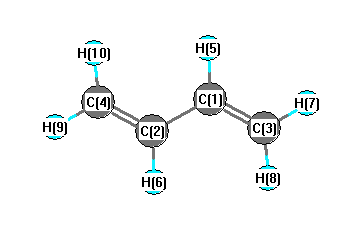 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
124.592 |
|
C1 |
C2 |
H6 |
116.613 |
| C1 |
C3 |
H7 |
121.191 |
|
C1 |
C3 |
H8 |
121.043 |
| C2 |
C1 |
C3 |
124.592 |
|
C2 |
C1 |
H5 |
116.613 |
| C2 |
C4 |
H9 |
121.191 |
|
C2 |
C4 |
H10 |
121.043 |
| C3 |
C1 |
H5 |
118.787 |
|
C4 |
C2 |
H6 |
118.787 |
| H7 |
C3 |
H8 |
117.761 |
|
H9 |
C4 |
H10 |
117.761 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
