return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHCCH2CH3 (1-Butyne)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-155.673586
Energy at 298.15K-155.678633
HF Energy-154.955372
Nuclear repulsion energy103.324512
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3510 3332 58.94      
2 A' 3179 3018 20.12      
3 A' 3102 2945 21.58      
4 A' 3095 2938 12.30      
5 A' 2158 2049 2.03      
6 A' 1533 1455 3.46      
7 A' 1502 1426 3.37      
8 A' 1423 1351 2.19      
9 A' 1364 1295 7.19      
10 A' 1108 1052 2.91      
11 A' 1054 1001 0.42      
12 A' 868 824 0.00      
13 A' 650 617 43.23      
14 A' 518 492 5.47      
15 A' 197 187 1.21      
16 A" 3183 3022 17.11      
17 A" 3127 2969 3.45      
18 A" 1525 1448 6.77      
19 A" 1310 1243 0.00      
20 A" 1122 1065 0.78      
21 A" 796 756 0.87      
22 A" 652 619 43.46      
23 A" 365 346 5.63      
24 A" 237 225 1.02      

Unscaled Zero Point Vibrational Energy (zpe) 18787.3 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 17836.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.91532 0.15326 0.13791

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.646 -1.858 0.000
C2 0.000 -0.835 0.000
C3 0.748 0.411 0.000
C4 -0.167 1.630 0.000
H5 -1.209 -2.752 0.000
H6 1.397 0.435 0.872
H7 1.397 0.435 -0.872
H8 0.416 2.544 0.000
H9 -0.804 1.626 -0.878
H10 -0.804 1.626 0.878

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10
C11.21062.66403.52081.05603.19333.19334.52913.59643.5964
C21.21061.45362.47012.26662.07972.07973.40462.73332.7333
C32.66401.45361.52353.71991.08721.08722.15832.15712.1571
C43.52082.47011.52354.50372.15242.15241.08491.08431.0843
H51.05602.26663.71994.50374.20854.20855.54004.48304.4830
H63.19332.07971.08722.15244.20851.74392.48373.05322.5022
H73.19332.07971.08722.15244.20851.74392.48372.50223.0532
H84.52913.40462.15831.08495.54002.48372.48371.76161.7616
H93.59642.73332.15711.08434.48303.05322.50221.76161.7551
H103.59642.73332.15711.08434.48302.50223.05321.76161.7551

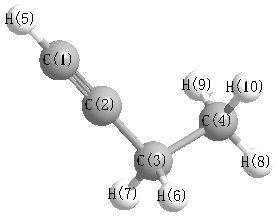
picture of 1-Butyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 178.701 C2 C1 H5 179.943
C2 C3 C4 112.113 C2 C3 H6 109.026
C2 C3 H7 109.026 C3 C4 H8 110.542
C3 C4 H9 110.483 C3 C4 H10 110.483
C4 C3 H6 109.935 C4 C3 H7 109.935
H6 C3 H7 106.647 H8 C4 H9 108.602
H8 C4 H10 108.602 H9 C4 H10 108.056
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability