Jump to
S1C2
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -191.600357 |
| Energy at 298.15K | |
| HF Energy | -190.833310 |
| Nuclear repulsion energy | 103.476775 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3289 |
3123 |
2.59 |
58.72 |
0.41 |
0.58 |
| 2 |
A' |
3251 |
3087 |
0.17 |
95.55 |
0.33 |
0.50 |
| 3 |
A' |
3200 |
3038 |
2.61 |
72.26 |
0.11 |
0.19 |
| 4 |
A' |
2962 |
2812 |
85.39 |
102.14 |
0.25 |
0.40 |
| 5 |
A' |
1767 |
1677 |
144.88 |
61.02 |
0.39 |
0.56 |
| 6 |
A' |
1684 |
1599 |
5.29 |
8.56 |
0.11 |
0.20 |
| 7 |
A' |
1476 |
1402 |
10.27 |
3.10 |
0.71 |
0.83 |
| 8 |
A' |
1412 |
1340 |
4.65 |
18.97 |
0.34 |
0.51 |
| 9 |
A' |
1305 |
1239 |
4.28 |
6.80 |
0.21 |
0.34 |
| 10 |
A' |
1188 |
1128 |
32.25 |
10.47 |
0.60 |
0.75 |
| 11 |
A' |
937 |
890 |
17.13 |
0.86 |
0.05 |
0.10 |
| 12 |
A' |
577 |
548 |
4.02 |
4.92 |
0.29 |
0.45 |
| 13 |
A' |
320 |
304 |
9.85 |
0.45 |
0.39 |
0.56 |
| 14 |
A" |
1057 |
1004 |
11.17 |
0.87 |
0.75 |
0.86 |
| 15 |
A" |
1044 |
991 |
10.99 |
2.20 |
0.75 |
0.86 |
| 16 |
A" |
1011 |
960 |
35.39 |
1.08 |
0.75 |
0.86 |
| 17 |
A" |
632 |
600 |
9.70 |
1.51 |
0.75 |
0.86 |
| 18 |
A" |
172 |
163 |
4.24 |
0.77 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13642.8 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 12952.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.151 |
-0.735 |
0.000 |
| C2 |
0.000 |
0.721 |
0.000 |
| C3 |
1.215 |
1.268 |
0.000 |
| O4 |
-1.217 |
-1.314 |
0.000 |
| H5 |
0.799 |
-1.292 |
0.000 |
| H6 |
-0.899 |
1.311 |
0.000 |
| H7 |
1.364 |
2.333 |
0.000 |
| H8 |
2.092 |
0.640 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4637 | 2.4246 | 1.2131 | 1.1014 | 2.1789 | 3.4218 | 2.6314 |
C2 | 1.4637 | | 1.3326 | 2.3710 | 2.1657 | 1.0758 | 2.1117 | 2.0939 | C3 | 2.4246 | 1.3326 | | 3.5471 | 2.5936 | 2.1148 | 1.0756 | 1.0786 | O4 | 1.2131 | 2.3710 | 3.5471 | | 2.0164 | 2.6445 | 4.4679 | 3.8433 | H5 | 1.1014 | 2.1657 | 2.5936 | 2.0164 | | 3.1084 | 3.6690 | 2.3251 | H6 | 2.1789 | 1.0758 | 2.1148 | 2.6445 | 3.1084 | | 2.4829 | 3.0659 | H7 | 3.4218 | 2.1117 | 1.0756 | 4.4679 | 3.6690 | 2.4829 | | 1.8432 | H8 | 2.6314 | 2.0939 | 1.0786 | 3.8433 | 2.3251 | 3.0659 | 1.8432 | |
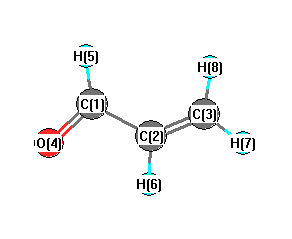 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.163 |
|
C1 |
C2 |
H6 |
117.369 |
| C2 |
C1 |
O4 |
124.426 |
|
C2 |
C1 |
H5 |
114.452 |
| C2 |
C3 |
H7 |
122.172 |
|
C2 |
C3 |
H8 |
120.172 |
| C3 |
C2 |
H6 |
122.468 |
|
O4 |
C1 |
H5 |
121.122 |
| H7 |
C3 |
H8 |
117.656 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -191.597102 |
| Energy at 298.15K | |
| HF Energy | -190.829755 |
| Nuclear repulsion energy | 105.059403 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3299 |
3132 |
0.85 |
47.69 |
0.55 |
0.71 |
| 2 |
A' |
3241 |
3077 |
7.04 |
120.26 |
0.24 |
0.38 |
| 3 |
A' |
3209 |
3046 |
2.51 |
54.18 |
0.13 |
0.24 |
| 4 |
A' |
2996 |
2845 |
123.04 |
162.63 |
0.28 |
0.44 |
| 5 |
A' |
1765 |
1676 |
65.50 |
11.89 |
0.53 |
0.69 |
| 6 |
A' |
1688 |
1602 |
44.53 |
38.15 |
0.18 |
0.31 |
| 7 |
A' |
1460 |
1386 |
34.38 |
6.31 |
0.72 |
0.84 |
| 8 |
A' |
1453 |
1379 |
5.03 |
9.65 |
0.17 |
0.30 |
| 9 |
A' |
1323 |
1256 |
1.58 |
13.18 |
0.33 |
0.50 |
| 10 |
A' |
1080 |
1025 |
2.86 |
3.35 |
0.75 |
0.86 |
| 11 |
A' |
956 |
908 |
56.55 |
3.11 |
0.16 |
0.28 |
| 12 |
A' |
687 |
652 |
10.80 |
1.03 |
0.58 |
0.73 |
| 13 |
A' |
290 |
276 |
6.80 |
2.69 |
0.45 |
0.62 |
| 14 |
A" |
1059 |
1005 |
9.65 |
4.55 |
0.75 |
0.86 |
| 15 |
A" |
1047 |
994 |
26.84 |
0.79 |
0.75 |
0.86 |
| 16 |
A" |
1021 |
970 |
13.51 |
0.62 |
0.75 |
0.86 |
| 17 |
A" |
583 |
553 |
10.18 |
2.52 |
0.75 |
0.86 |
| 18 |
A" |
155 |
147 |
5.80 |
0.26 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13655.9 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 12964.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.887 |
-0.283 |
0.000 |
| C2 |
0.000 |
0.897 |
0.000 |
| C3 |
1.325 |
0.761 |
0.000 |
| O4 |
-0.493 |
-1.431 |
0.000 |
| H5 |
-1.963 |
-0.067 |
0.000 |
| H6 |
-0.463 |
1.869 |
0.000 |
| H7 |
1.981 |
1.613 |
0.000 |
| H8 |
1.755 |
-0.226 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4758 | 2.4454 | 1.2137 | 1.0977 | 2.1927 | 3.4379 | 2.6426 |
C2 | 1.4758 | | 1.3318 | 2.3794 | 2.1869 | 1.0763 | 2.1068 | 2.0841 | C3 | 2.4454 | 1.3318 | | 2.8474 | 3.3905 | 2.1031 | 1.0755 | 1.0772 | O4 | 1.2137 | 2.3794 | 2.8474 | | 2.0052 | 3.2996 | 3.9224 | 2.5504 | H5 | 1.0977 | 2.1869 | 3.3905 | 2.0052 | | 2.4489 | 4.2873 | 3.7217 | H6 | 2.1927 | 1.0763 | 2.1031 | 3.2996 | 2.4489 | | 2.4578 | 3.0513 | H7 | 3.4379 | 2.1068 | 1.0755 | 3.9224 | 4.2873 | 2.4578 | | 1.8531 | H8 | 2.6426 | 2.0841 | 1.0772 | 2.5504 | 3.7217 | 3.0513 | 1.8531 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.061 |
|
C1 |
C2 |
H6 |
117.601 |
| C2 |
C1 |
O4 |
124.142 |
|
C2 |
C1 |
H5 |
115.597 |
| C2 |
C3 |
H7 |
121.765 |
|
C2 |
C3 |
H8 |
119.418 |
| C3 |
C2 |
H6 |
121.338 |
|
O4 |
C1 |
H5 |
120.261 |
| H7 |
C3 |
H8 |
118.816 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
