Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -191.894814 |
| Energy at 298.15K | |
| HF Energy | -191.894814 |
| Nuclear repulsion energy | 103.712299 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3252 |
3113 |
4.74 |
66.52 |
0.50 |
0.67 |
| 2 |
A' |
3202 |
3065 |
1.38 |
103.81 |
0.32 |
0.49 |
| 3 |
A' |
3155 |
3020 |
5.16 |
81.13 |
0.11 |
0.19 |
| 4 |
A' |
2901 |
2776 |
94.94 |
107.25 |
0.27 |
0.43 |
| 5 |
A' |
1821 |
1743 |
246.50 |
63.29 |
0.42 |
0.59 |
| 6 |
A' |
1702 |
1629 |
2.44 |
37.77 |
0.18 |
0.31 |
| 7 |
A' |
1455 |
1392 |
13.13 |
5.80 |
0.61 |
0.76 |
| 8 |
A' |
1391 |
1331 |
6.35 |
19.21 |
0.36 |
0.53 |
| 9 |
A' |
1299 |
1243 |
3.97 |
7.19 |
0.22 |
0.36 |
| 10 |
A' |
1175 |
1125 |
38.04 |
11.93 |
0.59 |
0.74 |
| 11 |
A' |
925 |
885 |
19.22 |
1.17 |
0.06 |
0.10 |
| 12 |
A' |
574 |
550 |
5.23 |
5.32 |
0.28 |
0.44 |
| 13 |
A' |
312 |
299 |
10.13 |
1.02 |
0.38 |
0.55 |
| 14 |
A" |
1044 |
999 |
9.49 |
0.77 |
0.75 |
0.86 |
| 15 |
A" |
1037 |
992 |
3.13 |
3.45 |
0.75 |
0.86 |
| 16 |
A" |
1007 |
964 |
44.31 |
0.98 |
0.75 |
0.86 |
| 17 |
A" |
617 |
591 |
9.92 |
1.52 |
0.75 |
0.86 |
| 18 |
A" |
170 |
162 |
4.26 |
1.06 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13519.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 12939.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.153 |
-0.738 |
0.000 |
| C2 |
0.000 |
0.720 |
0.000 |
| C3 |
1.208 |
1.268 |
0.000 |
| O4 |
-1.209 |
-1.311 |
0.000 |
| H5 |
0.803 |
-1.299 |
0.000 |
| H6 |
-0.908 |
1.307 |
0.000 |
| H7 |
1.358 |
2.337 |
0.000 |
| H8 |
2.094 |
0.646 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4660 | 2.4240 | 1.2023 | 1.1081 | 2.1803 | 3.4262 | 2.6388 |
C2 | 1.4660 | | 1.3262 | 2.3642 | 2.1725 | 1.0812 | 2.1117 | 2.0950 | C3 | 2.4240 | 1.3262 | | 3.5351 | 2.5984 | 2.1157 | 1.0794 | 1.0826 | O4 | 1.2023 | 2.3642 | 3.5351 | | 2.0128 | 2.6360 | 4.4613 | 3.8398 | H5 | 1.1081 | 2.1725 | 2.5984 | 2.0128 | | 3.1174 | 3.6778 | 2.3341 | H6 | 2.1803 | 1.0812 | 2.1157 | 2.6360 | 3.1174 | | 2.4887 | 3.0734 | H7 | 3.4262 | 2.1117 | 1.0794 | 4.4613 | 3.6778 | 2.4887 | | 1.8439 | H8 | 2.6388 | 2.0950 | 1.0826 | 3.8398 | 2.3341 | 3.0734 | 1.8439 | |
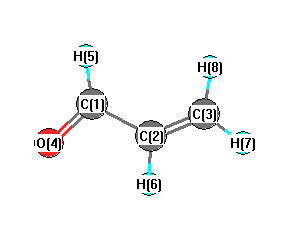 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.401 |
|
C1 |
C2 |
H6 |
116.923 |
| C2 |
C1 |
O4 |
124.458 |
|
C2 |
C1 |
H5 |
114.401 |
| C2 |
C3 |
H7 |
122.426 |
|
C2 |
C3 |
H8 |
120.520 |
| C3 |
C2 |
H6 |
122.676 |
|
O4 |
C1 |
H5 |
121.141 |
| H7 |
C3 |
H8 |
117.054 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.135 |
|
|
|
| 2 |
C |
-0.092 |
|
|
|
| 3 |
C |
-0.280 |
|
|
|
| 4 |
O |
-0.265 |
|
|
|
| 5 |
H |
0.055 |
|
|
|
| 6 |
H |
0.158 |
|
|
|
| 7 |
H |
0.150 |
|
|
|
| 8 |
H |
0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.438 |
2.079 |
0.000 |
3.204 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.323 |
-2.572 |
0.000 |
| y |
-2.572 |
-23.802 |
0.000 |
| z |
0.000 |
0.000 |
-24.709 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.067 |
-2.572 |
0.000 |
| y |
-2.572 |
0.714 |
0.000 |
| z |
0.000 |
0.000 |
-0.646 |
|
| Polar |
|---|
| 3z2-r2 | -1.293 |
|---|
| x2-y2 | -0.520 |
|---|
| xy | -2.572 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.196 |
1.661 |
0.000 |
| y |
1.661 |
6.666 |
0.000 |
| z |
0.000 |
0.000 |
3.193 |
<r2> (average value of r
2) Å
2
| <r2> |
82.516 |
| (<r2>)1/2 |
9.084 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -191.891547 |
| Energy at 298.15K | |
| HF Energy | -191.891547 |
| Nuclear repulsion energy | 105.283453 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3263 |
3123 |
1.83 |
55.94 |
0.60 |
0.75 |
| 2 |
A' |
3192 |
3055 |
10.07 |
124.10 |
0.25 |
0.41 |
| 3 |
A' |
3165 |
3029 |
5.09 |
64.64 |
0.12 |
0.22 |
| 4 |
A' |
2930 |
2804 |
139.96 |
173.27 |
0.30 |
0.46 |
| 5 |
A' |
1821 |
1743 |
118.51 |
14.96 |
0.61 |
0.76 |
| 6 |
A' |
1693 |
1620 |
65.54 |
51.44 |
0.17 |
0.30 |
| 7 |
A' |
1438 |
1376 |
38.37 |
6.86 |
0.72 |
0.84 |
| 8 |
A' |
1427 |
1366 |
8.01 |
8.69 |
0.20 |
0.33 |
| 9 |
A' |
1316 |
1260 |
2.43 |
16.91 |
0.34 |
0.51 |
| 10 |
A' |
1069 |
1023 |
4.36 |
4.15 |
0.71 |
0.83 |
| 11 |
A' |
943 |
902 |
62.30 |
4.37 |
0.16 |
0.27 |
| 12 |
A' |
683 |
654 |
12.19 |
0.76 |
0.72 |
0.84 |
| 13 |
A' |
283 |
270 |
6.97 |
3.30 |
0.42 |
0.59 |
| 14 |
A" |
1047 |
1002 |
1.30 |
6.83 |
0.75 |
0.86 |
| 15 |
A" |
1034 |
990 |
36.18 |
0.14 |
0.75 |
0.86 |
| 16 |
A" |
1022 |
978 |
11.50 |
0.18 |
0.75 |
0.86 |
| 17 |
A" |
563 |
539 |
9.68 |
2.70 |
0.75 |
0.86 |
| 18 |
A" |
156 |
149 |
6.47 |
0.29 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13521.2 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 12941.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.883 |
-0.290 |
0.000 |
| C2 |
0.000 |
0.894 |
0.000 |
| C3 |
1.320 |
0.766 |
0.000 |
| O4 |
-0.490 |
-1.427 |
0.000 |
| H5 |
-1.966 |
-0.074 |
0.000 |
| H6 |
-0.476 |
1.866 |
0.000 |
| H7 |
1.978 |
1.622 |
0.000 |
| H8 |
1.762 |
-0.220 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4771 | 2.4425 | 1.2031 | 1.1051 | 2.1936 | 3.4407 | 2.6460 |
C2 | 1.4771 | | 1.3258 | 2.3727 | 2.1917 | 1.0817 | 2.1076 | 2.0851 | C3 | 2.4425 | 1.3258 | | 2.8438 | 3.3916 | 2.1054 | 1.0794 | 1.0812 | O4 | 1.2031 | 2.3727 | 2.8438 | | 2.0031 | 3.2929 | 3.9227 | 2.5556 | H5 | 1.1051 | 2.1917 | 3.3916 | 2.0031 | | 2.4456 | 4.2933 | 3.7317 | H6 | 2.1936 | 1.0817 | 2.1054 | 3.2929 | 2.4456 | | 2.4662 | 3.0594 | H7 | 3.4407 | 2.1076 | 1.0794 | 3.9227 | 4.2933 | 2.4662 | | 1.8542 | H8 | 2.6460 | 2.0851 | 1.0812 | 2.5556 | 3.7317 | 3.0594 | 1.8542 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.163 |
|
C1 |
C2 |
H6 |
117.187 |
| C2 |
C1 |
O4 |
124.253 |
|
C2 |
C1 |
H5 |
115.402 |
| C2 |
C3 |
H7 |
122.055 |
|
C2 |
C3 |
H8 |
119.713 |
| C3 |
C2 |
H6 |
121.650 |
|
O4 |
C1 |
H5 |
120.345 |
| H7 |
C3 |
H8 |
118.232 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.130 |
|
|
|
| 2 |
C |
-0.136 |
|
|
|
| 3 |
C |
-0.252 |
|
|
|
| 4 |
O |
-0.265 |
|
|
|
| 5 |
H |
0.073 |
|
|
|
| 6 |
H |
0.147 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.210 |
2.651 |
0.000 |
2.659 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.129 |
-0.629 |
0.000 |
| y |
-0.629 |
-25.997 |
0.000 |
| z |
0.000 |
0.000 |
-24.688 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.214 |
-0.629 |
0.000 |
| y |
-0.629 |
-3.588 |
0.000 |
| z |
0.000 |
0.000 |
-1.625 |
|
| Polar |
|---|
| 3z2-r2 | -3.250 |
|---|
| x2-y2 | 5.868 |
|---|
| xy | -0.629 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.388 |
0.517 |
0.000 |
| y |
0.517 |
6.005 |
0.000 |
| z |
0.000 |
0.000 |
3.188 |
<r2> (average value of r
2) Å
2
| <r2> |
74.187 |
| (<r2>)1/2 |
8.613 |
