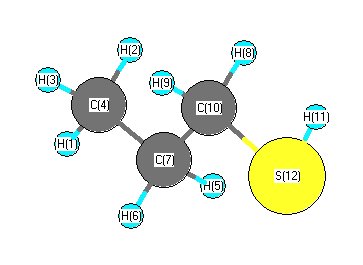
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3132 |
2998 |
29.64 |
|
|
|
| 2 |
A' |
3076 |
2944 |
37.52 |
|
|
|
| 3 |
A' |
3058 |
2926 |
9.66 |
|
|
|
| 4 |
A' |
3053 |
2922 |
20.57 |
|
|
|
| 5 |
A' |
2728 |
2611 |
4.41 |
|
|
|
| 6 |
A' |
1508 |
1443 |
6.35 |
|
|
|
| 7 |
A' |
1493 |
1429 |
0.50 |
|
|
|
| 8 |
A' |
1486 |
1422 |
2.95 |
|
|
|
| 9 |
A' |
1409 |
1349 |
2.54 |
|
|
|
| 10 |
A' |
1372 |
1314 |
1.84 |
|
|
|
| 11 |
A' |
1260 |
1206 |
22.72 |
|
|
|
| 12 |
A' |
1139 |
1090 |
3.85 |
|
|
|
| 13 |
A' |
1066 |
1020 |
0.02 |
|
|
|
| 14 |
A' |
934 |
894 |
2.41 |
|
|
|
| 15 |
A' |
849 |
813 |
2.25 |
|
|
|
| 16 |
A' |
746 |
714 |
1.63 |
|
|
|
| 17 |
A' |
348 |
333 |
0.39 |
|
|
|
| 18 |
A' |
209 |
200 |
1.84 |
|
|
|
| 19 |
A" |
3134 |
3000 |
44.82 |
|
|
|
| 20 |
A" |
3121 |
2988 |
9.06 |
|
|
|
| 21 |
A" |
3089 |
2956 |
1.16 |
|
|
|
| 22 |
A" |
1496 |
1432 |
8.19 |
|
|
|
| 23 |
A" |
1323 |
1266 |
0.04 |
|
|
|
| 24 |
A" |
1242 |
1189 |
0.52 |
|
|
|
| 25 |
A" |
1062 |
1017 |
2.51 |
|
|
|
| 26 |
A" |
865 |
828 |
0.00 |
|
|
|
| 27 |
A" |
748 |
716 |
3.42 |
|
|
|
| 28 |
A" |
225 |
216 |
0.08 |
|
|
|
| 29 |
A" |
124 |
119 |
2.62 |
|
|
|
| 30 |
A" |
67 |
64 |
14.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22680.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 21707.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.123 |
|
|
|
| 2 |
H |
0.114 |
|
|
|
| 3 |
H |
0.114 |
|
|
|
| 4 |
C |
-0.362 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
H |
0.129 |
|
|
|
| 7 |
C |
-0.217 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
| 9 |
H |
0.131 |
|
|
|
| 10 |
C |
-0.205 |
|
|
|
| 11 |
H |
0.109 |
|
|
|
| 12 |
S |
-0.195 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.311 |
0.985 |
0.000 |
1.639 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.045 |
-2.506 |
0.000 |
| y |
-2.506 |
-32.568 |
0.000 |
| z |
0.000 |
0.000 |
-35.162 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.180 |
-2.506 |
0.000 |
| y |
-2.506 |
2.536 |
0.000 |
| z |
0.000 |
0.000 |
-1.356 |
|
| Polar |
|---|
| 3z2-r2 | -2.712 |
|---|
| x2-y2 | -2.477 |
|---|
| xy | -2.506 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.523 |
-0.283 |
0.000 |
| y |
-0.283 |
7.648 |
0.000 |
| z |
0.000 |
0.000 |
6.666 |
<r2> (average value of r
2) Å
2
| <r2> |
155.902 |
| (<r2>)1/2 |
12.486 |