Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Geometric Data calculated at MP2=FULL/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -997.957001 |
| Energy at 298.15K | -997.961914 |
| HF Energy | -997.132220 |
| Nuclear repulsion energy | 204.294013 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3182 |
3021 |
0.60 |
|
|
|
| 2 |
A |
3138 |
2980 |
20.48 |
|
|
|
| 3 |
A |
1496 |
1420 |
0.35 |
|
|
|
| 4 |
A |
1360 |
1291 |
15.86 |
|
|
|
| 5 |
A |
1252 |
1189 |
0.91 |
|
|
|
| 6 |
A |
1082 |
1027 |
1.44 |
|
|
|
| 7 |
A |
982 |
933 |
9.42 |
|
|
|
| 8 |
A |
705 |
670 |
14.28 |
|
|
|
| 9 |
A |
269 |
255 |
0.78 |
|
|
|
| 10 |
A |
125 |
119 |
0.69 |
|
|
|
| 11 |
B |
3195 |
3033 |
2.92 |
|
|
|
| 12 |
B |
3132 |
2974 |
3.14 |
|
|
|
| 13 |
B |
1493 |
1418 |
12.07 |
|
|
|
| 14 |
B |
1336 |
1269 |
30.45 |
|
|
|
| 15 |
B |
1187 |
1127 |
0.86 |
|
|
|
| 16 |
B |
916 |
870 |
16.34 |
|
|
|
| 17 |
B |
729 |
692 |
17.50 |
|
|
|
| 18 |
B |
419 |
398 |
6.45 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12999.5 cm
-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 12341.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.293 |
0.691 |
0.895 |
| C2 |
-0.293 |
-0.691 |
0.895 |
| Cl3 |
-0.293 |
1.659 |
-0.471 |
| Cl4 |
0.293 |
-1.659 |
-0.471 |
| H5 |
0.000 |
1.207 |
1.802 |
| H6 |
1.373 |
0.653 |
0.832 |
| H7 |
-0.000 |
-1.207 |
1.802 |
| H8 |
-1.373 |
-0.653 |
0.832 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5016 | 1.7743 | 2.7185 | 1.0835 | 1.0818 | 2.1238 | 2.1417 |
C2 | 1.5016 | | 2.7185 | 1.7743 | 2.1238 | 2.1417 | 1.0835 | 1.0818 | Cl3 | 1.7743 | 2.7185 | | 3.3694 | 2.3363 | 2.3415 | 3.6696 | 2.8650 | Cl4 | 2.7185 | 1.7743 | 3.3694 | | 3.6696 | 2.8650 | 2.3363 | 2.3415 | H5 | 1.0835 | 2.1238 | 2.3363 | 3.6696 | | 1.7698 | 2.4135 | 2.5073 | H6 | 1.0818 | 2.1417 | 2.3415 | 2.8650 | 1.7698 | | 2.5073 | 3.0403 | H7 | 2.1238 | 1.0835 | 3.6696 | 2.3363 | 2.4135 | 2.5073 | | 1.7698 | H8 | 2.1417 | 1.0818 | 2.8650 | 2.3415 | 2.5073 | 3.0403 | 1.7698 | |
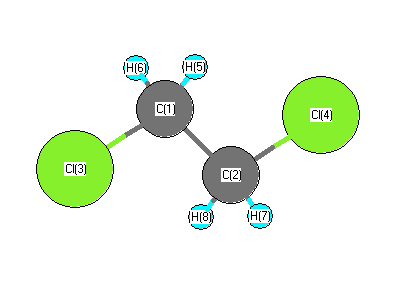 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
111.903 |
|
C1 |
C2 |
H7 |
109.411 |
| C1 |
C2 |
H8 |
110.949 |
|
C2 |
C1 |
Cl3 |
111.903 |
| C2 |
C1 |
H5 |
109.411 |
|
C2 |
C1 |
H6 |
110.949 |
| Cl3 |
C1 |
H5 |
107.189 |
|
Cl3 |
C1 |
H6 |
107.656 |
| Cl4 |
C2 |
H7 |
107.189 |
|
Cl4 |
C2 |
H8 |
107.656 |
| H5 |
C1 |
H6 |
109.637 |
|
H7 |
C2 |
H8 |
109.637 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
