Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -999.156914 |
| Energy at 298.15K | -999.161599 |
| Nuclear repulsion energy | 195.515850 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3121 |
2987 |
0.00 |
|
|
|
| 2 |
Ag |
1491 |
1427 |
0.00 |
|
|
|
| 3 |
Ag |
1329 |
1272 |
0.00 |
|
|
|
| 4 |
Ag |
1086 |
1039 |
0.00 |
|
|
|
| 5 |
Ag |
779 |
746 |
0.00 |
|
|
|
| 6 |
Ag |
300 |
287 |
0.00 |
|
|
|
| 7 |
Au |
3200 |
3063 |
2.87 |
|
|
|
| 8 |
Au |
1140 |
1091 |
1.47 |
|
|
|
| 9 |
Au |
776 |
743 |
2.62 |
|
|
|
| 10 |
Au |
119 |
114 |
6.79 |
|
|
|
| 11 |
Bg |
3176 |
3040 |
0.00 |
|
|
|
| 12 |
Bg |
1295 |
1239 |
0.00 |
|
|
|
| 13 |
Bg |
1008 |
965 |
0.00 |
|
|
|
| 14 |
Bu |
3129 |
2995 |
14.77 |
|
|
|
| 15 |
Bu |
1493 |
1429 |
7.70 |
|
|
|
| 16 |
Bu |
1248 |
1195 |
28.73 |
|
|
|
| 17 |
Bu |
742 |
711 |
95.67 |
|
|
|
| 18 |
Bu |
209 |
200 |
9.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12821.0 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 12271.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.473 |
0.585 |
0.000 |
| C2 |
-0.473 |
-0.585 |
0.000 |
| Cl3 |
-0.473 |
2.100 |
0.000 |
| Cl4 |
0.473 |
-2.100 |
0.000 |
| H5 |
1.097 |
0.590 |
0.886 |
| H6 |
1.097 |
0.590 |
-0.886 |
| H7 |
-1.097 |
-0.590 |
0.886 |
| H8 |
-1.097 |
-0.590 |
-0.886 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5038 | 1.7864 | 2.6851 | 1.0838 | 1.0838 | 2.1519 | 2.1519 |
C2 | 1.5038 | | 2.6851 | 1.7864 | 2.1519 | 2.1519 | 1.0838 | 1.0838 | Cl3 | 1.7864 | 2.6851 | | 4.3058 | 2.3515 | 2.3515 | 2.9009 | 2.9009 | Cl4 | 2.6851 | 1.7864 | 4.3058 | | 2.9009 | 2.9009 | 2.3515 | 2.3515 | H5 | 1.0838 | 2.1519 | 2.3515 | 2.9009 | | 1.7711 | 2.4925 | 3.0577 | H6 | 1.0838 | 2.1519 | 2.3515 | 2.9009 | 1.7711 | | 3.0577 | 2.4925 | H7 | 2.1519 | 1.0838 | 2.9009 | 2.3515 | 2.4925 | 3.0577 | | 1.7711 | H8 | 2.1519 | 1.0838 | 2.9009 | 2.3515 | 3.0577 | 2.4925 | 1.7711 | |
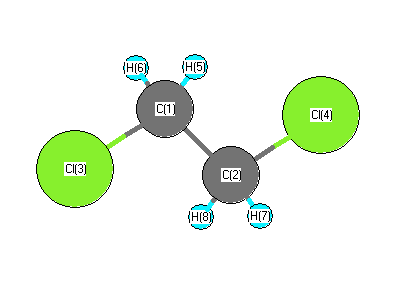 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.087 |
|
C1 |
C2 |
H7 |
111.505 |
| C1 |
C2 |
H8 |
111.505 |
|
C2 |
C1 |
Cl3 |
109.087 |
| C2 |
C1 |
H5 |
111.505 |
|
C2 |
C1 |
H6 |
111.505 |
| Cl3 |
C1 |
H5 |
107.493 |
|
Cl3 |
C1 |
H6 |
107.493 |
| Cl4 |
C2 |
H7 |
107.493 |
|
Cl4 |
C2 |
H8 |
107.493 |
| H5 |
C1 |
H6 |
109.592 |
|
H7 |
C2 |
H8 |
109.592 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.160 |
|
|
|
| 2 |
C |
-0.160 |
|
|
|
| 3 |
Cl |
-0.156 |
|
|
|
| 4 |
Cl |
-0.156 |
|
|
|
| 5 |
H |
0.158 |
|
|
|
| 6 |
H |
0.158 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
| 8 |
H |
0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.063 |
1.966 |
0.000 |
| y |
1.966 |
-44.530 |
0.000 |
| z |
0.000 |
0.000 |
-37.305 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.855 |
1.966 |
0.000 |
| y |
1.966 |
-7.347 |
0.000 |
| z |
0.000 |
0.000 |
3.492 |
|
| Polar |
|---|
| 3z2-r2 | 6.983 |
|---|
| x2-y2 | 7.468 |
|---|
| xy | 1.966 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.319 |
-1.347 |
0.000 |
| y |
-1.347 |
9.770 |
0.000 |
| z |
0.000 |
0.000 |
5.296 |
<r2> (average value of r
2) Å
2
| <r2> |
198.480 |
| (<r2>)1/2 |
14.088 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -999.154604 |
| Energy at 298.15K | -999.159413 |
| HF Energy | -999.154604 |
| Nuclear repulsion energy | 203.343897 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3161 |
3025 |
0.69 |
|
|
|
| 2 |
A |
3107 |
2973 |
22.46 |
|
|
|
| 3 |
A |
1469 |
1406 |
0.77 |
|
|
|
| 4 |
A |
1338 |
1281 |
15.84 |
|
|
|
| 5 |
A |
1229 |
1177 |
0.57 |
|
|
|
| 6 |
A |
1060 |
1014 |
1.75 |
|
|
|
| 7 |
A |
959 |
918 |
11.54 |
|
|
|
| 8 |
A |
681 |
652 |
17.59 |
|
|
|
| 9 |
A |
255 |
244 |
0.93 |
|
|
|
| 10 |
A |
107 |
102 |
0.81 |
|
|
|
| 11 |
B |
3175 |
3039 |
4.28 |
|
|
|
| 12 |
B |
3099 |
2966 |
3.21 |
|
|
|
| 13 |
B |
1467 |
1405 |
12.74 |
|
|
|
| 14 |
B |
1313 |
1256 |
33.57 |
|
|
|
| 15 |
B |
1163 |
1113 |
0.60 |
|
|
|
| 16 |
B |
898 |
859 |
18.91 |
|
|
|
| 17 |
B |
702 |
672 |
24.18 |
|
|
|
| 18 |
B |
407 |
390 |
7.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12793.8 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 12244.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.293 |
0.691 |
0.893 |
| C2 |
-0.293 |
-0.691 |
0.893 |
| Cl3 |
-0.293 |
1.675 |
-0.470 |
| Cl4 |
0.293 |
-1.675 |
-0.470 |
| H5 |
0.002 |
1.208 |
1.803 |
| H6 |
1.375 |
0.661 |
0.826 |
| H7 |
-0.002 |
-1.208 |
1.803 |
| H8 |
-1.375 |
-0.661 |
0.826 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5019 | 1.7805 | 2.7311 | 1.0865 | 1.0843 | 2.1268 | 2.1485 |
C2 | 1.5019 | | 2.7311 | 1.7805 | 2.1268 | 2.1485 | 1.0865 | 1.0843 | Cl3 | 1.7805 | 2.7311 | | 3.4019 | 2.3397 | 2.3434 | 3.6833 | 2.8819 | Cl4 | 2.7311 | 1.7805 | 3.4019 | | 3.6833 | 2.8819 | 2.3397 | 2.3434 | H5 | 1.0865 | 2.1268 | 2.3397 | 3.6833 | | 1.7722 | 2.4158 | 2.5189 | H6 | 1.0843 | 2.1485 | 2.3434 | 2.8819 | 1.7722 | | 2.5189 | 3.0511 | H7 | 2.1268 | 1.0865 | 3.6833 | 2.3397 | 2.4158 | 2.5189 | | 1.7722 | H8 | 2.1485 | 1.0843 | 2.8819 | 2.3434 | 2.5189 | 3.0511 | 1.7722 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.342 |
|
C1 |
C2 |
H7 |
109.455 |
| C1 |
C2 |
H8 |
111.325 |
|
C2 |
C1 |
Cl3 |
112.342 |
| C2 |
C1 |
H5 |
109.455 |
|
C2 |
C1 |
H6 |
111.325 |
| Cl3 |
C1 |
H5 |
106.876 |
|
Cl3 |
C1 |
H6 |
107.262 |
| Cl4 |
C2 |
H7 |
106.876 |
|
Cl4 |
C2 |
H8 |
107.262 |
| H5 |
C1 |
H6 |
109.441 |
|
H7 |
C2 |
H8 |
109.441 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.164 |
|
|
|
| 2 |
C |
-0.164 |
|
|
|
| 3 |
Cl |
-0.140 |
|
|
|
| 4 |
Cl |
-0.140 |
|
|
|
| 5 |
H |
0.148 |
|
|
|
| 6 |
H |
0.156 |
|
|
|
| 7 |
H |
0.148 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.735 |
2.735 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.225 |
1.155 |
0.000 |
| y |
1.155 |
-41.877 |
0.000 |
| z |
0.000 |
0.000 |
-35.105 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.266 |
1.155 |
0.000 |
| y |
1.155 |
-5.712 |
0.000 |
| z |
0.000 |
0.000 |
4.445 |
|
| Polar |
|---|
| 3z2-r2 | 8.891 |
|---|
| x2-y2 | 4.652 |
|---|
| xy | 1.155 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.692 |
-0.492 |
0.000 |
| y |
-0.492 |
7.872 |
0.000 |
| z |
0.000 |
0.000 |
7.254 |
<r2> (average value of r
2) Å
2
| <r2> |
161.431 |
| (<r2>)1/2 |
12.706 |
