return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2CH2CH3 (1-Propanamine)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-174.167643
Energy at 298.15K-174.178129
HF Energy-173.352110
Nuclear repulsion energy132.082408
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3540 3361 0.80      
2 A' 3168 3008 24.88      
3 A' 3090 2934 65.94      
4 A' 3087 2931 15.92      
5 A' 3077 2921 3.30      
6 A' 1665 1581 21.71      
7 A' 1533 1456 6.14      
8 A' 1519 1442 0.56      
9 A' 1505 1428 0.04      
10 A' 1423 1351 3.55      
11 A' 1409 1338 5.71      
12 A' 1334 1266 5.05      
13 A' 1162 1103 3.68      
14 A' 1129 1072 14.56      
15 A' 1069 1015 0.30      
16 A' 933 886 87.35      
17 A' 858 815 106.87      
18 A' 454 431 2.93      
19 A' 264 251 3.85      
20 A" 3632 3448 1.38      
21 A" 3158 2998 41.54      
22 A" 3127 2969 31.15      
23 A" 3100 2944 0.29      
24 A" 1528 1451 7.55      
25 A" 1401 1330 0.33      
26 A" 1340 1272 0.67      
27 A" 1256 1192 0.01      
28 A" 1045 992 0.05      
29 A" 869 825 1.16      
30 A" 759 720 1.94      
31 A" 291 276 35.41      
32 A" 240 228 8.90      
33 A" 138 131 0.22      

Unscaled Zero Point Vibrational Energy (zpe) 27050.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 25681.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.83862 0.12504 0.11782

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.424 1.269 0.000
C2 0.000 0.747 0.000
C3 -0.055 -0.770 0.000
N4 -1.384 -1.363 0.000
H5 1.455 2.354 0.000
H6 1.962 0.920 0.877
H7 1.962 0.920 -0.877
H8 -0.533 1.119 0.873
H9 -0.533 1.119 -0.873
H10 0.480 -1.146 -0.870
H11 0.480 -1.146 0.870
H12 -1.899 -1.039 -0.807
H13 -1.899 -1.039 0.807

Atom - Atom Distances (Å)
  C1 C2 C3 N4 H5 H6 H7 H8 H9 H10 H11 H12 H13
C11.51672.51923.84881.08481.08651.08652.14862.14862.73502.73504.12584.1258
C21.51671.51832.52362.16712.15622.15621.08851.08852.13762.13762.72922.7292
C32.51921.51831.45513.46932.77392.77392.13512.13511.08781.08782.03062.0306
N43.84882.52361.45514.67664.14464.14462.76502.76502.06782.06781.01051.0105
H51.08482.16713.46934.67661.75551.75552.49792.49793.73523.73524.83834.8383
H61.08652.15622.77394.14461.75551.75362.50363.05453.08472.54274.64564.3305
H71.08652.15622.77394.14461.75551.75363.05452.50362.54273.08474.33054.6456
H82.14861.08852.13512.76502.49792.50363.05451.74623.03162.48063.05662.5546
H92.14861.08852.13512.76502.49793.05452.50361.74622.48063.03162.55463.0566
H102.73502.13761.08782.06783.73523.08472.54273.03162.48061.73952.38182.9119
H112.73502.13761.08782.06783.73522.54273.08472.48063.03161.73952.91192.3818
H124.12582.72922.03061.01054.83834.64564.33053.05662.55462.38182.91191.6131
H134.12582.72922.03061.01054.83834.33054.64562.55463.05662.91192.38181.6131

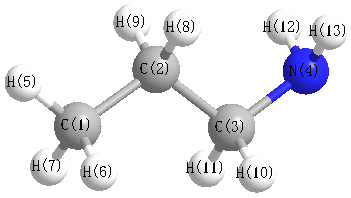
picture of 1-Propanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 112.210 C1 C2 H8 110.040
C1 C2 H9 110.040 C2 C1 H5 111.747
C2 C1 H6 110.763 C2 C1 H7 110.763
C2 C3 N4 116.133 C2 C3 H10 109.099
C2 C3 H11 109.099 C3 C2 H8 108.863
C3 C2 H9 108.863 C3 N4 H12 109.572
C3 N4 H13 109.572 N4 C3 H10 107.941
N4 C3 H11 107.941 H5 C1 H6 107.902
H5 C1 H7 107.902 H6 C1 H7 107.605
H8 C2 H9 106.657 H10 C3 H11 106.177
H12 N4 H13 105.917
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability