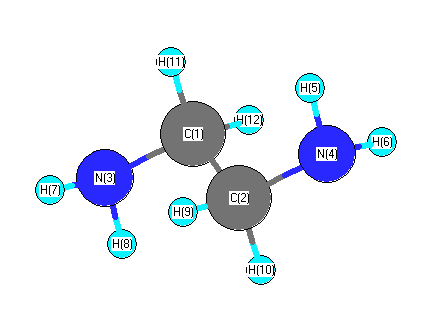
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3528 |
3384 |
0.00 |
|
|
|
| 2 |
Ag |
3027 |
2904 |
0.00 |
|
|
|
| 3 |
Ag |
1666 |
1598 |
0.00 |
|
|
|
| 4 |
Ag |
1482 |
1422 |
0.00 |
|
|
|
| 5 |
Ag |
1401 |
1344 |
0.00 |
|
|
|
| 6 |
Ag |
1129 |
1083 |
0.00 |
|
|
|
| 7 |
Ag |
1071 |
1027 |
0.00 |
|
|
|
| 8 |
Ag |
830 |
796 |
0.00 |
|
|
|
| 9 |
Ag |
459 |
441 |
0.00 |
|
|
|
| 10 |
Au |
3608 |
3461 |
0.70 |
|
|
|
| 11 |
Au |
3084 |
2959 |
66.77 |
|
|
|
| 12 |
Au |
1398 |
1341 |
1.19 |
|
|
|
| 13 |
Au |
1076 |
1032 |
0.18 |
|
|
|
| 14 |
Au |
757 |
726 |
0.96 |
|
|
|
| 15 |
Au |
247 |
237 |
93.58 |
|
|
|
| 16 |
Au |
146 |
140 |
0.00 |
|
|
|
| 17 |
Bg |
3609 |
3462 |
0.00 |
|
|
|
| 18 |
Bg |
3055 |
2930 |
0.00 |
|
|
|
| 19 |
Bg |
1376 |
1320 |
0.00 |
|
|
|
| 20 |
Bg |
1303 |
1250 |
0.00 |
|
|
|
| 21 |
Bg |
943 |
905 |
0.00 |
|
|
|
| 22 |
Bg |
281 |
270 |
0.00 |
|
|
|
| 23 |
Bu |
3527 |
3383 |
1.49 |
|
|
|
| 24 |
Bu |
3037 |
2913 |
89.26 |
|
|
|
| 25 |
Bu |
1665 |
1597 |
45.61 |
|
|
|
| 26 |
Bu |
1500 |
1439 |
1.28 |
|
|
|
| 27 |
Bu |
1331 |
1276 |
17.99 |
|
|
|
| 28 |
Bu |
1104 |
1059 |
20.51 |
|
|
|
| 29 |
Bu |
863 |
828 |
386.52 |
|
|
|
| 30 |
Bu |
267 |
256 |
18.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24385.6 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 23390.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.079 |
|
|
|
| 2 |
C |
-0.079 |
|
|
|
| 3 |
N |
-0.302 |
|
|
|
| 4 |
N |
-0.302 |
|
|
|
| 5 |
H |
0.118 |
|
|
|
| 6 |
H |
0.118 |
|
|
|
| 7 |
H |
0.118 |
|
|
|
| 8 |
H |
0.118 |
|
|
|
| 9 |
H |
0.072 |
|
|
|
| 10 |
H |
0.072 |
|
|
|
| 11 |
H |
0.072 |
|
|
|
| 12 |
H |
0.072 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.186 |
-4.572 |
0.000 |
| y |
-4.572 |
-34.956 |
0.000 |
| z |
0.000 |
0.000 |
-23.293 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.938 |
-4.572 |
0.000 |
| y |
-4.572 |
-11.216 |
0.000 |
| z |
0.000 |
0.000 |
6.278 |
|
| Polar |
|---|
| 3z2-r2 | 12.556 |
|---|
| x2-y2 | 10.770 |
|---|
| xy | -4.572 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.199 |
-0.510 |
0.000 |
| y |
-0.510 |
6.826 |
0.000 |
| z |
0.000 |
0.000 |
5.829 |
<r2> (average value of r
2) Å
2
| <r2> |
105.906 |
| (<r2>)1/2 |
10.291 |