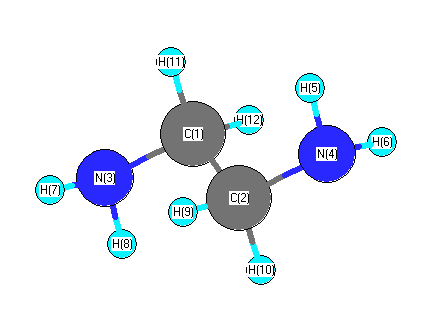
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3525 |
3374 |
0.00 |
|
|
|
| 2 |
Ag |
3033 |
2903 |
0.00 |
|
|
|
| 3 |
Ag |
1663 |
1591 |
0.00 |
|
|
|
| 4 |
Ag |
1482 |
1418 |
0.00 |
|
|
|
| 5 |
Ag |
1396 |
1336 |
0.00 |
|
|
|
| 6 |
Ag |
1129 |
1080 |
0.00 |
|
|
|
| 7 |
Ag |
1073 |
1027 |
0.00 |
|
|
|
| 8 |
Ag |
828 |
792 |
0.00 |
|
|
|
| 9 |
Ag |
455 |
436 |
0.00 |
|
|
|
| 10 |
Au |
3606 |
3451 |
0.67 |
|
|
|
| 11 |
Au |
3090 |
2958 |
68.96 |
|
|
|
| 12 |
Au |
1393 |
1333 |
1.32 |
|
|
|
| 13 |
Au |
1071 |
1025 |
0.19 |
|
|
|
| 14 |
Au |
754 |
721 |
0.80 |
|
|
|
| 15 |
Au |
246 |
235 |
93.65 |
|
|
|
| 16 |
Au |
144 |
138 |
0.04 |
|
|
|
| 17 |
Bg |
3607 |
3452 |
0.00 |
|
|
|
| 18 |
Bg |
3061 |
2930 |
0.00 |
|
|
|
| 19 |
Bg |
1372 |
1313 |
0.00 |
|
|
|
| 20 |
Bg |
1299 |
1243 |
0.00 |
|
|
|
| 21 |
Bg |
938 |
898 |
0.00 |
|
|
|
| 22 |
Bg |
280 |
268 |
0.00 |
|
|
|
| 23 |
Bu |
3524 |
3373 |
1.47 |
|
|
|
| 24 |
Bu |
3042 |
2912 |
91.41 |
|
|
|
| 25 |
Bu |
1662 |
1590 |
45.10 |
|
|
|
| 26 |
Bu |
1500 |
1436 |
0.99 |
|
|
|
| 27 |
Bu |
1324 |
1268 |
19.48 |
|
|
|
| 28 |
Bu |
1105 |
1058 |
21.17 |
|
|
|
| 29 |
Bu |
860 |
823 |
384.06 |
|
|
|
| 30 |
Bu |
258 |
247 |
18.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24358.7 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 23313.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.150 |
|
|
|
| 2 |
C |
-0.150 |
|
|
|
| 3 |
N |
-0.346 |
|
|
|
| 4 |
N |
-0.346 |
|
|
|
| 5 |
H |
0.144 |
|
|
|
| 6 |
H |
0.144 |
|
|
|
| 7 |
H |
0.144 |
|
|
|
| 8 |
H |
0.144 |
|
|
|
| 9 |
H |
0.104 |
|
|
|
| 10 |
H |
0.104 |
|
|
|
| 11 |
H |
0.104 |
|
|
|
| 12 |
H |
0.104 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.224 |
-4.511 |
0.000 |
| y |
-4.511 |
-34.955 |
0.000 |
| z |
0.000 |
0.000 |
-23.342 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.924 |
-4.511 |
0.000 |
| y |
-4.511 |
-11.172 |
0.000 |
| z |
0.000 |
0.000 |
6.248 |
|
| Polar |
|---|
| 3z2-r2 | 12.496 |
|---|
| x2-y2 | 10.731 |
|---|
| xy | -4.511 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.214 |
-0.517 |
0.000 |
| y |
-0.517 |
6.812 |
0.000 |
| z |
0.000 |
0.000 |
5.839 |
<r2> (average value of r
2) Å
2
| <r2> |
105.584 |
| (<r2>)1/2 |
10.275 |