Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
Geometric Data calculated at mPW1PW91/cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -207.978209 |
| Energy at 298.15K | -207.981568 |
| HF Energy | -207.978209 |
| Nuclear repulsion energy | 102.683478 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3876 |
3717 |
50.84 |
58.56 |
0.23 |
0.38 |
| 2 |
A |
3118 |
2991 |
2.77 |
77.97 |
0.37 |
0.54 |
| 3 |
A |
3037 |
2913 |
21.46 |
117.12 |
0.12 |
0.22 |
| 4 |
A |
2383 |
2286 |
0.64 |
66.58 |
0.20 |
0.33 |
| 5 |
A |
1487 |
1426 |
4.81 |
9.19 |
0.61 |
0.76 |
| 6 |
A |
1418 |
1361 |
42.32 |
3.93 |
0.75 |
0.85 |
| 7 |
A |
1377 |
1321 |
2.43 |
3.30 |
0.72 |
0.84 |
| 8 |
A |
1231 |
1181 |
15.09 |
2.74 |
0.46 |
0.63 |
| 9 |
A |
1109 |
1063 |
104.73 |
5.09 |
0.31 |
0.48 |
| 10 |
A |
991 |
950 |
21.61 |
0.62 |
0.33 |
0.50 |
| 11 |
A |
910 |
873 |
13.78 |
1.96 |
0.08 |
0.15 |
| 12 |
A |
593 |
569 |
1.31 |
1.84 |
0.19 |
0.33 |
| 13 |
A |
383 |
367 |
29.61 |
1.01 |
0.73 |
0.85 |
| 14 |
A |
296 |
283 |
107.67 |
2.62 |
0.75 |
0.85 |
| 15 |
A |
215 |
206 |
12.61 |
3.34 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11210.8 cm
-1
Scaled (by 0.9592) Zero Point Vibrational Energy (zpe) 10753.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.566 |
0.584 |
0.038 |
| C2 |
0.822 |
0.111 |
-0.006 |
| O3 |
-1.504 |
-0.447 |
-0.109 |
| H4 |
-0.701 |
1.149 |
0.967 |
| H5 |
-0.722 |
1.276 |
-0.789 |
| H6 |
-1.395 |
-1.073 |
0.609 |
| N7 |
1.902 |
-0.279 |
-0.015 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4672 | 1.4013 | 1.0952 | 1.0897 | 1.9388 | 2.6153 |
C2 | 1.4672 | | 2.3938 | 2.0840 | 2.0864 | 2.5876 | 1.1484 | O3 | 1.4013 | 2.3938 | | 2.0853 | 2.0108 | 0.9590 | 3.4109 | H4 | 1.0952 | 2.0840 | 2.0853 | | 1.7609 | 2.3548 | 3.1270 | H5 | 1.0897 | 2.0864 | 2.0108 | 1.7609 | | 2.8155 | 3.1466 | H6 | 1.9388 | 2.5876 | 0.9590 | 2.3548 | 2.8155 | | 3.4482 | N7 | 2.6153 | 1.1484 | 3.4109 | 3.1270 | 3.1466 | 3.4482 | |
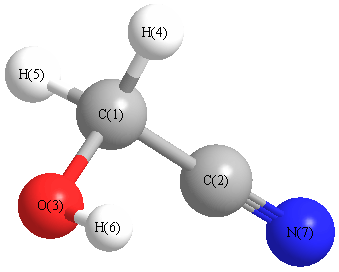 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.337 |
|
C1 |
O3 |
H6 |
109.015 |
| C2 |
C1 |
O3 |
113.105 |
|
C2 |
C1 |
H4 |
107.956 |
| C2 |
C1 |
H5 |
108.463 |
|
O3 |
C1 |
H4 |
112.714 |
| O3 |
C1 |
H5 |
106.982 |
|
H4 |
C1 |
H5 |
107.399 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.073 |
|
|
|
| 2 |
C |
-0.116 |
|
|
|
| 3 |
O |
-0.301 |
|
|
|
| 4 |
H |
0.084 |
|
|
|
| 5 |
H |
0.109 |
|
|
|
| 6 |
H |
0.205 |
|
|
|
| 7 |
N |
-0.054 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.516 |
1.128 |
1.261 |
3.031 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.561 |
1.226 |
-1.828 |
| y |
1.226 |
-20.511 |
-1.718 |
| z |
-1.828 |
-1.718 |
-21.734 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.438 |
1.226 |
-1.828 |
| y |
1.226 |
6.137 |
-1.718 |
| z |
-1.828 |
-1.718 |
4.302 |
|
| Polar |
|---|
| 3z2-r2 | 8.604 |
|---|
| x2-y2 | -11.050 |
|---|
| xy | 1.226 |
|---|
| xz | -1.828 |
|---|
| yz | -1.718 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.863 |
-0.412 |
-0.035 |
| y |
-0.412 |
4.074 |
-0.094 |
| z |
-0.035 |
-0.094 |
3.550 |
<r2> (average value of r
2) Å
2
| <r2> |
78.108 |
| (<r2>)1/2 |
8.838 |
