return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (2-Propen-1-ol)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-192.797511
Energy at 298.15K-192.803784
HF Energy-191.996260
Nuclear repulsion energy117.329199
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3881 3684 34.26      
2 A 3284 3117 9.44      
3 A 3222 3059 2.80      
4 A 3200 3038 7.45      
5 A 3087 2930 35.06      
6 A 3051 2896 47.25      
7 A 1722 1635 0.95      
8 A 1539 1462 1.73      
9 A 1494 1418 14.72      
10 A 1440 1367 3.77      
11 A 1322 1255 0.49      
12 A 1300 1234 11.92      
13 A 1250 1186 52.34      
14 A 1181 1121 16.03      
15 A 1093 1037 88.54      
16 A 1047 994 20.14      
17 A 983 933 11.93      
18 A 973 924 32.13      
19 A 937 890 1.37      
20 A 673 639 5.27      
21 A 450 427 3.53      
22 A 338 321 9.61      
23 A 254 241 112.50      
24 A 113 107 1.85      

Unscaled Zero Point Vibrational Energy (zpe) 18915.5 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 17958.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.91883 0.14619 0.14102

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.583 0.447 0.298
C2 -0.662 -0.360 0.266
C3 -1.814 0.080 -0.227
O4 1.605 -0.298 -0.349
H5 0.409 1.403 -0.196
H6 0.856 0.648 1.335
H7 -0.591 -1.352 0.682
H8 -2.701 -0.529 -0.211
H9 -1.891 1.065 -0.658
H10 2.439 0.146 -0.183

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.48462.48131.42061.08981.09102.18313.46392.72291.9405
C21.48461.32812.34972.11412.11301.07792.10092.09623.1738
C32.48131.32813.44172.58693.14542.09101.07591.07774.2535
O41.42062.34973.44172.08482.07172.64554.31423.76430.9593
H51.08982.11412.58692.08481.76513.05993.66092.36952.3879
H61.09102.11303.14542.07171.76512.55394.05343.41922.2494
H72.18311.07792.09102.64553.05992.55392.43453.05383.4891
H83.46392.10091.07594.31423.66094.05342.43451.84275.1840
H92.72292.09621.07773.76432.36953.41923.05381.84274.4514
H101.94053.17384.25350.95932.38792.24943.48915.18404.4514

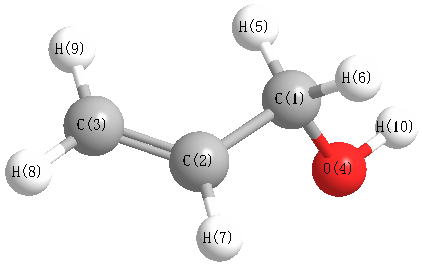
picture of 2-Propen-1-ol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.717 C1 C2 H7 115.935
C1 O4 H10 107.667 C2 C1 O4 107.932
C2 C1 H5 109.453 C2 C1 H6 109.294
C2 C3 H8 121.472 C2 C3 H9 120.866
C3 C2 H7 120.346 O4 C1 H5 111.616
O4 C1 H6 110.456 H5 C1 H6 108.066
H8 C3 H9 117.662
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability