Jump to
S1C2
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -612.714625 |
| Energy at 298.15K | |
| HF Energy | -611.906131 |
| Nuclear repulsion energy | 142.919527 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
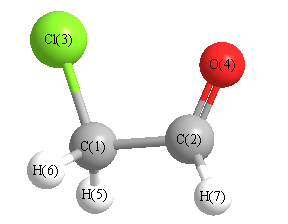
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.017 |
0.687 |
0.138 |
| C2 |
1.158 |
-0.310 |
0.225 |
| Cl3 |
-1.536 |
-0.174 |
-0.083 |
| O4 |
2.252 |
-0.083 |
-0.237 |
| H5 |
-0.047 |
1.250 |
1.070 |
| H6 |
0.176 |
1.365 |
-0.696 |
| H7 |
0.924 |
-1.251 |
0.752 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5169 | 1.7899 | 2.3927 | 1.0904 | 1.0864 | 2.2261 |
C2 | 1.5169 | | 2.7147 | 1.2086 | 2.1440 | 2.1485 | 1.1042 | Cl3 | 1.7899 | 2.7147 | | 3.7919 | 2.3609 | 2.3822 | 2.8127 | O4 | 2.3927 | 1.2086 | 3.7919 | | 2.9608 | 2.5724 | 2.0258 | H5 | 1.0904 | 2.1440 | 2.3609 | 2.9608 | | 1.7834 | 2.7015 | H6 | 1.0864 | 2.1485 | 2.3822 | 2.5724 | 1.7834 | | 3.0825 | H7 | 2.2261 | 1.1042 | 2.8127 | 2.0258 | 2.7015 | 3.0825 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
122.374 |
|
C1 |
C2 |
H7 |
115.376 |
| C2 |
C1 |
Cl3 |
110.089 |
|
C2 |
C1 |
H5 |
109.551 |
| C2 |
C1 |
H6 |
110.142 |
|
Cl3 |
C1 |
H5 |
107.613 |
| Cl3 |
C1 |
H6 |
109.381 |
|
O4 |
C2 |
H7 |
122.244 |
| H5 |
C1 |
H6 |
110.025 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -612.712790 |
| Energy at 298.15K | -612.716226 |
| HF Energy | -611.903475 |
| Nuclear repulsion energy | 146.781203 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3068 |
2991 |
|
|
|
|
| 2 |
A' |
2918 |
2845 |
|
|
|
|
| 3 |
A' |
1809 |
1764 |
|
|
|
|
| 4 |
A' |
1448 |
1411 |
|
|
|
|
| 5 |
A' |
1410 |
1374 |
|
|
|
|
| 6 |
A' |
1310 |
1277 |
|
|
|
|
| 7 |
A' |
957 |
933 |
|
|
|
|
| 8 |
A' |
777 |
758 |
|
|
|
|
| 9 |
A' |
637 |
621 |
|
|
|
|
| 10 |
A' |
211 |
205 |
|
|
|
|
| 11 |
A" |
3119 |
3041 |
|
|
|
|
| 12 |
A" |
1209 |
1179 |
|
|
|
|
| 13 |
A" |
1033 |
1007 |
|
|
|
|
| 14 |
A" |
696 |
678 |
|
|
|
|
| 15 |
A" |
157 |
153 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10379.9 cm
-1
Scaled (by 0.9748) Zero Point Vibrational Energy (zpe) 10118.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.925 |
0.000 |
| C2 |
1.365 |
0.268 |
0.000 |
| Cl3 |
-1.344 |
-0.232 |
0.000 |
| O4 |
1.579 |
-0.918 |
0.000 |
| H5 |
-0.080 |
1.562 |
0.883 |
| H6 |
-0.080 |
1.562 |
-0.883 |
| H7 |
2.190 |
1.007 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5150 | 1.7735 | 2.4272 | 1.0914 | 1.0914 | 2.1918 |
C2 | 1.5150 | | 2.7543 | 1.2051 | 2.1314 | 2.1314 | 1.1082 | Cl3 | 1.7735 | 2.7543 | | 3.0020 | 2.3655 | 2.3655 | 3.7450 | O4 | 2.4272 | 1.2051 | 3.0020 | | 3.1120 | 3.1120 | 2.0201 | H5 | 1.0914 | 2.1314 | 2.3655 | 3.1120 | | 1.7652 | 2.4985 | H6 | 1.0914 | 2.1314 | 2.3655 | 3.1120 | 1.7652 | | 2.4985 | H7 | 2.1918 | 1.1082 | 3.7450 | 2.0201 | 2.4985 | 2.4985 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
125.956 |
|
C1 |
C2 |
H7 |
112.423 |
| C2 |
C1 |
Cl3 |
113.532 |
|
C2 |
C1 |
H5 |
108.634 |
| C2 |
C1 |
H6 |
108.634 |
|
Cl3 |
C1 |
H5 |
108.981 |
| Cl3 |
C1 |
H6 |
108.981 |
|
O4 |
C2 |
H7 |
121.620 |
| H5 |
C1 |
H6 |
107.936 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
