Jump to
S1C2
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -613.490540 |
| Energy at 298.15K | -613.493786 |
| HF Energy | -613.490540 |
| Nuclear repulsion energy | 143.742706 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3176 |
3040 |
0.48 |
|
|
|
| 2 |
A |
3096 |
2963 |
8.58 |
|
|
|
| 3 |
A |
2947 |
2821 |
61.93 |
|
|
|
| 4 |
A |
1855 |
1775 |
171.84 |
|
|
|
| 5 |
A |
1443 |
1381 |
12.52 |
|
|
|
| 6 |
A |
1402 |
1342 |
4.74 |
|
|
|
| 7 |
A |
1260 |
1206 |
15.59 |
|
|
|
| 8 |
A |
1191 |
1140 |
4.39 |
|
|
|
| 9 |
A |
1056 |
1011 |
37.28 |
|
|
|
| 10 |
A |
1031 |
987 |
2.96 |
|
|
|
| 11 |
A |
820 |
785 |
27.12 |
|
|
|
| 12 |
A |
699 |
669 |
10.28 |
|
|
|
| 13 |
A |
463 |
444 |
9.90 |
|
|
|
| 14 |
A |
273 |
261 |
11.32 |
|
|
|
| 15 |
A |
53 |
50 |
15.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10382.5 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 9937.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.021 |
0.665 |
0.124 |
| C2 |
1.153 |
-0.330 |
0.177 |
| Cl3 |
-1.538 |
-0.163 |
-0.071 |
| O4 |
2.257 |
-0.062 |
-0.194 |
| H5 |
-0.013 |
1.217 |
1.061 |
| H6 |
0.168 |
1.357 |
-0.697 |
| H7 |
0.893 |
-1.318 |
0.596 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5076 | 1.7766 | 2.3720 | 1.0881 | 1.0838 | 2.2166 |
C2 | 1.5076 | | 2.7075 | 1.1950 | 2.1293 | 2.1399 | 1.1043 | Cl3 | 1.7766 | 2.7075 | | 3.7982 | 2.3485 | 2.3690 | 2.7723 | O4 | 2.3720 | 1.1950 | 3.7982 | | 2.8917 | 2.5749 | 2.0156 | H5 | 1.0881 | 2.1293 | 2.3485 | 2.8917 | | 1.7729 | 2.7318 | H6 | 1.0838 | 2.1399 | 2.3690 | 2.5749 | 1.7729 | | 3.0580 | H7 | 2.2166 | 1.1043 | 2.7723 | 2.0156 | 2.7318 | 3.0580 | |
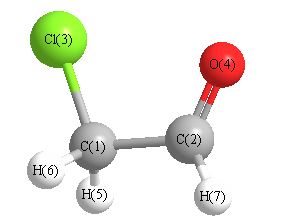 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
122.311 |
|
C1 |
C2 |
H7 |
115.259 |
| C2 |
C1 |
Cl3 |
110.787 |
|
C2 |
C1 |
H5 |
109.171 |
| C2 |
C1 |
H6 |
110.266 |
|
Cl3 |
C1 |
H5 |
107.694 |
| Cl3 |
C1 |
H6 |
109.435 |
|
O4 |
C2 |
H7 |
122.425 |
| H5 |
C1 |
H6 |
109.440 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.138 |
|
|
|
| 2 |
C |
0.136 |
|
|
|
| 3 |
Cl |
-0.143 |
|
|
|
| 4 |
O |
-0.236 |
|
|
|
| 5 |
H |
0.149 |
|
|
|
| 6 |
H |
0.152 |
|
|
|
| 7 |
H |
0.080 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.821 |
0.603 |
0.907 |
1.364 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.577 |
-0.275 |
1.254 |
| y |
-0.275 |
-28.190 |
0.023 |
| z |
1.254 |
0.023 |
-29.082 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.941 |
-0.275 |
1.254 |
| y |
-0.275 |
5.639 |
0.023 |
| z |
1.254 |
0.023 |
4.302 |
|
| Polar |
|---|
| 3z2-r2 | 8.603 |
|---|
| x2-y2 | -10.387 |
|---|
| xy | -0.275 |
|---|
| xz | 1.254 |
|---|
| yz | 0.023 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.350 |
0.459 |
-0.133 |
| y |
0.459 |
5.101 |
-0.205 |
| z |
-0.133 |
-0.205 |
4.117 |
<r2> (average value of r
2) Å
2
| <r2> |
121.203 |
| (<r2>)1/2 |
11.009 |
Jump to
S1C1
Energy calculated at B1B95/cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -613.489124 |
| Energy at 298.15K | -613.492585 |
| HF Energy | -613.489124 |
| Nuclear repulsion energy | 147.819708 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3078 |
2946 |
19.30 |
|
|
|
| 2 |
A' |
2897 |
2773 |
101.38 |
|
|
|
| 3 |
A' |
1873 |
1792 |
150.42 |
|
|
|
| 4 |
A' |
1432 |
1371 |
28.10 |
|
|
|
| 5 |
A' |
1407 |
1346 |
13.98 |
|
|
|
| 6 |
A' |
1302 |
1246 |
15.89 |
|
|
|
| 7 |
A' |
954 |
913 |
6.30 |
|
|
|
| 8 |
A' |
783 |
750 |
8.14 |
|
|
|
| 9 |
A' |
644 |
616 |
45.32 |
|
|
|
| 10 |
A' |
204 |
196 |
2.98 |
|
|
|
| 11 |
A" |
3125 |
2991 |
0.44 |
|
|
|
| 12 |
A" |
1208 |
1156 |
1.46 |
|
|
|
| 13 |
A" |
1035 |
990 |
0.57 |
|
|
|
| 14 |
A" |
684 |
655 |
1.80 |
|
|
|
| 15 |
A" |
180 |
172 |
1.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10402.6 cm
-1
Scaled (by 0.9571) Zero Point Vibrational Energy (zpe) 9956.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.914 |
0.000 |
| C2 |
1.356 |
0.266 |
0.000 |
| Cl3 |
-1.337 |
-0.232 |
0.000 |
| O4 |
1.571 |
-0.907 |
0.000 |
| H5 |
-0.080 |
1.554 |
0.878 |
| H6 |
-0.080 |
1.554 |
-0.878 |
| H7 |
2.179 |
1.008 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5028 | 1.7603 | 2.4047 | 1.0891 | 1.0891 | 2.1809 |
C2 | 1.5028 | | 2.7379 | 1.1919 | 2.1193 | 2.1193 | 1.1085 | Cl3 | 1.7603 | 2.7379 | | 2.9851 | 2.3529 | 2.3529 | 3.7277 | O4 | 2.4047 | 1.1919 | 2.9851 | | 3.0903 | 3.0903 | 2.0089 | H5 | 1.0891 | 2.1193 | 2.3529 | 3.0903 | | 1.7556 | 2.4842 | H6 | 1.0891 | 2.1193 | 2.3529 | 3.0903 | 1.7556 | | 2.4842 | H7 | 2.1809 | 1.1085 | 3.7277 | 2.0089 | 2.4842 | 2.4842 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
125.961 |
|
C1 |
C2 |
H7 |
112.393 |
| C2 |
C1 |
Cl3 |
113.852 |
|
C2 |
C1 |
H5 |
108.650 |
| C2 |
C1 |
H6 |
108.650 |
|
Cl3 |
C1 |
H5 |
109.045 |
| Cl3 |
C1 |
H6 |
109.045 |
|
O4 |
C2 |
H7 |
121.646 |
| H5 |
C1 |
H6 |
107.403 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.145 |
|
|
|
| 2 |
C |
0.117 |
|
|
|
| 3 |
Cl |
-0.115 |
|
|
|
| 4 |
O |
-0.219 |
|
|
|
| 5 |
H |
0.142 |
|
|
|
| 6 |
H |
0.142 |
|
|
|
| 7 |
H |
0.078 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.517 |
3.365 |
0.000 |
3.404 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.117 |
3.676 |
0.000 |
| y |
3.676 |
-29.446 |
0.000 |
| z |
0.000 |
0.000 |
-29.133 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.827 |
3.676 |
0.000 |
| y |
3.676 |
1.178 |
0.000 |
| z |
0.000 |
0.000 |
1.648 |
|
| Polar |
|---|
| 3z2-r2 | 3.297 |
|---|
| x2-y2 | -2.670 |
|---|
| xy | 3.676 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.603 |
0.583 |
0.000 |
| y |
0.583 |
6.039 |
0.000 |
| z |
0.000 |
0.000 |
3.906 |
<r2> (average value of r
2) Å
2
| <r2> |
105.096 |
| (<r2>)1/2 |
10.252 |
