return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6S (Thiirane, methyl-)

using model chemistry: MP2=FULL/cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/cc-pVTZ
 hartrees
Energy at 0K-515.438243
Energy at 298.15K-515.444973
HF Energy-514.662664
Nuclear repulsion energy161.184084
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3265 3100 1.58      
2 A 3206 3044 8.48      
3 A 3181 3020 9.77      
4 A 3174 3013 8.06      
5 A 3157 2998 8.87      
6 A 3085 2929 20.87      
7 A 1522 1445 6.54      
8 A 1511 1435 3.79      
9 A 1499 1423 5.81      
10 A 1429 1357 6.34      
11 A 1395 1325 5.41      
12 A 1217 1156 2.06      
13 A 1203 1142 1.47      
14 A 1117 1060 13.41      
15 A 1074 1020 7.86      
16 A 1035 982 3.22      
17 A 951 903 3.10      
18 A 939 891 1.79      
19 A 907 861 2.20      
20 A 675 641 4.47      
21 A 651 618 13.78      
22 A 388 368 0.12      
23 A 311 295 1.46      
24 A 252 239 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 18570.7 cm-1
Scaled (by 0.9494) Zero Point Vibrational Energy (zpe) 17631.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/cc-pVTZ
ABC
0.39265 0.16465 0.13356

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/cc-pVTZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.760 -0.297 -0.179
C2 -0.510 0.155 0.504
C3 0.375 1.138 -0.149
S4 1.079 -0.523 -0.056
H5 -1.609 -0.381 -1.251
H6 -2.090 -1.263 0.192
H7 -2.555 0.424 0.001
H8 -0.590 0.230 1.577
H9 0.101 1.486 -1.134
H10 0.859 1.897 0.447

Atom - Atom Distances (Å)
  C1 C2 C3 S4 H5 H6 H7 H8 H9 H10
C11.49412.57242.85051.08571.08601.08812.17472.74843.4734
C21.49411.47451.81612.13852.14572.12251.07882.19652.2161
C32.57241.47451.80662.73103.45763.01922.17541.07991.0793
S42.85051.81611.80662.94493.26343.75562.45322.48112.4812
H51.08572.13852.73102.94491.75771.76383.06712.53483.7635
H61.08602.14573.45763.26341.75771.76002.52923.75684.3297
H71.08812.12253.01923.75561.76381.76002.52593.07733.7450
H82.17471.07882.17542.45323.06712.52922.52593.06612.4811
H92.74842.19651.07992.48112.53483.75683.07733.06611.8002
H103.47342.21611.07932.48123.76354.32973.74502.48111.8002

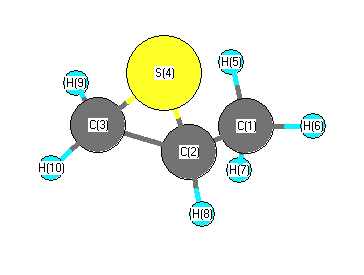
picture of Thiirane, methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.115 C1 C2 S4 118.560
C1 C2 H8 114.425 C2 C1 H5 110.986
C2 C1 H6 111.555 C2 C1 H7 109.564
C2 C3 S4 66.323 C2 C3 H9 117.779
C2 C3 H10 119.606 C2 S4 C3 48.033
C3 C2 S4 65.644 C3 C2 H8 115.995
S4 C2 H8 113.396 S4 C3 H9 116.247
S4 C3 H10 116.302 H5 C1 H6 108.068
H5 C1 H7 108.467 H6 C1 H7 108.101
H9 C3 H10 112.962
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability